




-
The genus Hedychium (Zingiberaceae) comprises approximately 80 species of perennial herbs, with a major center of diversity in the Himalayan region and a natural distribution extending across tropical and subtropical Asia[1,2]. Valued for their striking floral morphology, rich fragrance, and diverse biological activities, Hedychium species have been widely introduced and cultivated worldwide, with over 110 cultivars developed for ornamental, medicinal, and aromatic applications[3]. However, taxonomic delineation of the genus based on traditional morphology remains contentious, largely because of ambiguous interspecific morphological boundaries and frequent intra- and interspecific hybridization. These complexities have resulted in unclear species limits and uncertain genetic origins for many cultivars[4−6]. Previous molecular studies using markers such as isozymes, random amplified polymorphic DNA (RAPD), inter-simple sequence repeat (ISSR), and simple sequence repeats (SSRs) have provided preliminary insights into the genetic diversity of Hedychium[7−12]. Nevertheless, the low resolution and limited genome-wide coverage of these conventional markers have constrained a deeper understanding of the population's genetic structure and complex breeding history, particularly within the cultivated gene pool. Consequently, several fundamental questions remain unresolved: (1) Which sampled Hedychium species show close genetic affinities with the major groups of modern Hedychium cultivars; (2) how admixture patterns potentially associated with historical hybridization are reflected in the contemporary genetic architecture of modern Hedychium cultivars; and (3) whether high-density genomic markers can resolve genetic relationships among major germplasm resources and clarify the placement of unidentified accessions.
Restriction site-associated DNA sequencing (RAD-seq) has emerged as an efficient approach for developing genome-wide molecular markers, offering high throughput, accuracy, and rapid turnaround[13,14]. By generating high-density single-nucleotide polymorphism (SNP) datasets, RAD-seq enables robust analyses of population structure, phylogenetic relationships, and genetic admixture, making it particularly suitable for elucidating complex breeding histories[15]. This method has also been widely used to investigate admixture, introgression, and allele sharing patterns in nonmodel plant groups[16,17]. Despite its advantages, a comprehensive RAD-seq-based population genomic study examining genetic structure and genetic affinities between species accessions and cultivated Hedychium germplasm is still lacking.
In this study, we initially collected 168 individual samples representing 35 Hedychium germplasm accessions and applied RAD-seq to generate a comprehensive set of polymorphic SNP loci. Using these high-quality SNPs, we conducted phylogenetic reconstruction, principal component analysis (PCA), population structure inference, and D-statistics analysis to elucidate the genetic relationships, population subdivision, and asymmetric allele sharing among Hedychium accessions. Our objectives were to identify Hedychium species showing close genetic affinities with the major cultivated groups, characterize admixture patterns in modern cultivars and assess the relationships of these patterns with known or possible hybrid origins, and clarify the genetic placement of major and unidentified germplasm resources. The findings from this study will provide a crucial molecular foundation for the taxonomic clarification, breeding, conservation, and utilization of Hedychium germplasm.
-
In total, 35 Hedychium germplasm accessions were selected for this study, comprising eight species accessions, eighteen cultivars, five artificial hybrids, and four accessions of unknown identity. These accessions, originating from China, the USA and Thailand, are maintained in the Zingiberales Garden at the Xiamen Botanical Garden (Supplementary Table S1). Accessions with uncertain identities, including Unknown 1–4 and provisional Hedychium sp. materials, were retained in the analysis to evaluate their genetic placement within the broader germplasm collection; their available background information is provided in Supplementary Table S1. From these accessions, 168 individual plant samples were initially collected, with six individual plants sampled for most accessions and one to five for those with limited availability. The 'Orange Brush' cultivar was an exception, with 11 individual plants collected. Approximately 0.5 g of healthy young leaf tissue was sampled from each plant, rapidly frozen in liquid nitrogen, and stored at −80 °C for subsequent DNA extraction.
Genomic DNA extraction and quality assessment
-
Total genomic DNA was isolated from the Hedychium leaf samples using a plant DNA extraction kit (DNA BR). The integrity of the extracted DNA was verified by electrophoresis on a 1% agarose gel. DNA concentration was measured using either a Qubit Fluorometer or a microplate reader. Only DNA samples meeting quality standards were used for subsequent library construction.
Library construction and sequencing
-
Library preparation and sequencing were performed by BGI-Shenzhen (Shenzhen, China). The specific procedure was as follows: Genomic DNA (0.1–1 μg in a 50-μL reaction volume) was digested with the EcoRI enzyme (20 U per reaction) at 37 °C for 15 min, followed by ligation to a Solexa P1 adapter containing a sample-specific barcode. The barcoded samples were then pooled and physically sheared to fragment sizes ranging from 300 to 500 bp. Target fragments within this size range were recovered by excision from a 1% agarose gel. The recovered fragments underwent end repair, A-tailing, and ligation to a Y-shaped P2 adapter. A 5-μL aliquot of the ligation product was used for polymerase chain reaction (PCR) amplification. Finally, the library's fragments, sized between 350 and 550 bp, were purified via gel electrophoresis[18]. Library quality was assessed using an Agilent 2100 Bioanalyzer (Agilent Technologies, USA) and quantitative PCR (qPCR). Libraries passing quality control were subjected to sequencing.
Raw data processing and quality control
-
Raw sequencing data underwent stringent quality control to obtain high-quality clean data for downstream analysis. Reads were first demultiplexed and assigned to their respective samples according to their unique 5' barcodes (4–8 bp in length). The barcode sequences were then trimmed, and the presence of the correct EcoRI cut-site remnant (AATTC) was verified; only reads containing the correct site were retained. Further filtering removed adapter-contaminated reads, reads in which more than 40% of bases had a Phred quality score of ≤ 15, and reads containing more than 10% ambiguous bases (N)[19].
SNP calling and preprocessing
-
The demultiplexing and quality-control procedures described above were implemented using the process_radtags module of the STACKS pipeline (v1.44)[20]. Subsequently, the ustacks module was used to assemble loci within each sample, followed by cstacks to build a catalog of loci across all samples. The sstacks module was then used to match loci from each sample against the catalog. Finally, genotyping was performed using the populations module, yielding a dataset of 647,693 high-quality SNPs. This dataset was used for constructing the neighbor-joining (NJ) tree.
To address potential biases from linked loci in analyses that assume interlocus independence, a separate SNP dataset was generated for PCA and admixture analyses. Starting from the original STACKS variant call format (VCF) output, loci were filtered using a site-level genotype missingness threshold of < 50%, a minimum genotype depth of 5×, and a requirement of at least two observed alleles, yielding 1,441,743 SNPs. Linkage disequilibrium (LD) pruning was then performed using PLINK v2.0 with a sliding window of 50 SNPs, a step size of 10 SNPs, and an r2 threshold of 0.2, resulting in a final set of 265,365 LD-pruned SNPs for PCA and admixture analyses[21]. The original VCF was used as the source dataset for D-statistic analysis. Because Sample 30-6 yielded only 0.07 Gb of clean data, it was removed from both the LD-pruned dataset used for PCA and admixture analyses and the VCF used as input for D-statistic analysis. Both updated datasets therefore contained 167 samples.
Population genetic structure analysis
-
The corresponding filtered SNP datasets were used for population genetic analyses as described below. An NJ tree was constructed from the 647,693 high-quality SNP dataset using TreeBest (
https://treesoft.sourceforge.net/treebest.shtml ), based on a pairwise genetic distance matrix calculated with the p-distance method, to visualize the genetic similarity and clustering patterns among accessions. Topology robustness was assessed with 1,000 bootstrap replicates. Because RAD-seq data from a closely related external genus were not available in this study, H. villosum was used as a functional outgroup to root the NJ tree. This choice was based on a previous molecular phylogenetic study of Thai Hedychium, in which H. villosum was separated from the group containing H. coronarium, H. flavescens, H. coccineum, and H. forrestii[9]. The NJ tree was visualized and annotated using iTOL (https://itol.embl.de/ )[22], with Sample 30-6 removed from the final visualization because it had insufficient sequencing data.PCA was performed using PLINK v2.0 on the 265,365 SNPs after LD pruning. Because H. villosum (the designated outgroup) and Unknown 3 were extreme outliers that compressed the spatial distribution of the remaining samples in the preliminary analysis (Supplementary Fig. S1), these two accessions were excluded, and the final PCA was conducted on the remaining 165 samples. The PCA results were visualized using the ggplot2 package in R v4.4.1[23].
Ancestral genetic components and population structure were inferred using ADMIXTURE software on the 167 sample LD-pruned SNP dataset containing 265,365 SNPs[24]. Cross-validation (CV) error was calculated for K values from 2 to 20 to evaluate the model's fit. Because the lowest CV error at high K values may reflect fine-scale differentiation among closely related cultivars or repeated individuals rather than the major genetic groupings, the main value for biological interpretation was determined by integrating the CV error trends with the concordance of inferred components with the groupings identified by the NJ tree and PCA. The results were visualized using TBtools (version 2.357)[25].
D-statistic analysis
-
D-statistic analysis was performed using Dsuite (v0.5) to test for asymmetric allele sharing among Hedychium accessions under the ABBA-BABA framework[26−28]. The test was based on the four-taxon model (((P1, P2), P3), outgroup), in which significant deviations from equal ABBA and BABA site patterns indicate allele sharing inconsistent with a strictly bifurcating tree. Such signals may reflect historical introgression, incomplete lineage sorting, or other sources of discordance among taxa. Therefore, the D-statistic was used here as a complementary test for asymmetric allele sharing rather than as a direct estimator of the parental contribution or the hybridization direction.
To balance computational efficiency with statistical power, 100,000 biallelic SNP sites were randomly sampled from the genome-wide SNP set using BCFtools (v1.9). The DtriosParallel module in Dsuite was then used to calculate genome-wide D-statistics, Z-scores, and p-values for all possible ingroup trios (P1, P2, P3), with H. villosum used as the outgroup. H. villosum was selected because it was designated as the functional outgroup in the NJ tree analysis, whereas Unknown 3 was not used as an outgroup because of its uncertain identity and highly divergent genetic position. Test combinations involving Unknown 3 were excluded from downstream interpretation because they produced unstable statistical estimates. Results with |Z| > 3 were considered to be significant and were further interpreted as complementary evidence for asymmetric allele sharing among accessions.
-
Following demultiplexing and stringent quality control, 856.92 Gb of high-quality clean data was obtained in total from the raw sequencing data. The average output per sample was 5.1 Gb of high-quality sequence data. The average percentage of bases with a Phred quality score ≥ 30 (Q30) and the average guanine-cytosine (GC) content across all samples were 93.13% and 38.12%, respectively (Supplementary Table S2), indicating high sequencing quality suitable for subsequent bioinformatic analyses. Notably, Sample 30-6 yielded substantially less data (0.07 Gb) than other samples, suggesting a potential issue during library construction or sequencing. Because of this insufficient sequencing output, Sample 30-6 was excluded from the PCA, admixture, and D-statistic datasets and was removed from the final NJ tree visualization.
SNP detection results
-
In total, 42,212,223 raw SNPs were identified across the 168 Hedychium samples. The number of SNPs detected per sample varied widely, ranging from 4,445 (30-6) to 510,277 (4-6), with an average of 251,263 SNPs per sample. The homozygous rate across all samples ranged from 9.10% to 91.90%, with an overall average of 57.63%. Conversely, the heterozygous rate ranged from 8.10% to 90.90%, with an average of 42.37% (Supplementary Table S3). The STACKS pipeline generated a final set of 647,693 high-quality SNP loci, which was used for constructing the NJ tree. Independent filtering of the original VCF based on the genotype missing rate, sequencing depth, and the number of allele types yielded 1,441,743 SNPs. After LD pruning, 265,365 SNPs were retained for PCA and admixture analyses.
Genetic clustering reveals distinct groups and broad correspondence with floral traits
-
After Sample 30-6 was pruned, the resulting NJ tree contained 167 Hedychium samples and resolved the major genetic relationships among the retained accessions (Fig. 1). The tree was rooted using H. villosum as a functional outgroup. This choice was supported by a previous molecular phylogenetic study of Thai Hedychium, in which H. villosum was separated from the group containing H. coronarium, H. flavescens, H. coccineum, and H. forrestii. Following the outgroup, Hedychium sp. (thick-leaved) was the most genetically distinct accession among the remaining samples, separating from all other ingroup accessions as an independent lineage. Unknown 3 diverged next and exhibited the longest branch length among all accessions, suggesting that it may represent a distinct genetic lineage not fully represented in this study, potentially a species not included in the current collection. The remaining 163 samples were further grouped into eight major clusters (Clades 1–8), a pattern highly consistent with the subsequent PCA and population structure analyses.
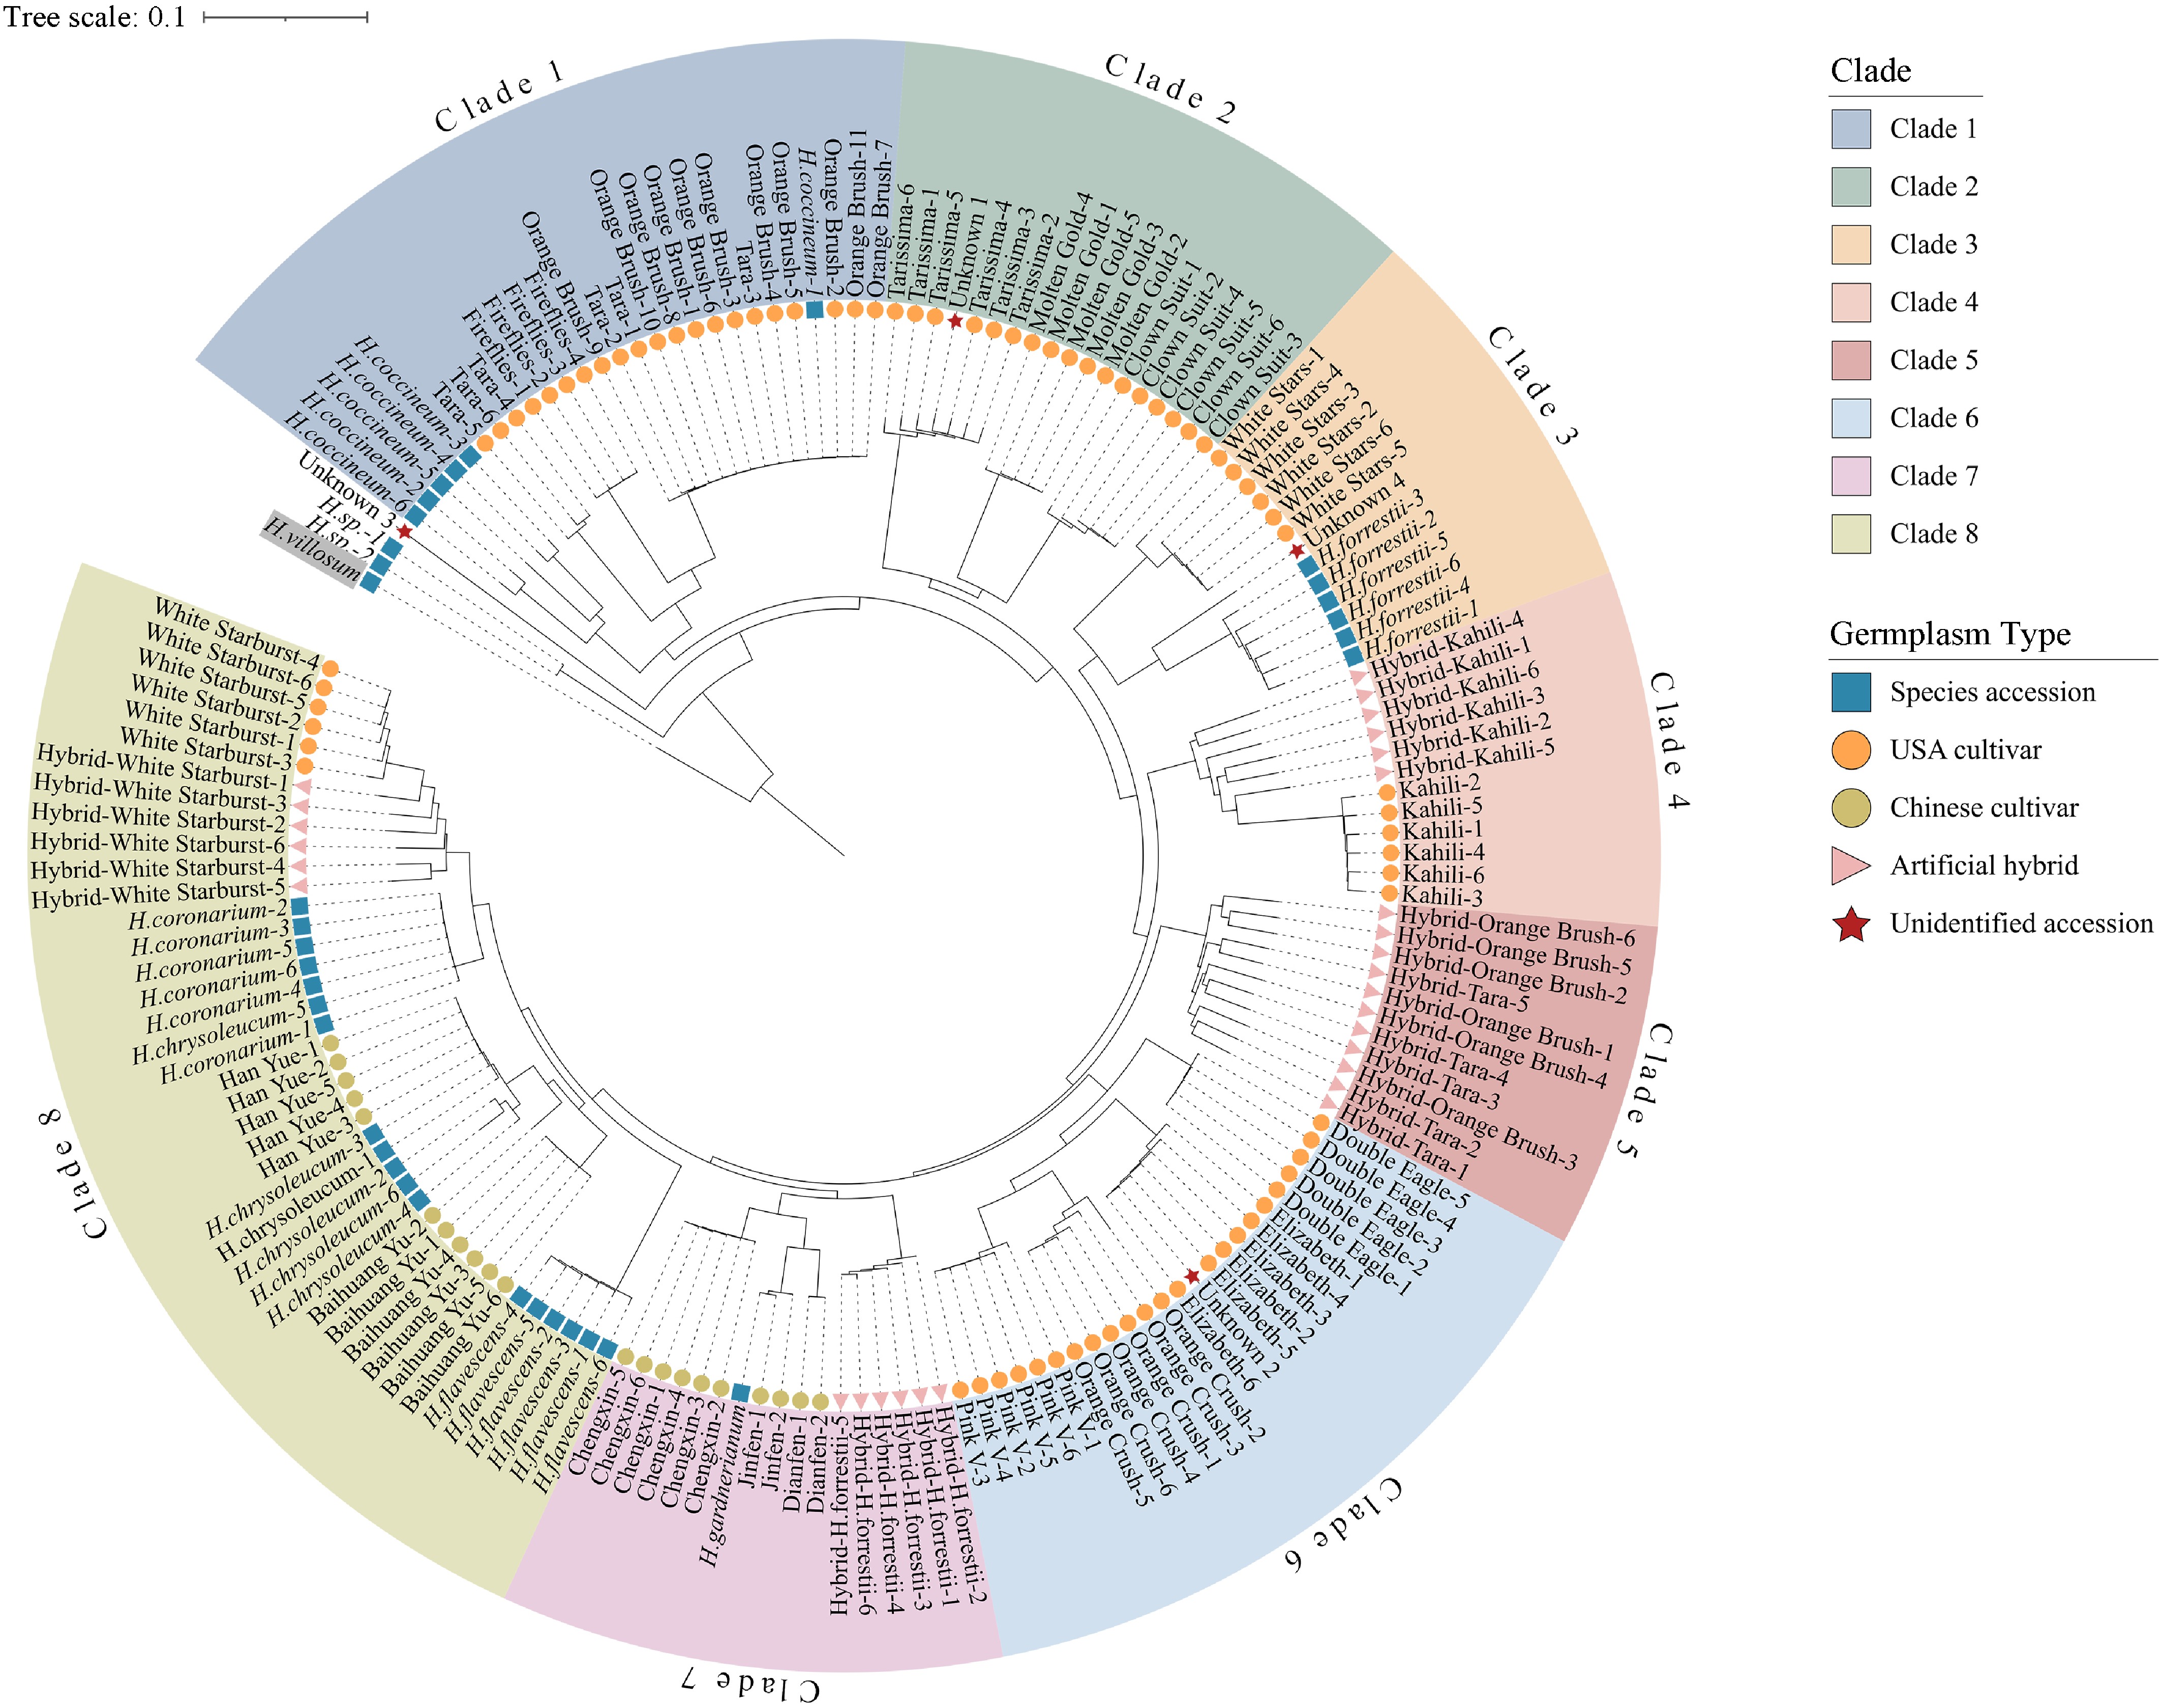
Figure 1.
Pruned NJ tree based on 647,693 high-quality SNPs. The original tree included 168 samples; Sample 30-6 was removed from the displayed topology, leaving 167 samples. The tree was rooted using H. villosum as a functional outgroup. Colors indicate the eight major clades.
The clustering patterns showed a broad correspondence with several visible ornamental traits, especially flower color and floral architecture (Supplementary Fig. S2). Clades 1 and 2 predominantly consisted of germplasms with orange-red flowers. Specifically, Clade 1 was centered on H. coccineum and clustered with several USA cultivars ('Fireflies', 'Tara', and 'Orange Brush'). Their close genetic proximity, together with their shared orange-red coloration and small petal traits, suggests a close genetic affinity between H. coccineum and these cultivars. In contrast, Clade 8 comprised a group with white or pale-yellow flowers centered on H. coronarium, including closely related species (H. flavescens and H. chrysoleucum) and associated cultivars ('Baihuang Yu', 'Han Yue'), which generally feature large petals and imbricate bracts. These observations indicate a broad descriptive correspondence between genetic clustering and visible floral traits, particularly flower color and floral architecture.
Furthermore, the phylogenetic positions of several unknown accessions provided key clues for their identification: Unknown 1 clustered closely with the cultivar 'Tarissima', Unknown 2 grouped within the same branch as 'Elizabeth', and Unknown 4 showed a sister group relationship with H. forrestii. These results provide strong evidence for close genetic affinities.
A notable observation was the intermingled clustering of individuals from the cultivars 'Tara' and 'Orange Brush' within the phylogenetic tree, where they failed to form distinct monophyletic groups. This pattern, together with their highly similar morphological characteristics, indicates an extremely close genetic relationship between these two cultivars, potentially reflecting a closely related breeding origin or a recent shared genetic background.
Population genetic structure of Hedychium germplasm elucidated by PCA
-
PCA was performed using 265,365 LD-pruned SNPs after excluding H. villosum and Unknown 3, which showed extreme divergence in the preliminary analysis of all 167 samples (Supplementary Fig. S1). In the final PCA of 165 samples, Principal Component (PC1) and PC2 explained 17.32% and 14.22% of the total genetic variation, respectively (Fig. 2). The PCA pattern largely supported the clustering pattern revealed by the NJ tree, with several major clades occupying distinguishable regions in the space defined by PC1 and PC2. In particular, Clades 1, 2, 3, 6, 7, and 8 showed clear separation to varying degrees, whereas Clades 4 and 5 were positioned close to each other near the center of the plot. Hedychium sp. (thick-leaved) occupied a clearly isolated position, further supporting its distinct genetic background relative to the other sampled accessions.
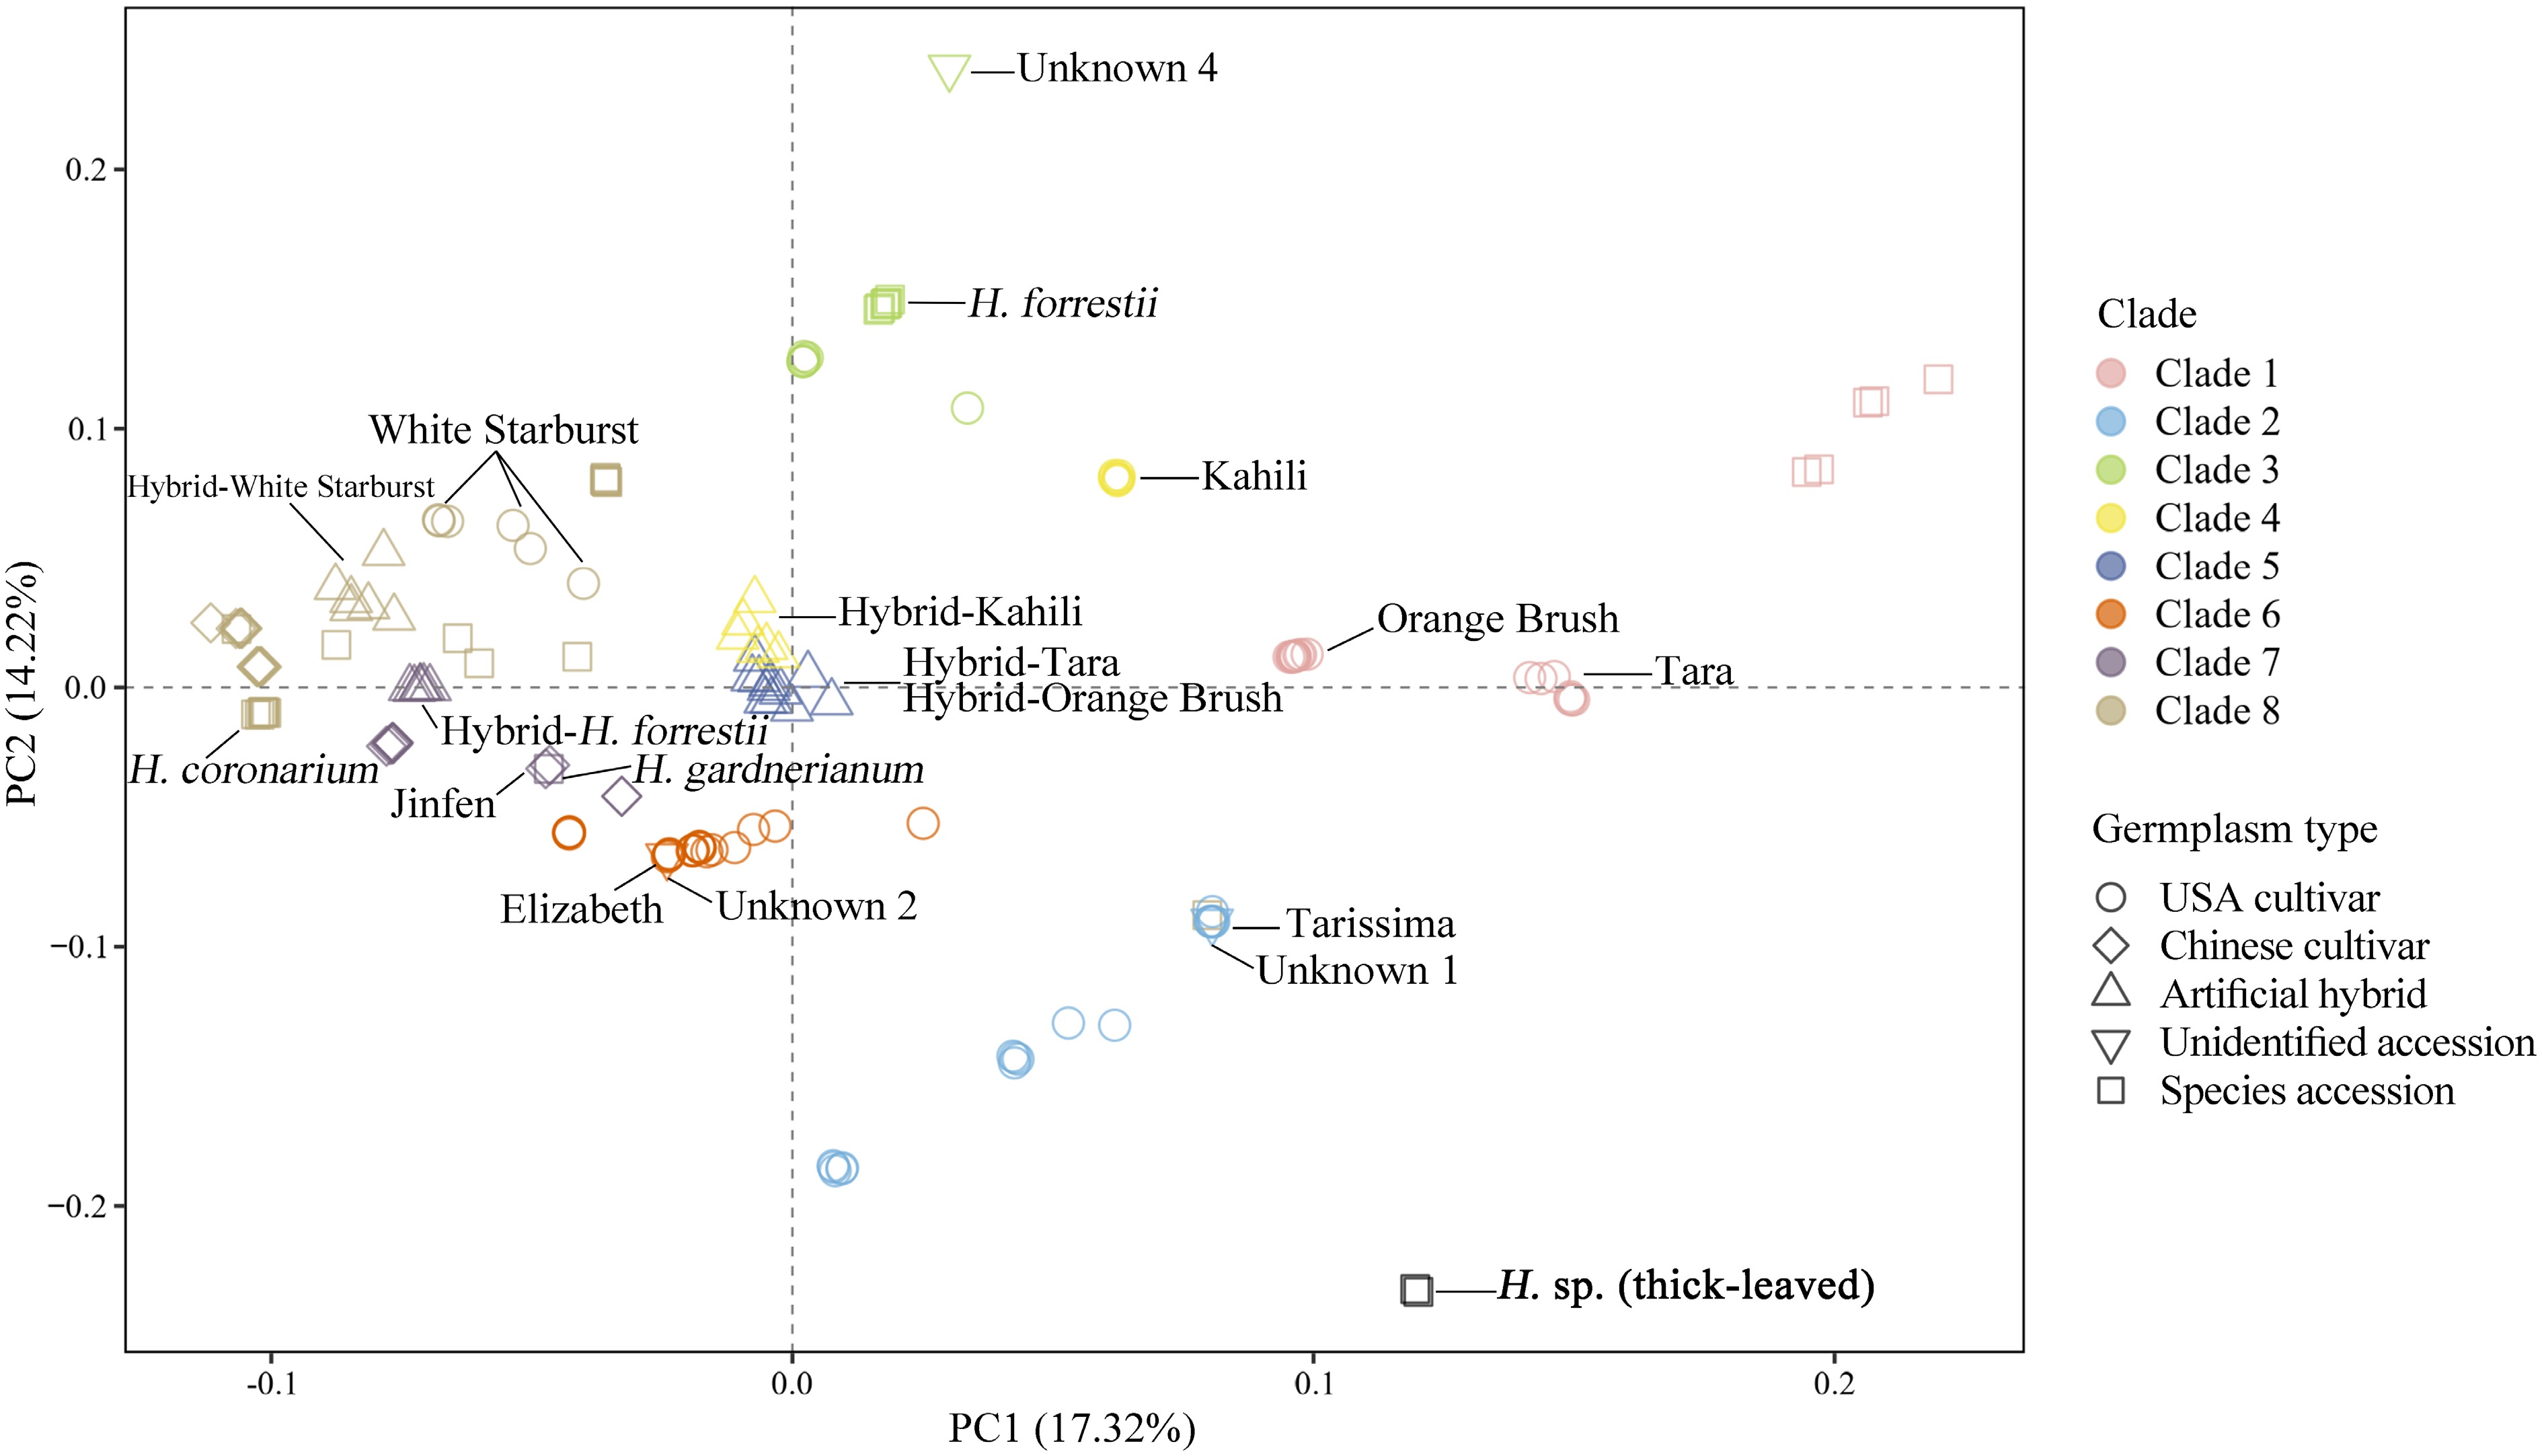
Figure 2.
PCA of 165 samples based on 265,365 SNPs after LD pruning. Colors indicate clades, and the shapes indicate germplasm types.
The PCA also provided informative evidence for hybrid and unknown accessions. The five artificial hybrid accessions were located between their corresponding parental groups, consistent with their expected hybrid origins. Unknown 1 almost overlapped with 'Tarissima', and Unknown 2 was positioned very close to 'Elizabeth', supporting their close genetic affinities with these cultivars. This genetic proximity between Unknown 2 and 'Elizabeth' is also consistent with their similar plant architecture and leaf morphology, as recorded in the living collection. Unknown 4 occupied a distinct position, although H. forrestii was the closest sampled accession in the PCA space, suggesting a possible genetic affinity rather than complete identity. In addition, H. gardnerianum and the Chinese cultivar 'Jinfen' almost overlapped in the PCA plot, suggesting a very close genetic affinity between these two accessions.
Dissecting patterns of genetic admixture in Hedychium germplasm
-
Admixture analysis was performed using the 167 samples in the LD-pruned SNP dataset. The CV error generally decreased as K increased from 2 to 20 and reached its lowest value at K = 18 (Fig. 3). However, the profiles at high K values mainly reflected fine-scale subdivisions among closely related cultivars and replicate individuals rather than the major genetic groupings. Therefore, K = 8 was used as the main value for biological interpretation, because it showed strong concordance with the eight major clades identified by the NJ tree and with the PCA results.
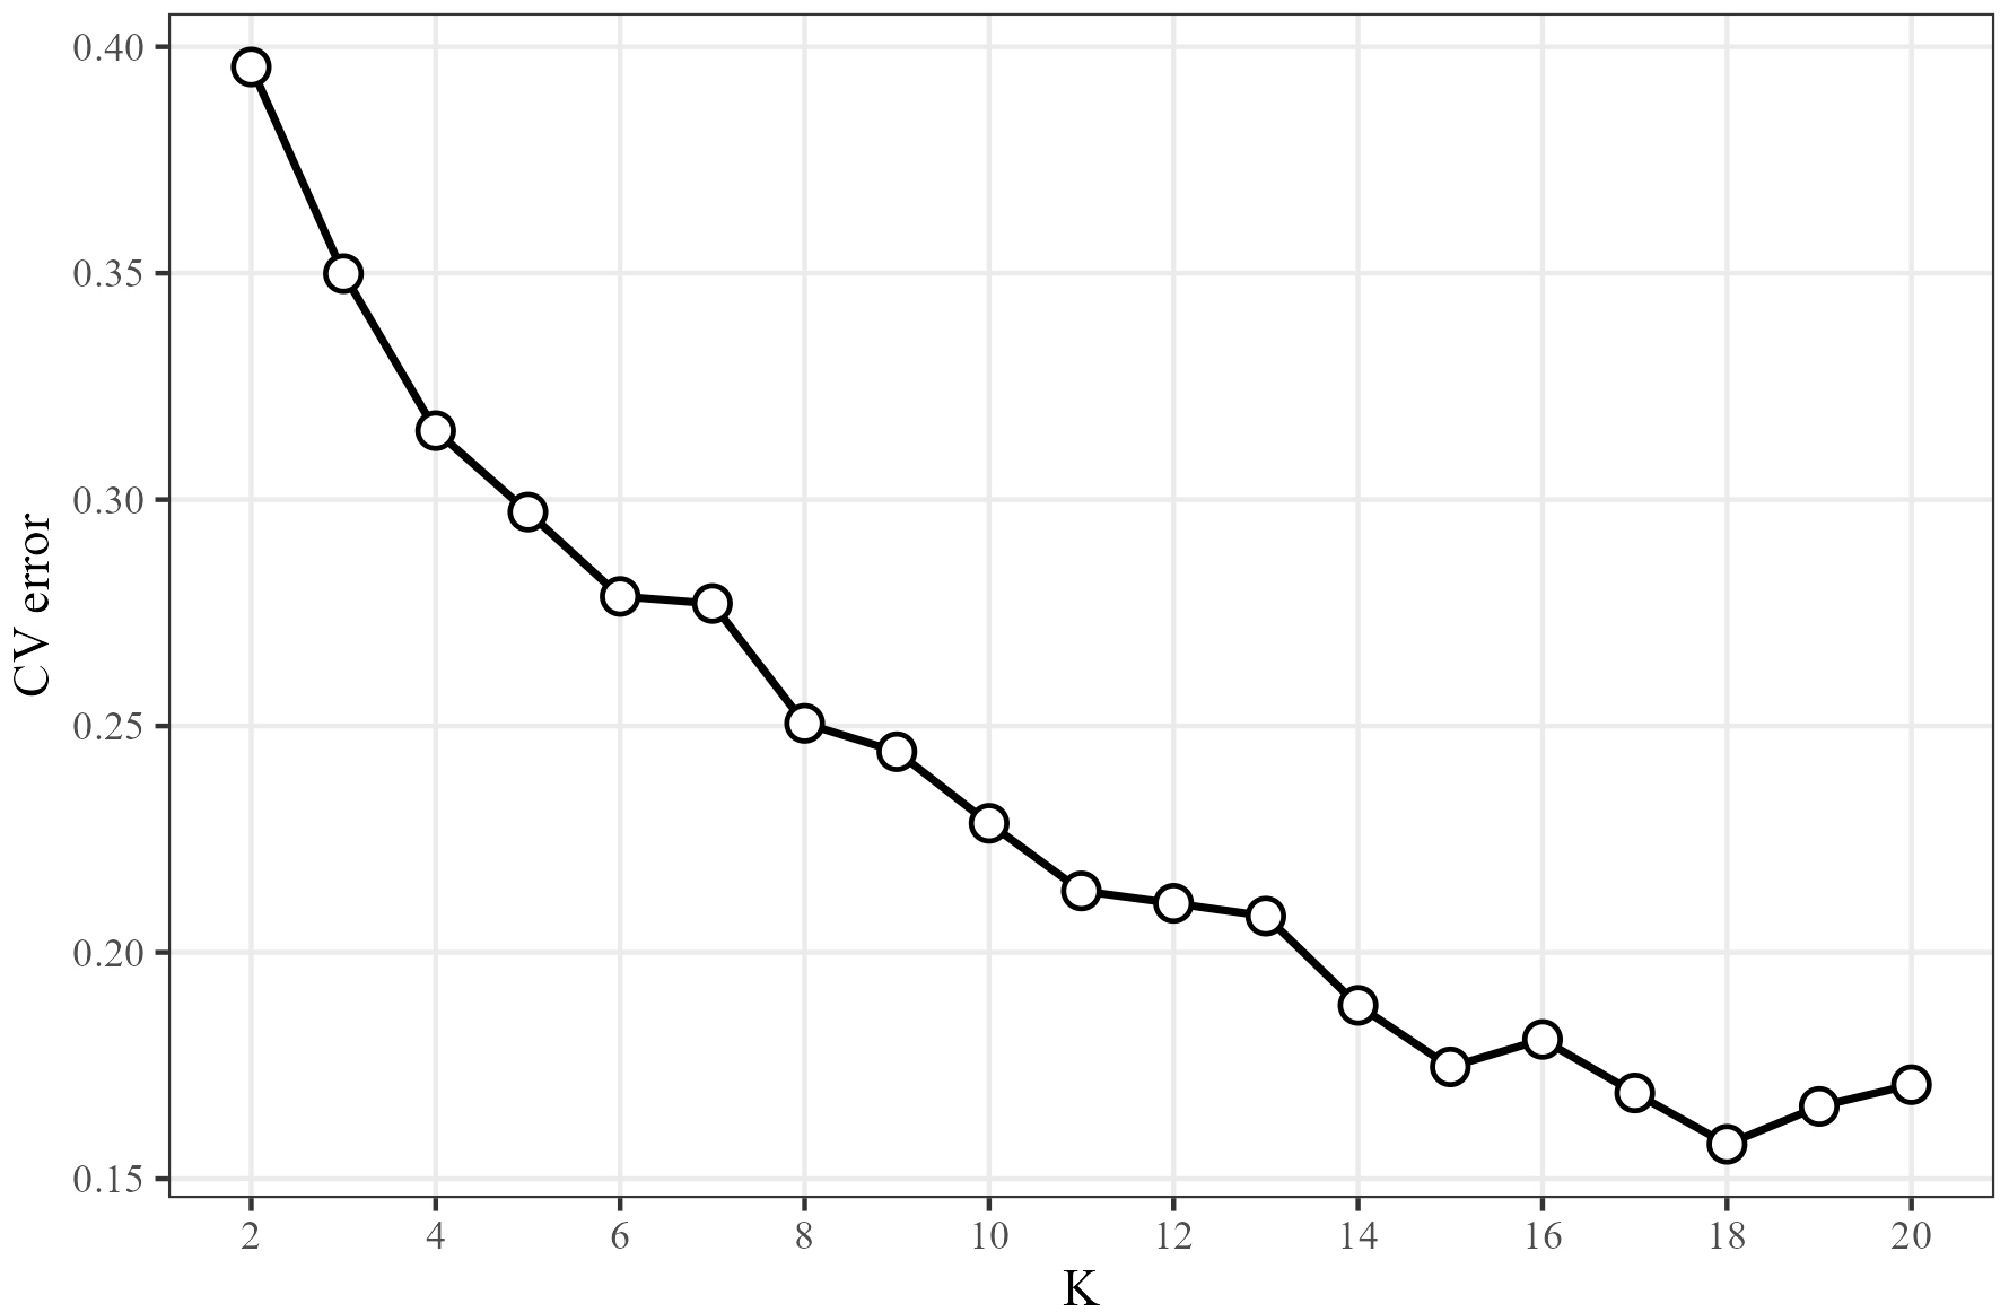
Figure 3.
CV error for different values of K.
At K = 8, most clades showed characteristic genetic component profiles, supporting the major clustering pattern revealed by the NJ tree (Fig. 4). Clade 4, consisting of 'Kahili' and hybrid 'Kahili', showed a distinct pattern in which 'Kahili' was dominated by a single major component, whereas hybrid 'Kahili' displayed a mixed profile. Clade 5, composed of hybrid 'Tara' and hybrid 'Orange Brush', was also admixed, with two major components and minor contributions from other components. These patterns are consistent with the recorded hybrid backgrounds of these materials.
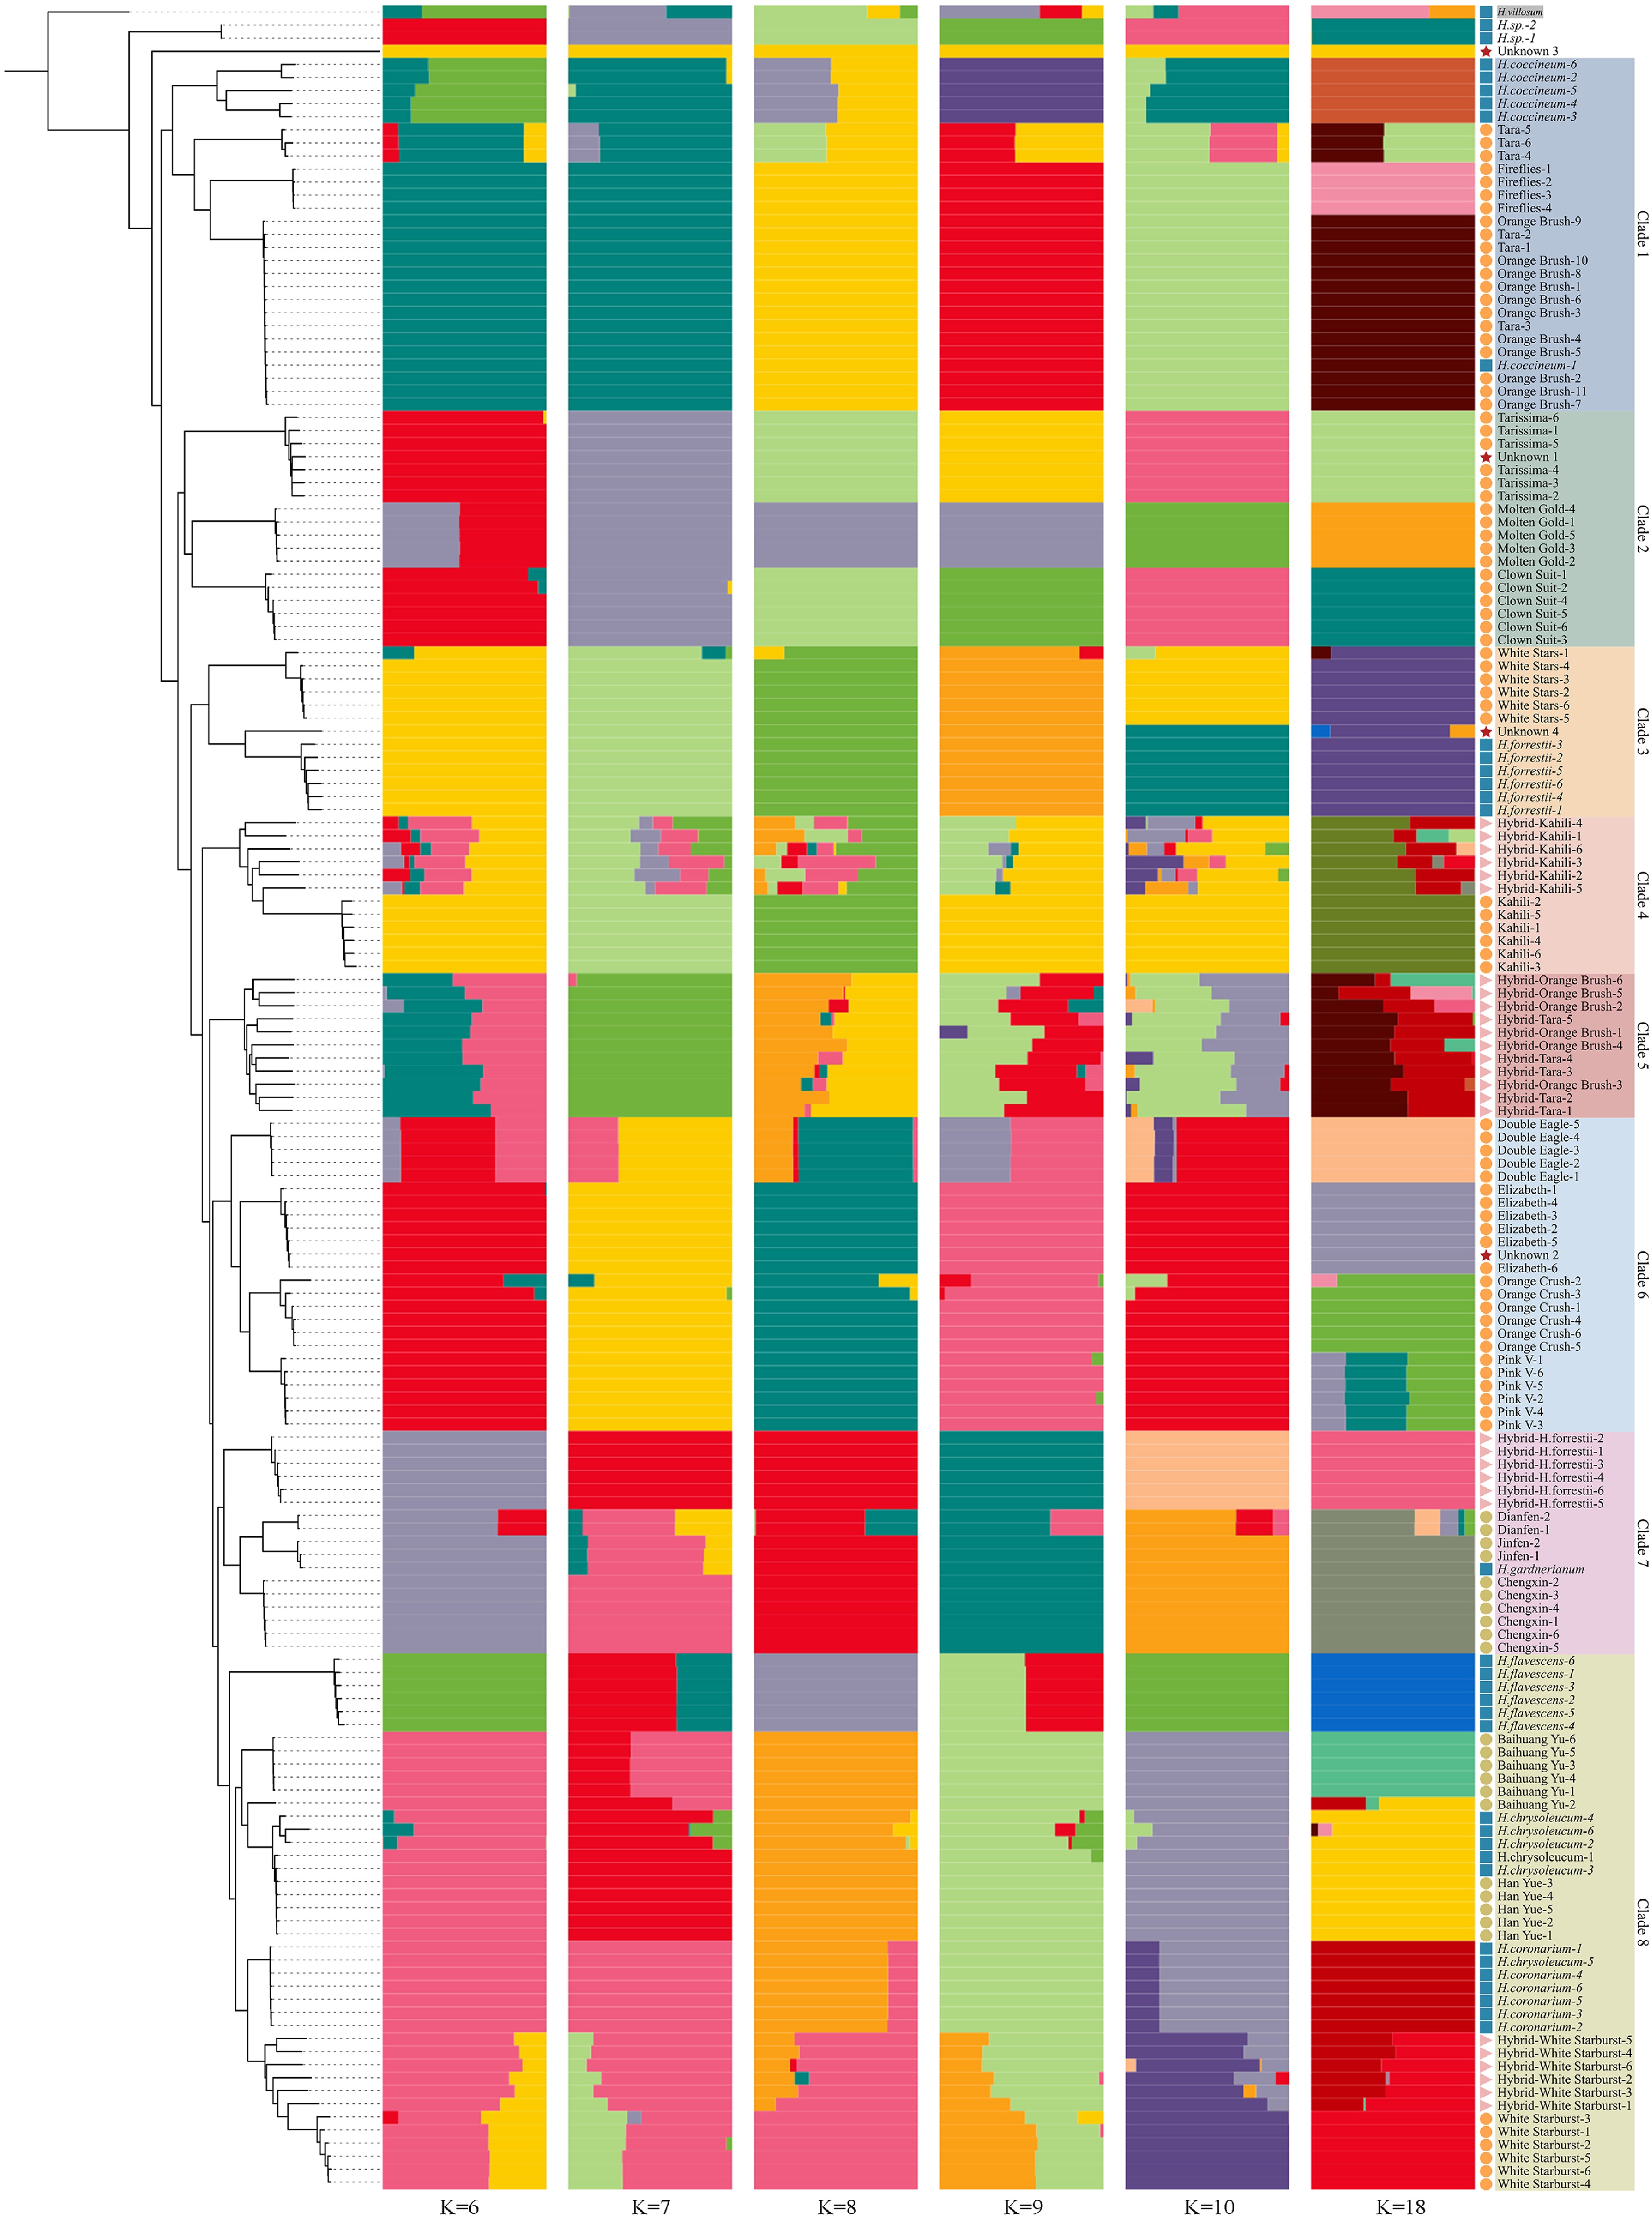
Figure 4.
Admixture results of the 167 Hedychium samples based on 265,365 SNPs after LD pruning. Results for K = 6, 7, 8, 9, 10, and 18 are shown. The sample order follows the NJ tree.
Clade 8 contained two major component patterns at K = 8. H. coronarium and materials related to 'White Starburst' were mainly characterized by one major component, whereas H. flavescens and 'Han Yue' were dominated by another component. 'Baihuang Yu' displayed a mixed profile between these two major components, whereas H. chrysoleucum showed a distinct combination of components. This pattern indicates that Clade 8 contains genetically differentiated but related white or pale-yellow flowered germplasm rather than a completely homogeneous group.
The admixture results also provided additional evidence for evaluating the genetic affinities of several unknown accessions. Unknown 1 and 'Tarissima' showed identical single-component profiles across all displayed K values, and Unknown 2 showed the same pattern as 'Elizabeth', supporting their very close genetic affinities with these cultivars. Unknown 4 exhibited the same component profile as H. forrestii at K = 6 to K = 8, whereas minor differences appeared at K = 9, K = 10, and K = 18. This pattern suggests a close genetic affinity between Unknown 4 and H. forrestii, while also indicating subtle differentiation at finer levels of population subdivision. In addition, H. gardnerianum and the Chinese cultivar 'Jinfen' exhibited highly similar component profiles, further supporting their close genetic affinity.
D-statistic analysis reveals asymmetric allele sharing among selected Hedychium accessions
-
D-statistic analysis was used as a complementary approach to examine asymmetric allele sharing among selected Hedychium accessions. Significant D-statistic signals (|Z| > 3) were detected in multiple combinations of species accessions and cultivars, indicating that the genetic relationships among these accessions cannot be fully explained by a strictly bifurcating tree model. Representative significant tests are summarized in Table 1, and the complete results are provided in Supplementary Tables S4 and S5.
Table 1. Representative D-statistic tests showing significant asymmetric allele sharing among selected Hedychium accessions.
P1 P2 P3 D Z-score p-value H. coccineum H. gardnerianum 'Molten Gold' 0.719 20.652 < 0.001 H. forrestii H. coronarium 'Molten Gold' 0.766 12.117 < 0.001 H. coronarium H. gardnerianum 'Fireflies' 0.468 6.578 < 0.001 H. coronarium H. coccineum 'Fireflies' 0.640 10.330 < 0.001 H. chrysoleucum H. coronarium 'White Starburst' 0.356 6.933 < 0.001 H. coronarium H. gardnerianum 'Orange Brush' 0.630 12.761 < 0.001 H. coronarium H. gardnerianum 'Tara' 0.604 11.966 < 0.001 H. forrestii H. coronarium 'Tarissima' 0.552 8.848 < 0.001 Note: Positive D-values indicate excess allele sharing between P3 and P2 relative to P3 and P1 under the reported Dsuite configuration. Significant results were defined as |Z| > 3. These results indicate significant asymmetric allele sharing and should not be interpreted as direct estimates of parental contribution. In the representative cultivar tests, several cultivars showed significant asymmetric allele sharing with different species accessions or closely related lineages. For example, 'Molten Gold' showed significant signals in tests involving H. gardnerianum and H. coronarium, whereas 'Fireflies' showed significant allele sharing patterns involving both H. coccineum and comparisons involving H. gardnerianum. 'White Starburst' showed a significant signal in the comparison involving H. coronarium, consistent with its placement within the related H. coronarium group in the NJ tree and admixture analyses. These results indicate complex patterns of asymmetric allele sharing among several cultivars, species accessions, and closely related lineages. However, these D-statistic results were interpreted as evidence of asymmetric allele sharing rather than direct quantitative estimates of parental contribution.
D-statistic profile comparisons further supported the interpretation of several unknown accessions (Table 2). The detailed paired comparisons underlying these D-statistic profile metrics are provided in Supplementary Tables S6–S8. Unknown 1 showed a highly similar D-statistic profile to 'Tarissima', and Unknown 2 showed the strongest similarity to 'Elizabeth', as indicated by high profile correlations and low absolute differences in their paired D-values. Unknown 4 was also broadly similar to H. forrestii, but its lower profile correlation and larger absolute differences suggest greater differentiation than observed for Unknown 1 and Unknown 2. These results are consistent with the NJ tree, PCA, and admixture analyses, supporting close genetic affinities between Unknown 1 and 'Tarissima', between Unknown 2 and 'Elizabeth', and between Unknown 4 and H. forrestii, while indicating that Unknown 4 may represent a more differentiated accession.
Table 2. D-statistic profile comparisons between unknown accessions and their closest reference accessions.
Unknown accession Compared reference No. of paired tests D profile correlation Mean |ΔD| Median |ΔD| Unknown 1 'Tarissima' 441 0.983439 0.033351 0.027903 Unknown 2 'Elizabeth' 426 0.971047 0.027970 0.017218 Unknown 4 H. forrestii 415 0.966883 0.038125 0.028380 Note: Paired tests were defined as D-statistic tests in which the unknown accession and the compared reference accession occupied the same position in the four-taxon configuration, with the other two taxa identical and in the same order. D profile correlation represents the Pearson correlation between paired D-values. Mean |ΔD| and median |ΔD| indicate the absolute differences between paired D-values. These metrics were calculated from the complete D-statistic result set and were used as descriptive measures of D-statistic profile similarity rather than formal tests of accessions' identity. Detailed paired D-statistic comparisons for Unknown 1, Unknown 2, and Unknown 4 are provided in Supplementary Tables S6, S7, and S8, respectively. -
The genus Hedychium harbors rich genetic diversity and a long history of cultivation, yet the genetic relationships and breeding backgrounds of many modern cultivars remain poorly understood, partly because species delimitation in this genus has long been complicated by morphological similarity, interspecific variation, and hybridization[4,5,9]. This study utilized SNP markers developed via RAD-seq to conduct a population genomic analysis of Hedychium germplasm representing 35 accessions, including major cultivated types, species accessions, artificial hybrids, and accessions of uncertain identity. Integrated analyses of their phylogenetic relationships, principal components, population structure, and D-statistics revealed a complex genetic architecture characterized by clear genetic differentiation, admixed component profiles, and asymmetric allele sharing among Hedychium germplasm. This work provides useful insights into the breeding history, germplasm classification, conservation, and future utilization of Hedychium genetic resources.
Admixture patterns associated with historical hybridization in modern Hedychium cultivars
-
Multiple lines of evidence from this study indicate that modern Hedychium cultivars possess complex genetic backgrounds in which admixed genomic profiles are likely associated with historical hybridization during cultivation and breeding. The intermediate positions of artificial hybrids in the PCA and the mixed component profiles detected by admixture were consistent with their recorded hybrid backgrounds, whereas the significant asymmetric allele sharing signals detected by D-statistic analysis provided complementary evidence for genetic discordance among selected species accessions and cultivars. Together, these results suggest that historical hybridization and related genomic mixing may have contributed to the genetic structure of cultivated Hedychium. This interpretation is consistent with the known breeding history of many ornamental plants, in which breeders have widely used interspecific and intercultivar hybridization to combine desirable traits from different parents, such as flower color, form, fragrance, and adaptability[29,30].
The admixed genetic profiles observed in several cultivars and artificial hybrids provide genomic evidence consistent with this breeding background. For example, 'Baihuang Yu' and several hybrid progenies showed mixed admixture component profiles, suggesting that they carry genetic components associated with more than one associated species or cultivated background. Widespread self-incompatibility and relatively high interspecific cross-compatibility in Hedychium may further facilitate hybridization under both natural and artificial conditions[7].
Despite the admixture patterns associated with possible historical hybridization, the sampled germplasm also showed clear genetic differentiation among several major groups. For example, groups associated with H. coccineum, H. forrestii and Hedychium sp. (thick-leaved) retained relatively distinct genetic positions in the NJ tree, PCA, and admixture analyses. This pattern suggests that hybridization has not completely homogenized the genetic structure of cultivated Hedychium. Instead, modern germplasm appears to comprise both admixed cultivated materials and genetically differentiated backgrounds represented by the sampled species accessions.
Multiple species accessions associated with cultivated Hedychium germplasm
-
The integrated analyses in this study indicate that cultivated Hedychium germplasm shows genetic affinities with multiple sampled species accessions or closely related taxa. The NJ tree, PCA, and admixture consistently revealed several distinct genetic groups, suggesting that modern cultivars were shaped by genetically diverse lineages rather than by a single ancestral source, and the D-statistic analysis provided complementary evidence of asymmetric allele sharing among selected accessions. This pattern is consistent with previous reports showing that hybridization has been used to combine desirable ornamental traits, such as floral scent, flower color, and floral morphology, in Hedychium breeding[29].
Several representative species accessions showed clear associations with different cultivated groups. The group associated with H. coronarium was mainly represented in Clade 8, which included H. coronarium, closely related taxa such as H. flavescens and H. chrysoleucum, cultivars with white or pale yellow flowers, and several hybrid materials. The internal heterogeneity of this clade in the admixture analysis suggests that this cultivated group involved multiple closely related species accessions or taxa rather than a single homogeneous background. In contrast, H. coccineum was closely associated with Clade 1, which included cultivars with orange to red flowers, such as 'Tara', 'Orange Brush', and 'Fireflies', indicating its close relationship with this cultivar group. The group associated with H. gardnerianum was mainly represented in Clade 7 and several Chinese cultivated materials, especially 'Jinfen', which showed very close genetic affinity with H. gardnerianum in the PCA and admixture analyses. In addition, H. forrestii formed a genetically distinct group in Clade 3 together with 'White Stars' and Unknown 4, suggesting that it represents another differentiated lineage within the sampled germplasm.
D-statistic analysis provided complementary evidence for asymmetric allele sharing among several species accessions and cultivars, but these results were not treated as direct quantitative estimates of parental contribution, introgression proportion, or hybridization direction[26−28]. Instead, they were interpreted together with the NJ tree, PCA, and admixture results. Taken together, these analyses suggest that modern Hedychium cultivars have complex and partially overlapping genetic backgrounds, with different species accessions or closely related taxa being associated with distinct genetic backgrounds in different cultivated groups.
Genetically differentiated accessions as potential resources for future breeding
-
In addition to the genetic backgrounds widely represented in cultivated groups, this study identified several species accessions and provisionally identified accessions that retain distinct genetic characteristics and may provide useful resources for future breeding, consistent with the broader importance of conserving and evaluating diverse Hedychium germplasm[31,32]. Among them, Hedychium sp. (thick-leaved) showed the most pronounced differentiation among the sampled accessions, with its distinct position consistently supported by the NJ tree, PCA, and admixture results. No significant allele sharing signal involving this accession was detected in the tested D-statistic combinations.
H. forrestii also represented a relatively differentiated genetic background. It was assigned to Clade 3 together with Unknown 4 and 'White Stars', and this clade was separated from the major cultivar groups in the NJ tree and PCA plot. In the admixture analysis, Clade 3 showed a distinct component profile from K = 8 onward. The significant allele sharing signals involving H. forrestii were limited to a small number of cultivars, suggesting that its genetic contribution to the current cultivar set may be relatively restricted.
Other species accessions, such as H. flavescens and H. chrysoleucum, may contribute to the internal differentiation of Clade 8. Their admixture profiles were similar to those of some accessions with pale flowers, indicating that this clade contains multiple related but genetically differentiated backgrounds. These materials may therefore be useful for further diversification of white- or pale yellow-flowered germplasm.
These findings highlight the value of conserving and evaluating both genetically differentiated species accessions and provisionally identified Hedychium germplasm. Although systematic phenotypic evaluation was not conducted in this study, these materials may retain allelic variation that has not been fully captured in the current cultivated backgrounds. After further assessment of their ornamental traits, ecological adaptation, fertility, and cross-compatibility, they could provide useful variation for broadening the genetic base of cultivated Hedychium[33]. Interspecific hybridization has been shown to improve floral scent, flower color, and morphological traits in Hedychium, and related strategies have also been applied in other ornamental or crop systems[30,34−36]. Therefore, controlled interspecific hybridization, combined, where necessary, with embryo rescue, marker development, and marker-assisted selection, may provide a feasible route for introducing valuable variation(s) from genetically differentiated species accessions into future breeding programs[37,38].
Genetic grouping, ornamental traits, and future perspectives
-
The genetic grouping identified in this study showed a broad correspondence with several visible ornamental traits, particularly flower color. For example, the clade associated with H. coccineum (Clade 1) consisted predominantly of accessions with orange to red flowers, whereas the group associated with H. coronarium (Clade 8) was mainly composed of accessions with white or pale yellow flowers. This pattern suggests that the use of genetically related parental backgrounds could have contributed to the clustering of cultivars with similar floral traits. However, because systematic phenotypic measurements and trait association tests were not conducted in this study, these observations should be interpreted as descriptive associations rather than direct evidence for the genetic control of ornamental traits.
The genome-wide SNP dataset generated in this study provides a useful foundation for future analyses of trait-related genetic variation in Hedychium. With expanded sampling and standardized phenotypic evaluation, this genomic framework could be used to investigate the genetic basis of key ornamental traits, such as flower color, floral scent, flowering time, and inflorescence architecture. Recent metabolomic, transcriptomic, and sensory omics studies in Hedychium have provided valuable insights into floral scent formation, volatile compound variation, and fragrance type differentiation, suggesting that population genomic resources could be integrated with multi-omics approaches in future trait studies[39−41]. Future studies integrating population genomics with genome-wide association analysis, transcriptomics, metabolomics, and volatile compound profiling would be particularly valuable for identifying candidate loci and biological pathways related to ornamental diversification[42−45]. Such integrative approaches may facilitate the transition of Hedychium breeding from phenotype-based selection toward more precise molecular breeding.
This study provides a population genomic framework for understanding the genetic differentiation, admixture patterns, and genetic affinities of Hedychium germplasm. The results indicate that modern cultivated materials are associated with multiple genetic backgrounds represented by the sampled species accessions or closely related taxa. Recorded artificial hybrids displayed mixed genetic components consistent with their documented origins, whereas several cultivars showed admixture patterns potentially associated with historical hybridization. Several genetically differentiated accessions, including the provisionally identified Hedychium sp. (thick-leaved) and H. forrestii, may represent useful resources for broadening the genetic base of future breeding after further phenotypic and reproductive evaluation. These findings provide useful information for germplasm classification, conservation, and breeding applications in Hedychium.
Certain limitations of this study should be noted. First, RAD-seq is a reduced representation sequencing method based on restriction enzyme digestion, which may not capture variants in genomic regions lacking the targeted restriction sites and may be affected by missing data or sampling bias in some genomic regions[15,46]. Future studies could utilize whole-genome resequencing to obtain more comprehensive genomic information[47−49]. Second, the NJ tree was rooted using H. villosum as a functional outgroup within the genus rather than a true external outgroup from a related Zingiberaceae genus; incorporating such data in future studies would further strengthen the phylogenetic framework. Third, the present study focused primarily on genomic relationships and did not include systematic phenotypic, cytological, or reproductive compatibility assessments. Integrating these data in future research would allow a more direct evaluation of the relationships among genetic background, ornamental traits, ploidy variation, and breeding potential[50,51]. Additionally, our samples were primarily from China and the USA, with limited representation of Hedychium species diversity from the Himalayan region and other parts of tropical and subtropical Asia. Expanding the collection to include a wider geographical range and more diverse sample types would offer a more complete depiction of the genetic landscape of Hedychium.
-
This study characterized the population genetic structure of Hedychium germplasm using RAD-seq. After Sample 30-6 was removed, the displayed NJ topology contained 167 samples. The admixture and D-statistic analyses were rerun using 167-sample datasets, and the final PCA included 165 samples after the exclusion of two extreme outliers. The NJ tree resolved eight major clades, which were broadly supported by the PCA and admixture results. D-statistic analysis detected asymmetric allele sharing in selected comparisons involving species accessions and cultivars, indicating complex and partially overlapping genetic backgrounds.
Modern cultivated materials showed genetic affinities with multiple backgrounds represented by the sampled species accessions, particularly those related to H. coronarium, H. coccineum, H. gardnerianum, and H. forrestii. Recorded artificial hybrids generally exhibited mixed genetic components consistent with their recorded origins, whereas several cultivars showed admixture patterns potentially associated with historical hybridization. Additionally, several genetically differentiated accessions, such as Hedychium sp. (thick-leaved) and H. forrestii, may provide useful genetic variation for broadening the genetic base of future breeding after further phenotypic and reproductive evaluation. Overall, this research establishes a population genomic framework for germplasm classification, conservation, and molecular breeding in Hedychium.
-
The authors confirm contribution to the paper as follows: study conception and design: Cai B, Bao P; writing – original draft, reviewing, and editing: Luo X, Cai B; investigation, data analysis, and supervision: Luo X, Bao P, Cai C, Wang M, Xie Y, Wang J; funding acquisition: Cai B. All authors reviewed the results and approved the final version of the manuscript.
-
The raw RAD-seq sequencing reads have been deposited in the NCBI SRA under BioProject accession PRJNA1484304, and the Data Availability section has been updated accordingly. As the data have not yet been publicly released, the deposited submission information can be viewed using the following NCBI reviewer link: https://dataview.ncbi.nlm.nih.gov/object/PRJNA1484304?reviewer=1jpnh7p4kkb2dg06puul4gsbu.
-
The authors declare that they have no conflict of interest.
-
This accompanies this paper online at: https://doi.org/10.48130/opr-0026-0026.
- Supplementary Table S1 Information of 35 Hedychium germplasm accessions.
- Supplementary Table S2 Summary statistics of RAD-seq reads for 168 Hedychium samples.
- Supplementary Table S3 SNP information statistics of 168 Hedychium samples.
- Supplementary Table S4 Significant D statistic results with |Z| > 3.
- Supplementary Table S5 Significant D statistic tests with P1 and P2 as wild species and P3 as cultivar.
- Supplementary Table S6 D statistic profile comparison between Unknown 1 and 'Tarissima'.
- Supplementary Table S7 D statistic profile comparison between Unknown 2 and 'Elizabeth'.
- Supplementary Table S8 D statistic profile comparison between Unknown 4 and H. forrestii.
- Supplementary Fig. S1 Preliminary PCA of all 167 Hedychium samples based on 265,365 SNPs after LD pruning.
- Supplementary Fig. S2 Representative floral morphology of selected species accessions and cultivars of Hedychium.
- Copyright: © 2026 by the author(s). Published by Maximum Academic Press, Fayetteville, GA. This article is an open access article distributed under Creative Commons Attribution License (CC BY 4.0), visit https://creativecommons.org/licenses/by/4.0/.
-
About this article
Cite this article
Luo X, Bao P, Cai C, Wang M, Xie Y, et al. 2026. Unveiling genetic differentiation and admixture in modern Hedychium germplasm: a population genomics perspective from RAD-seq. Ornamental Plant Research 6: e035 doi: 10.48130/opr-0026-0026 
Unveiling genetic differentiation and admixture in modern Hedychium germplasm: a population genomics perspective from RAD-seq
- Received: 14 February 2026
- Revised: 25 June 2026
- Accepted: 03 July 2026
- Published online: 09 October 2026
Abstract: The genus Hedychium has high ornamental and economic value, but its morphological classification remains contentious and the genetic backgrounds of many cultivars are unclear. In this study, we initially sequenced 168 Hedychium samples representing various species accessions, cultivars, artificial hybrids, and accessions of uncertain identity from China, the USA, and Thailand, using restriction site-associated DNA sequencing (RAD-seq). One low-quality sample was excluded from the principal component analysis (PCA), admixture, and D-statistic analyses, and its terminal branch was pruned from the displayed neighbor-joining (NJ) topology. In total, 856.92 Gb of high-quality data were generated, and 647,693 high-quality single-nucleotide polymorphisms (SNPs) were used for constructing the NJ tree, 265,365 linkage disequilibrium (LD)-pruned SNPs were used for PCA and admixture analyses, and 100,000 randomly sampled genome-wide SNPs were used for D-statistic analysis. The NJ tree resolved eight major clades, which were broadly supported by PCA and admixture analyses. D-statistic analysis further detected significant asymmetric allele sharing among selected accessions. Cultivated materials were associated with multiple genetic backgrounds represented by the sampled species accessions, including groups related to H. coronarium, H. coccineum, H. gardnerianum, and H. forrestii. Artificial hybrid accessions generally displayed mixed genetic components consistent with their recorded hybrid backgrounds. Several accessions of uncertain identity were assigned to closely related genetic groups, providing clues for their further identification. Our study provides a genome-scale population genetic framework for understanding genetic differentiation, admixture patterns, and genetic affinities within major Hedychium germplasm, offering molecular insights for the classification, conservation, and future breeding.
-
Key words:
- Hedychium /
- RAD-seq /
- SNP /
- Genetic diversity and structure