










-
Aquifers are an important component of terrestrial ecosystems and hold the largest accessible freshwater resource on Earth[1]. Groundwater contamination, exacerbated by economic development, geogenic processes, and climate change, has become a global issue that threatens drinking water safety, human health, and ecosystem stability[2−5]. Compared with surface water systems, the transport and transformation of groundwater contaminants are more difficult to predict due to the pronounced structural and chemical heterogeneities of aquifer media[6−8]. Researchers have devoted significant efforts to quantifying aquifer heterogeneity to better understand contaminant fate and to design effective, low-cost remediation strategies[6,9−11]. Structural heterogeneity plays a fundamental role in controlling the transport of groundwater contaminants. It is commonly characterized by hydraulic conductivity (k), which is widely used as a key parameter in contaminant transport modeling[12,13]. Hydraulic conductivity varies over several orders of magnitude, ranging from 1 × 10⁻¹¹ – 4.7 × 10⁻⁹ m/s for clay sediments to 9 × 10⁻⁷ – 6 × 10⁻³ m/s for coarse sands[14]. This structural heterogeneity mainly controls the advection and dispersion of groundwater contaminants[15], resulting in the retention of contaminants in low-permeability zones[16,17]. In addition to structural heterogeneity, contaminant transport and transformation are strongly influenced by the chemical heterogeneity of aquifer sediments, arising from diverse redox-active species such as Fe/Mn minerals, natural organic matter (NOM), and sulfur-containing compounds[18−20]. For example, electron-donating species in sediments (e.g., Fe(II) and reduced organic matter) can readily transform reducible contaminants such as Cr(VI) and nitroaromatic compounds[6,21−24]. Consequently, contaminant degradation rates may vary over several orders of magnitude depending on the reactivity of these redox-active components[25−27]. Similarly, the remediation of contaminated aquifers by in situ chemical oxidation (ISCO) is also strongly influenced by chemical heterogeneity, which constrains both the consumption and activation of injected oxidants[28,29].
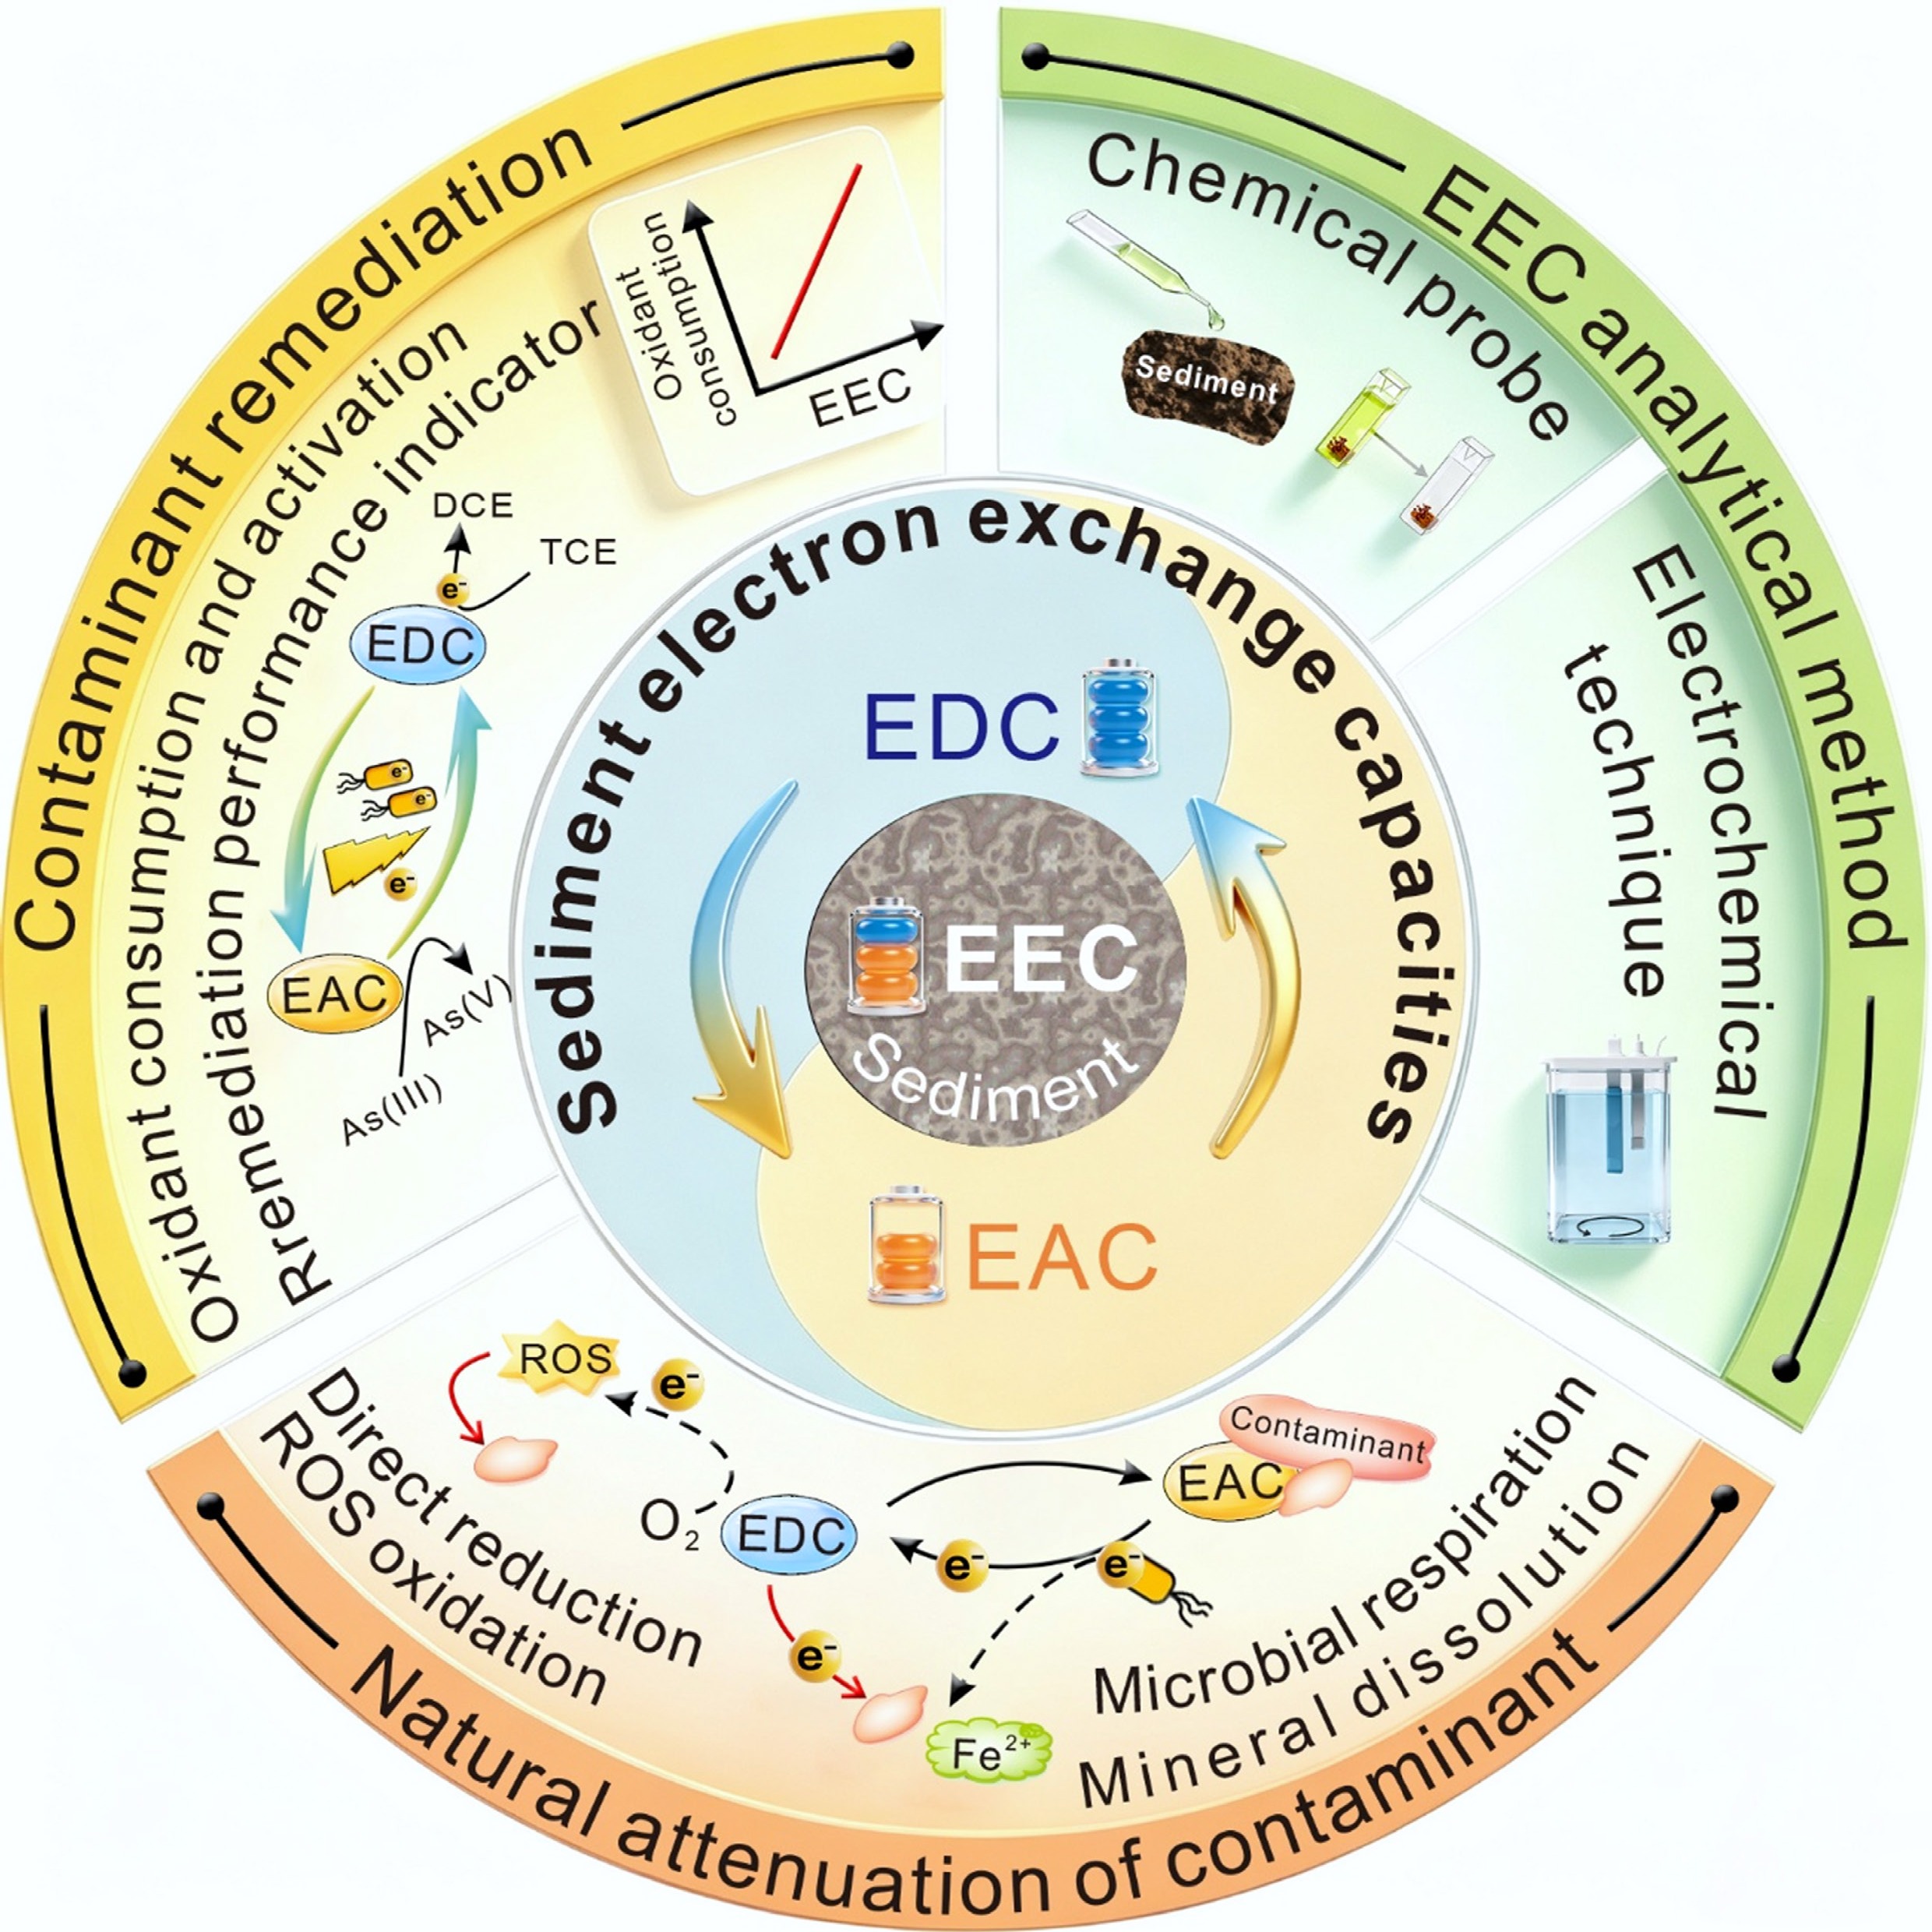
In contrast to well-characterized structural heterogeneity, the chemical heterogeneity of aquifer sediments remains far less well-quantified. This knowledge gap largely stems from the lack of appropriate methodologies[30]. Previous studies have typically measured the bulk contents of presumed redox-active components (e.g., Fe, organic matter, and S) to characterize sediment chemical heterogeneity[7,31]. However, such approaches cannot identify the specific species that actively donate or accept electrons[19]. In recent years, the concept of electron exchange capacity (EEC) has been introduced to better represent the chemical heterogeneity of aquifer sediments[19,32,33]. Sediment EEC comprises electron-donating capacity (EDC) and electron-accepting capacity (EAC), which quantify the amounts of electrons that can be transferred from and to sediments, respectively. Quantifying EEC provides valuable insights into sediment redox reactivity, including the rate, extent, and direction of electron transfer. Consequently, quantitative relationships for redox reactions can be established based on sediment EEC[34,35], which is often not achievable when using bulk component contents due to sediment chemical heterogeneity[36]. The development of EEC measurement methods[32,34] has opened new opportunities to understand biogeochemical processes and contaminant transformation in aquifers. The EEC has been incorporated into reactive transport models as a novel yet important parameter to better describe the complex reaction networks of biogeochemical processes[37,38]. In recent review papers, researchers have linked sediment EEC to biogeochemical–hydrological processes[18] and environmental impacts such as nitrogen cycling[33]. However, the role of sediment EEC in controlling the natural attenuation of groundwater contaminants and guiding sustainable remediation strategies remains unclear. In this review, we propose that sediment EEC can effectively represent the chemical heterogeneity of aquifers that governs contaminant transformation and remediation, compared with traditionally used parameters such as bulk elemental contents. To this end, we systematically review: (1) the development of methodologies for EEC measurement; (2) the composition of EEC in contaminated aquifers; (3) contaminant transformation processes driven by sediment EEC; and (4) the implications of sediment EEC for groundwater remediation. Finally, we discuss current knowledge gaps and future research directions, aiming to provide mechanistic insights into EEC-driven natural attenuation of groundwater contaminants and practical guidance for contamination remediation.
-
The EEC of aquifer sediments originates from redox-active species capable of donating or accepting electrons. Based on the elemental composition and abundance of soils and aquifer sediments, EEC is generally attributed to Fe/Mn-bearing minerals, natural organic matter (NOM), and sulfur-containing compounds[19,39−42]. Among these, Fe-bearing minerals and NOM are considered the most important contributors because of their high redox reactivity and widespread occurrence[19,35,39]. Therefore, suitable EEC measurement methods should efficiently oxidize or reduce these redox-active components[35]. However, not all presumed redox-active species (e.g., Fe) are sufficiently reactive to contribute to EEC. For example, Fe structurally constrained within non-expandable phyllosilicates is generally redox-inert and contributes little to sediment EEC[19,41]. These redox-inert Fe fractions also participate minimally in biogeochemical processes[19]. For heterogeneous NOM, humic acid fractions, particularly lignin-like compounds, are the dominant contributors to NOM-derived EEC[19]. Although the total EEC of aquifer systems includes contributions from both solid and aqueous phases, more than 99% of the total EEC is contributed by the solid phase[31]. Dissolved species in groundwater generally have negligible influence on sediment EEC measurements. Nevertheless, measuring the EEC of both sediments and the corresponding groundwater is encouraged to better characterize the distribution of EEC in subsurface environments.
EEC analytical methods
-
Methods for measuring EEC generally fall into two categories (Fig. 1). The first is chemical probe methods, in which sediments react with selected chemical probes, and the EEC is calculated from the consumption of these probes. The second is mediated electrochemical analysis, where sediments interact with electrodes through electron-transfer mediators, and the EEC is determined based on the resulting current–time curve. Several of these methods have been summarized in previous reviews[32,43]. In recent years, new techniques within these two categories have emerged[34], offering improvements in analytical accuracy, shorter analysis time, and lower detection limits.
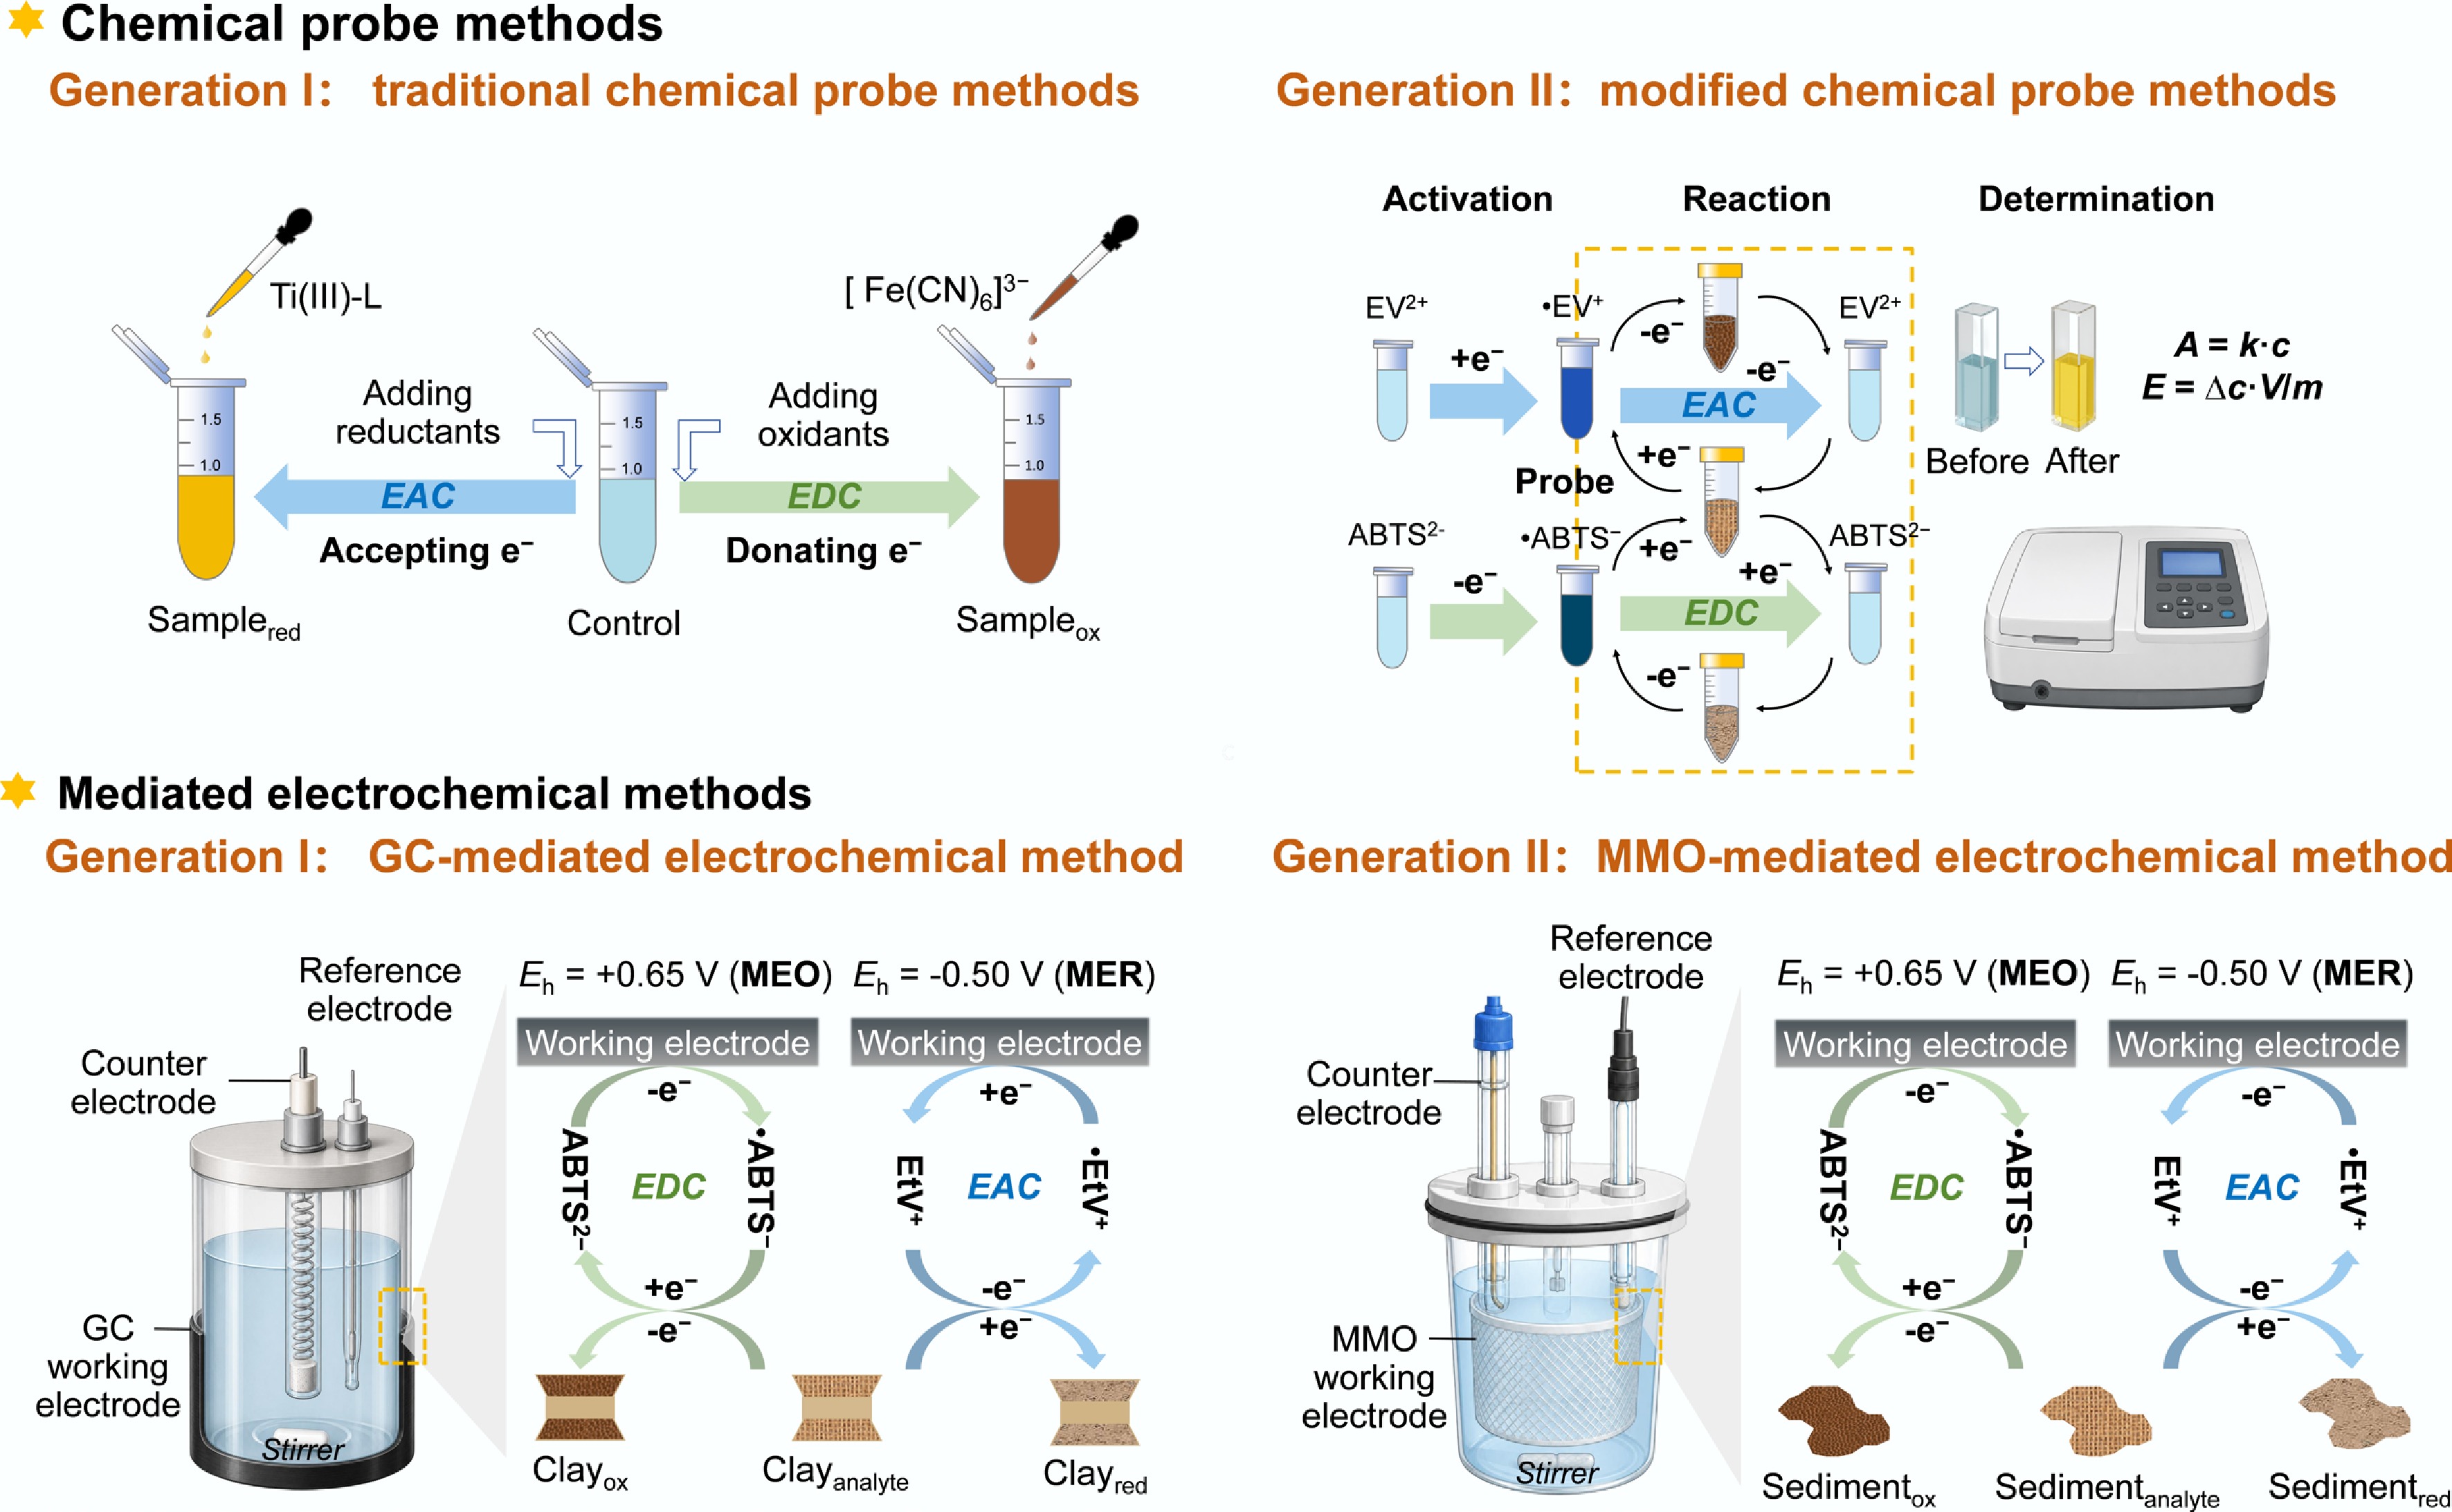
Figure 1.
EEC analysis by chemical probe methods and mediated electrochemical methods.
Chemical probe methods
Generation I: traditional chemical probe methods
-
Traditional chemical probe methods were among the earliest techniques used to characterize the redox capacity of sediments and NOM[44,45]. The basic principle involves reacting sediments with known amounts of an oxidant or reductant. The number of transferable electrons is then inferred from changes in the concentrations of the residual reagent or the reaction products[31,46]. For EDC measurement by chemical oxidation, commonly used oxidants include K3[Fe(CN)6], Fe(III)-citrate, and Cr(VI)[45−47]. The decrease in EDC after biogeochemical reduction has also been used to estimate the EAC of samples[48]. For EAC measurement by chemical reduction, a commonly used chemical probe is ligand-complexed titanous ion (Ti(III)-L)[44]. These methods are widely applied to determine the EEC of NOM[48,49]. One of the most important findings from these studies is that NOM can serve as an electron acceptor and function as an electron shuttle to promote microbial reduction of Fe(III)[48,50].
Despite their widespread use in early studies, traditional chemical probe methods have inherent limitations. First, many chemical redox reactions proceed slowly. Electron transfer is often constrained by mineral structure and surface binding sites, leading to slow reaction kinetics that may require tens of hours or longer to approach equilibrium[51,52]. Second, the redox potential of chemical reagents is highly sensitive to pH and complexation conditions[53,54]. The redox behavior of the same reagent may shift under different measurement environments. As a result, EEC measurements are strongly condition-dependent and difficult to compare across studies. Third, different chemical reagents exhibit varying selectivity toward functional groups and often capture only a portion of the EDC or EAC[47,55]. This suggests that the 'measured capacity' more accurately reflects the number of sites that react with a specific reagent, rather than the intrinsic, total redox capacity of the sample. For example, the EAC of humic acids measured by traditional chemical probe methods was approximately three times lower than that measured by mediated electrochemical methods[30].
Generation II: modified chemical probe methods
-
In contrast to traditional chemical probe methods, recent studies have developed modified approaches using radical-based chemical probes[34]. Ethyl viologen radical (•EV+, Eh0 ≈ −0.45 V vs SHE) and 2,2′-azino-bis(3-ethylbenzothiazoline-6-sulfonic acid) radical (•ABTS−, Eh0 ≈ +0.70 V vs SHE) are used as reductant and oxidant, respectively, to determine sediment EAC and EDC. •EV+ is prepared via Pd–H2 reduction of EV2+, while •ABTS− is generated through persulfate oxidation of ABTS2−[34]. This modified chemical probe method provides a simple and accurate approach for assessing the redox capacity of sediments, Fe/Mn minerals, and NOM. The detection limits of EAC and EDC are 0.6 and 0.4 μmol e−/L, respectively, which are three to four orders of magnitude lower than those of traditional chemical probes[34]. The reaction is also time-efficient, requiring only 15 min. In addition, relatively large sample dosages can be used, making the method well-suited for heterogeneous sediments and soils. Using this approach, the vertical distribution of sediment EAC and EDC was quantified in a riparian aquifer[34], reflecting redox condition changes across the groundwater table. A limitation of this method is that EAC and EDC can only be measured at the specified redox potentials (i.e., −0.45 and +0.70 V vs SHE). The fractions of EEC at other redox potentials, which correspond to different reactivities, cannot be determined due to the lack of suitable radical-based probes.
Mediated electrochemical methods
Generation I: GC-mediated electrochemical method
-
Electrochemical methods can measure the EEC of samples at a glassy carbon (GC) working electrode under defined thermodynamic boundary conditions (e.g., constant pH and Eh). The amount of electron transfer between the samples and the GC working electrode can be easily quantified using chronocoulometry. The GC electrode is chosen for its low background current and good chemical stability. However, direct electrochemical analysis is limited by slow electron transfer between the sample and the electrode[30]. To address this limitation, Sander's research group developed a novel approach, namely mediated electrochemical reduction (MER) and mediated electrochemical oxidation (MEO)[30,32]. MER and MEO enable fast (requiring only hours) and accurate measurements of EAC and EDC, respectively, using appropriate electron transfer mediators[32]. These mediators undergo rapid and reversible electron transfer in solution, acting as electron shuttles between the sample and the working electrode[32]. The most commonly used mediators are 2,2′-azino-bis(3-ethylbenzothiazoline-6-sulfonic acid) (ABTS2−/ABTS•−, +0.7 V) and ethylviologen (EV2+/EV•+, −0.45 V), which are employed for EDC and EAC measurements, respectively[56,57]. Other mediators with lower oxidizing or reducing potentials can also be applied to determine EEC, thereby capturing its reactivity under different applied potentials. The selection of mediators and corresponding potentials has been comprehensively summarized in previous studies[32,56]. Further studies have implemented a flow-injection analysis platform for mediated electrochemical measurements, allowing low-concentration injections and automated operation[58]. Mediated electrochemical methods have been widely applied to measure the EEC of Fe-bearing clay minerals[41,59], Fe (oxyhydr)oxides[60], NOM[30,61−63], soils[35,64], and sediments[19,39,65]. Method validation was supported by the comparable EEC values and Fe contents in standard Fe-bearing minerals[59,60]. Strictly speaking, mediated electrochemical methods are best suited for relatively homogeneous samples, including dissolved species, standard minerals, and fine fractions of soils and sediments, due to the small sample injection volumes required.
Generation II: MMO-mediated electrochemical method
-
In recent years, a key focus in the development of mediated electrochemical analysis has been the improvement of working electrodes and reaction systems. These advances aim to enhance the method's applicability to heterogeneous samples, such as whole soils and sediments. Conventional mediated electrochemical systems use GC electrodes as both the working electrode and the reaction cell, allowing only milligram-level or lower sample injections. However, such small sample amounts are often insufficient to represent the average redox properties of bulk soils or sediments. To address this limitation, our coworkers introduced mixed metal oxide (MMO) electrodes[66], providing a new approach for applying mediated electrochemical analysis to heterogeneous sediments. Compared with conventional GC electrodes, MMO electrodes exhibit higher electrochemical reactivity and a larger effective interfacial area, making them more suitable for analyzing the EEC of heterogeneous sediments (sample injection: 0.2–6 g)[66]. Similar to GC-mediated electrochemical methods, MMO-mediated methods can quantify EEC over a range of Eh values (−0.45 to +0.7 V)[66]. Therefore, MMO-mediated electrochemical analysis provides more accurate and reliable EEC measurements for heterogeneous soils and sediments, offering improved data for evaluating biogeochemical reactions in aquifers.
-
The transport and transformation of groundwater contaminants are closely linked to redox reactions. These electron transfer processes are governed by sediment EEC, which can influence microbial activity, chemical speciation, mobility of toxic elements, and both direct and indirect contaminant transformation. The specific roles of sediment EEC depend on its composition, the prevailing redox conditions, and the properties of the contaminants.
Sediment EAC as a terminal electron acceptor of anaerobic microbial respiration
-
Many microorganisms can survive in contaminated environments and play important roles in the biotransformation of toxic metals[67] and organic contaminants[68]. Under anoxic conditions, sediment EAC serves as a terminal electron acceptor for anaerobic microbial respiration, with organic matter acting as the carbon source. EAC derived from both Fe(III) and solid organic matter (SOM) can be bioavailable, depending on the physical accessibility and redox potential of the contributing species[19]. It has been reported that EACFe(III) and EACSOM in sediments can be microbially reduced to generate sediment EDC[19,35,39,65], with ΔEAC values ranging from 0.001–0.088 mmol e−/g for aquifer sediments[19] and ~0.02–0.6 mmol e−/g for lake sediments[39]. The relative contributions of Fe(III) and SOM to the overall decrease in EAC can be roughly estimated based on Fe(III)/Fe(II) variations[19,35]. In organic-rich environments such as peatlands, the EAC of peat particulate organic matter can serve as the terminal electron acceptor[69−71], with values reaching up to 27 × 106 mol e−/(km2·yr)[69]. Thus, sediment EAC can stimulate anaerobic microbial respiration, as reflected in the correlation between CO2 production and initial soil or sediment EAC[35]. In turn, microbial reduction of sediment EAC creates a more reduced environment[72,73], which favors the reductive transformation of contaminants (Fig. 2).
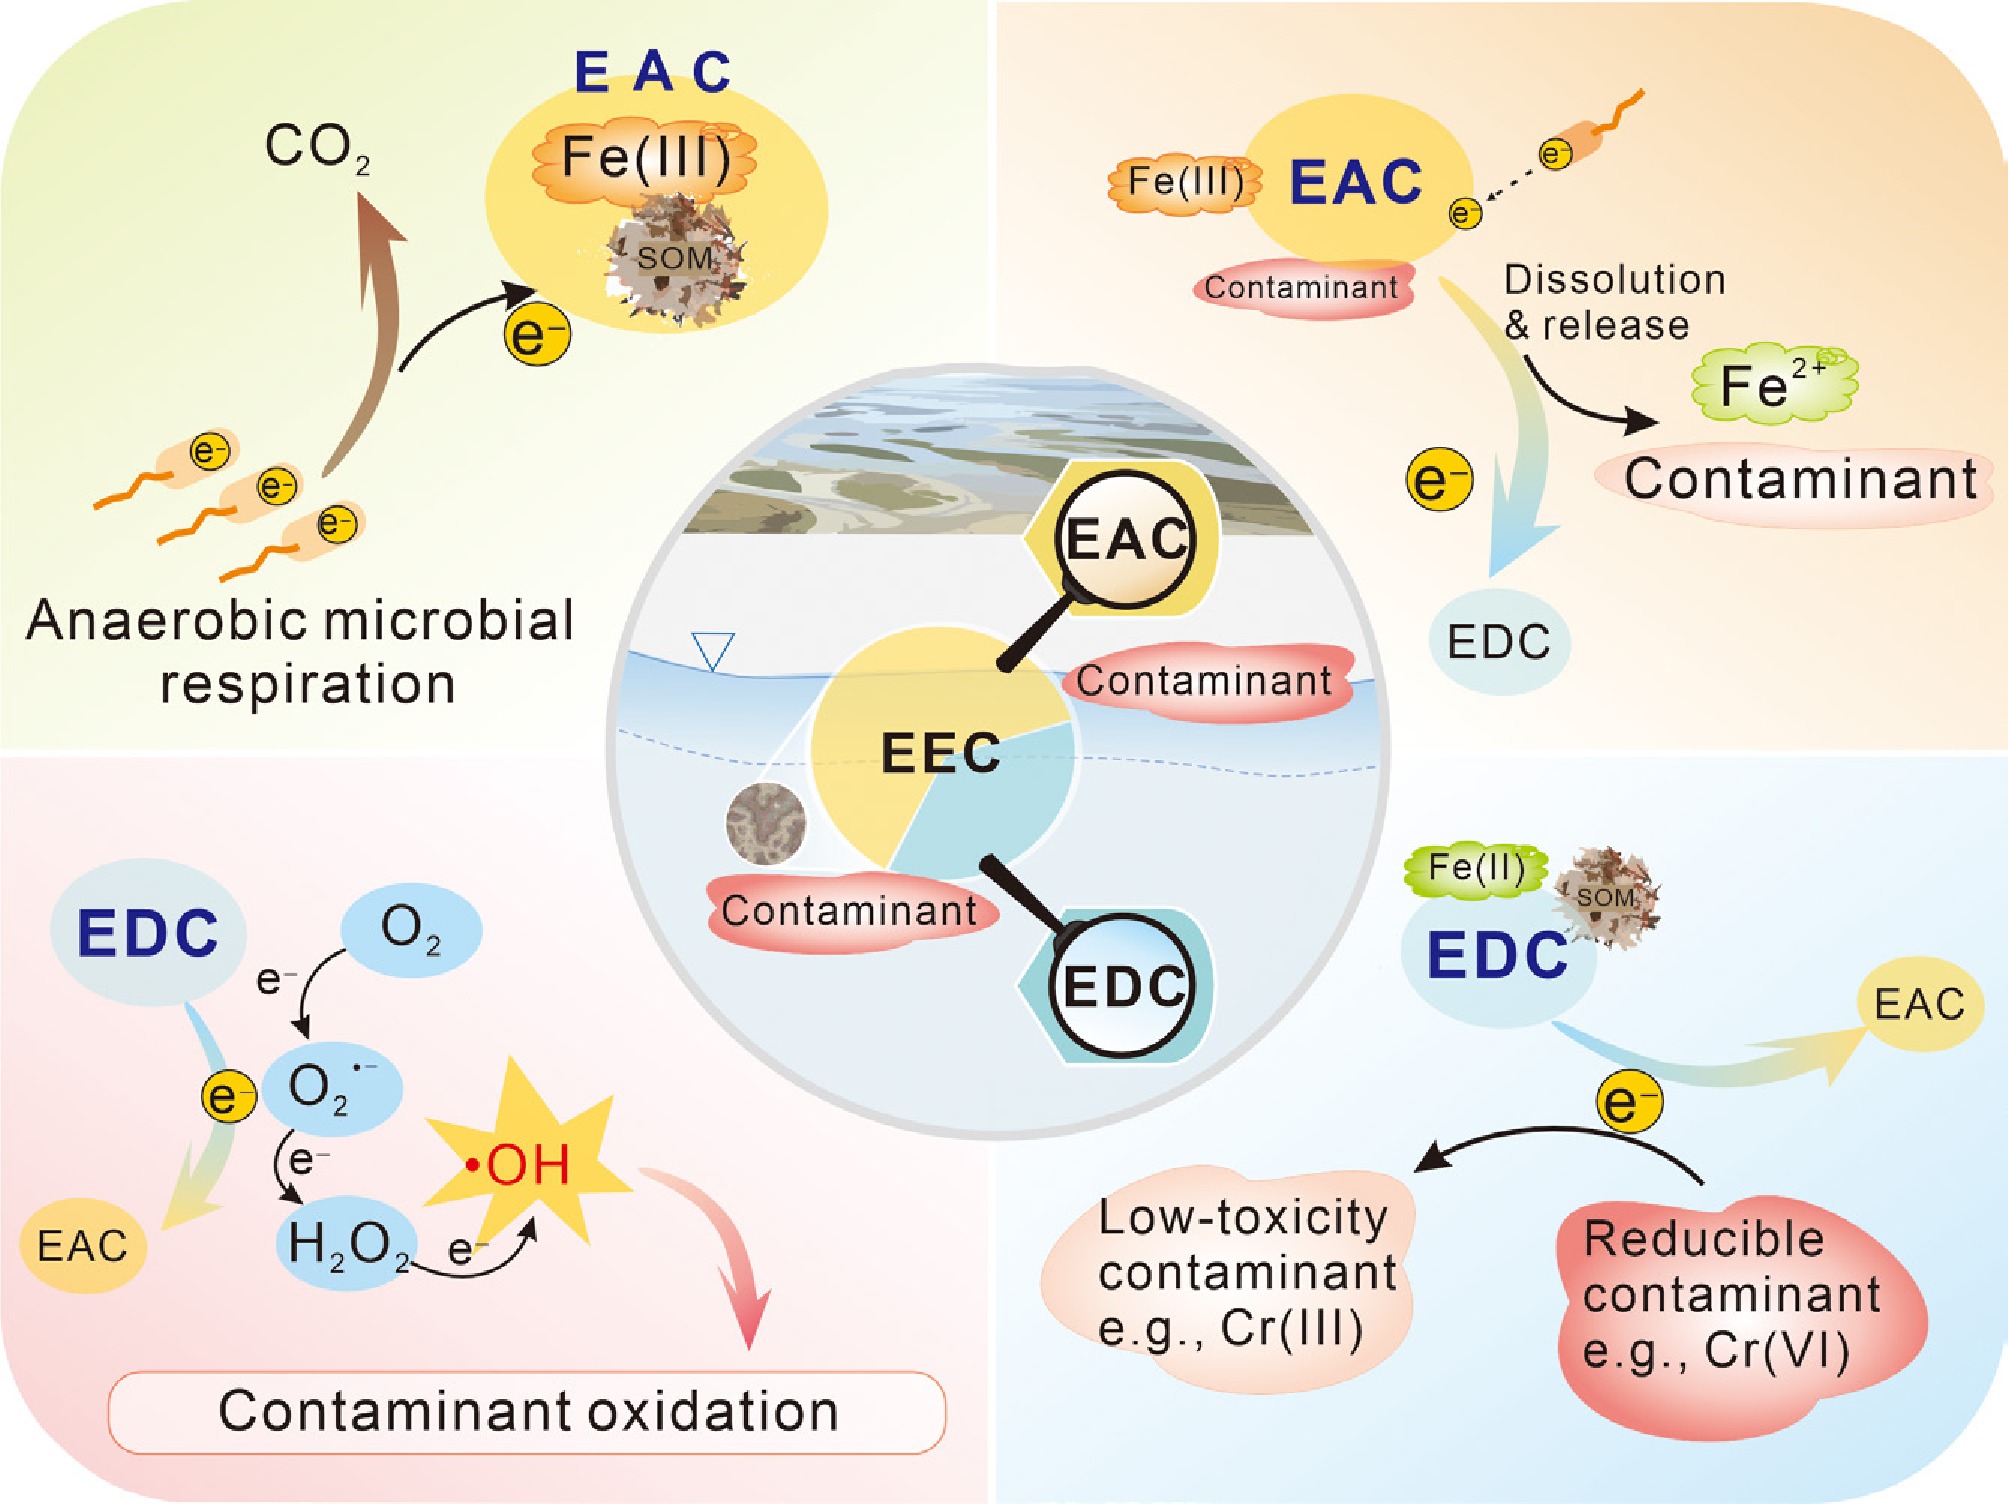
Figure 2.
Contaminant transport and transformation driven by EEC reactions in aquifers.
EAC reduction-driven mineral dissolution and the release of associated contaminants
-
Abiotic and microbial reduction of sediment EAC associated with Fe(III) minerals may induce mineral dissolution, depending on the mineral type. For expandable phyllosilicates (e.g., smectites) in sediments, the entire EAC can be accessed at an EAC/Fe(III) ratio of 1:1[59]. Reduction of smectite EAC generally causes limited mineral dissolution, although dissolution can reach up to 32% in highly reduced smectite[74,75]. In contrast, non-expandable phyllosilicates (e.g., chlorite and biotite) exhibit low EAC reactivity[19], low microbial reducibility[76], and minimal dissolution[77]. Compared with phyllosilicates, Fe-(oxyhydr)oxides are more susceptible to reductive dissolution[78]. The extent of EAC reduction-induced mineral dissolution decreases with increasing thermodynamic stability[60]. Low-crystallinity minerals such as ferrihydrite can be almost completely reductively dissolved, whereas highly crystalline minerals such as goethite and hematite are only partially dissolved[60,79,80]. Therefore, reduction of sediment EAC can lead to substantial mineral dissolution, as indicated by the production of dissolved Fe2+[81]. This process can release mineral-associated contaminants into surrounding aquatic environments. For example, reduction of lake sediment EAC can mobilize mineral-bound phosphorus, increasing phytoplankton productivity by up to 40%[65]. By extension, reduction of sediment EAC can also release toxic metals associated with minerals, such as arsenic (As)[82,83]. Because microbial reduction of sediment EAC is often mediated by dissolved electron shuttles such as dissolved organic matter (DOM), groundwater As concentrations have been observed to correlate with the EEC of DOM[84]. DOM with higher EEC promotes more extensive reduction of As-bearing Fe(III) oxides, resulting in greater As release to groundwater[84]. Therefore, sediment EAC and dissolved EEC can serve as useful predictors of mineral-associated contaminant release in aquifers (Fig. 2).
Direct reaction of sediment EDC with reducible contaminants
-
Sediment EDC represents electron sources that can be transferred to reducible contaminants under favorable thermodynamic conditions. In principle, previously reported contaminant reduction reactions mediated by sediments and their components[6,25,85] can be quantitatively described by EDC-based reactions. However, such quantitative studies remain scarce. For example, Lu et al. established a linear correlation between sediment EDC depletion and Cr(VI) reduction[34]. A slope of 0.3 accurately reflects the three-electron transfer involved in the reduction of Cr(VI) to Cr(III) (Eq. [1])[34]. Similar correlations have been reported between sediment EDC and the rate constants of nitrobenzene reduction[86], indicating direct electron transfer from sediment EDC to contaminants. In heterogeneous aquifers, low-permeability lenses enriched in EDC can release dissolved EDC into adjacent transmissive zones, where it can directly transform reducible contaminants in groundwater, such as Cr(VI)[38,87]. It has also been reported that dissolved EDC derived from humic acids exhibits a linear correlation with the reduction of the munition compound 3-nitro-1,2,4-triazol-5-one (NTO)[88]. The slope suggests that approximately half of the electrons stored in humic acids are accessible to NTO[88]. These findings indicate that sediment EDC, as well as its dissolved fraction, imposes a critical stoichiometric constraint on the transformation of redox-active contaminants (Fig. 2).
$ {\mathrm{Cr}}({\mathrm{VI}})+3{\mathrm{EDC}}\rightarrow {\mathrm{Cr}}({\mathrm{III}})+3{\mathrm{EAC}} $ (1) Reactive oxygen species generation from sediment EDC oxidation for contaminant transformation
-
In recent years, researchers have found that reactive oxygen species (ROS), including superoxide radical (O2•−), hydrogen peroxide (H2O2), and hydroxyl radical (•OH), can be generated through O2 activation at aquifer redox interfaces[89,90]. Field studies have increasingly documented the generation and distribution of ROS in shallow aquifers[91−94], paddy soils[95,96], and coastal soils[72], revealing a previously overlooked oxidizing force in subsurface redox-fluctuating environments. Among these ROS, •OH (standard reduction potential: 2.8 V[97]) is the most reactive species and plays a key role in the degradation of refractory organic contaminants, including phenol[98], polycyclic aromatic hydrocarbon[99], petroleum hydrocarbon[100], and antibiotics[57] in contaminated aquifers (Fig. 2). The formation of •OH involves three-electron transfer from reduced species to O2[89]. Fe(II)-bearing minerals and reduced organic matter have been recognized as the dominant electron donors in sediments[90,101]. Electron transfer from various Fe(II) species associated with sediments to O2 can be quantified by measuring changes in Fe(II) content[98,102,103]. In contrast, quantifying the contribution of SOM to sediment oxygenation and •OH production remains challenging[104]. By quantifying the EDC of SOM, electron transfer pathways from interior to surface sites and subsequently to O2 have been identified during oxygenation[104]. With these pathways and rates, the contributions of SOM and Fe(II) to total •OH production were evaluated, revealing a threshold at a molar ratio of SOC/Fe(II) of 39[104]. In addition, interfacial electron transfer results in dissolved EDC of DOM[105]. This EAC–EDC conversion of DOM contributes to long-distance electron transfer (e.g., at the decimeter scale) along redox gradients[20,106] and drives the remote production of •OH in subsurface environments[107].
The generated •OH can induce oxidative transformation of both organic contaminants and organic matter in subsurface environments. A correlation between sediment EDC and •OH production has been established[34], providing a quantitative basis for evaluating the potential for ROS generation and contaminant oxidation by aquifer sediments. Sediment EDC measurements can also help distinguish chemical and biotic contributions to contaminant degradation. Sediment oxygenation results in a rapid decrease in EDC and •OH-induced contaminant degradation[57], whereas biotic oxidation leads to minor EDC variation and slower contaminant transformation[108]. In this context, a recent study showed that •OH oxidation contributed less than 22.6%, while biotic oxidation accounted for 77.4% of total phenol degradation in sediments[108]. Similar to the oxidation of organic contaminants, •OH oxidation of organic matter represents a dark pathway in the soil/sediment carbon cycle[109−113]. It has been reported that soil EDC variation correlates well with CO2 production, with 5%–20% of dissolved organic matter (DOC) oxidized by •OH during soil oxygenation[114].
Overall, the indirect transformation of contaminants and organic matter mediated by •OH can be effectively predicted based on EDC reactions. This suggests that EDC or EAC quantification has unique value in elucidating electron transfer pathways, rates, and extents in complex biogeochemical processes. These processes typically involve multiple electron donors and acceptors, chain reactions with reactive intermediates, and complex, often unpredictable transformation products that are difficult to quantify with traditional methods.
-
Quantifying the heterogeneity of contaminated sites is a prerequisite for successful remediation. A statistical survey of 33 organic-contaminated sites in China revealed heterogeneous soil layers in all sites, with low-permeability media such as clay present in up to 83% of cases[115]. Regarding the chemical heterogeneity of contaminated sites, the EEC values of contaminated soils/sediments have been rarely reported[19,34]. Here, we collected chemical composition data for sediments from 21 contaminated aquifers in China with varying geological types and contamination characteristics (Supplementary Table S1). The results show that sediments in these aquifers widely contain redox-active components such as Fe, Mn, S, and solid organic carbon (SOC) (Fig. 3a). Specifically, Fe ranges from 2.8 to 99.49 g/kg, SOC from 0.465 to 51.95 g C/kg, Mn from 0.057 to 1.06 g/kg, and S from 0.106 to 7.28 g/kg (Supplementary Table S1). These redox-active substances are the main contributors to sediment EEC[19]. Thus, these contaminated aquifers are expected to exhibit wide EEC ranges, reflecting high chemical heterogeneity.
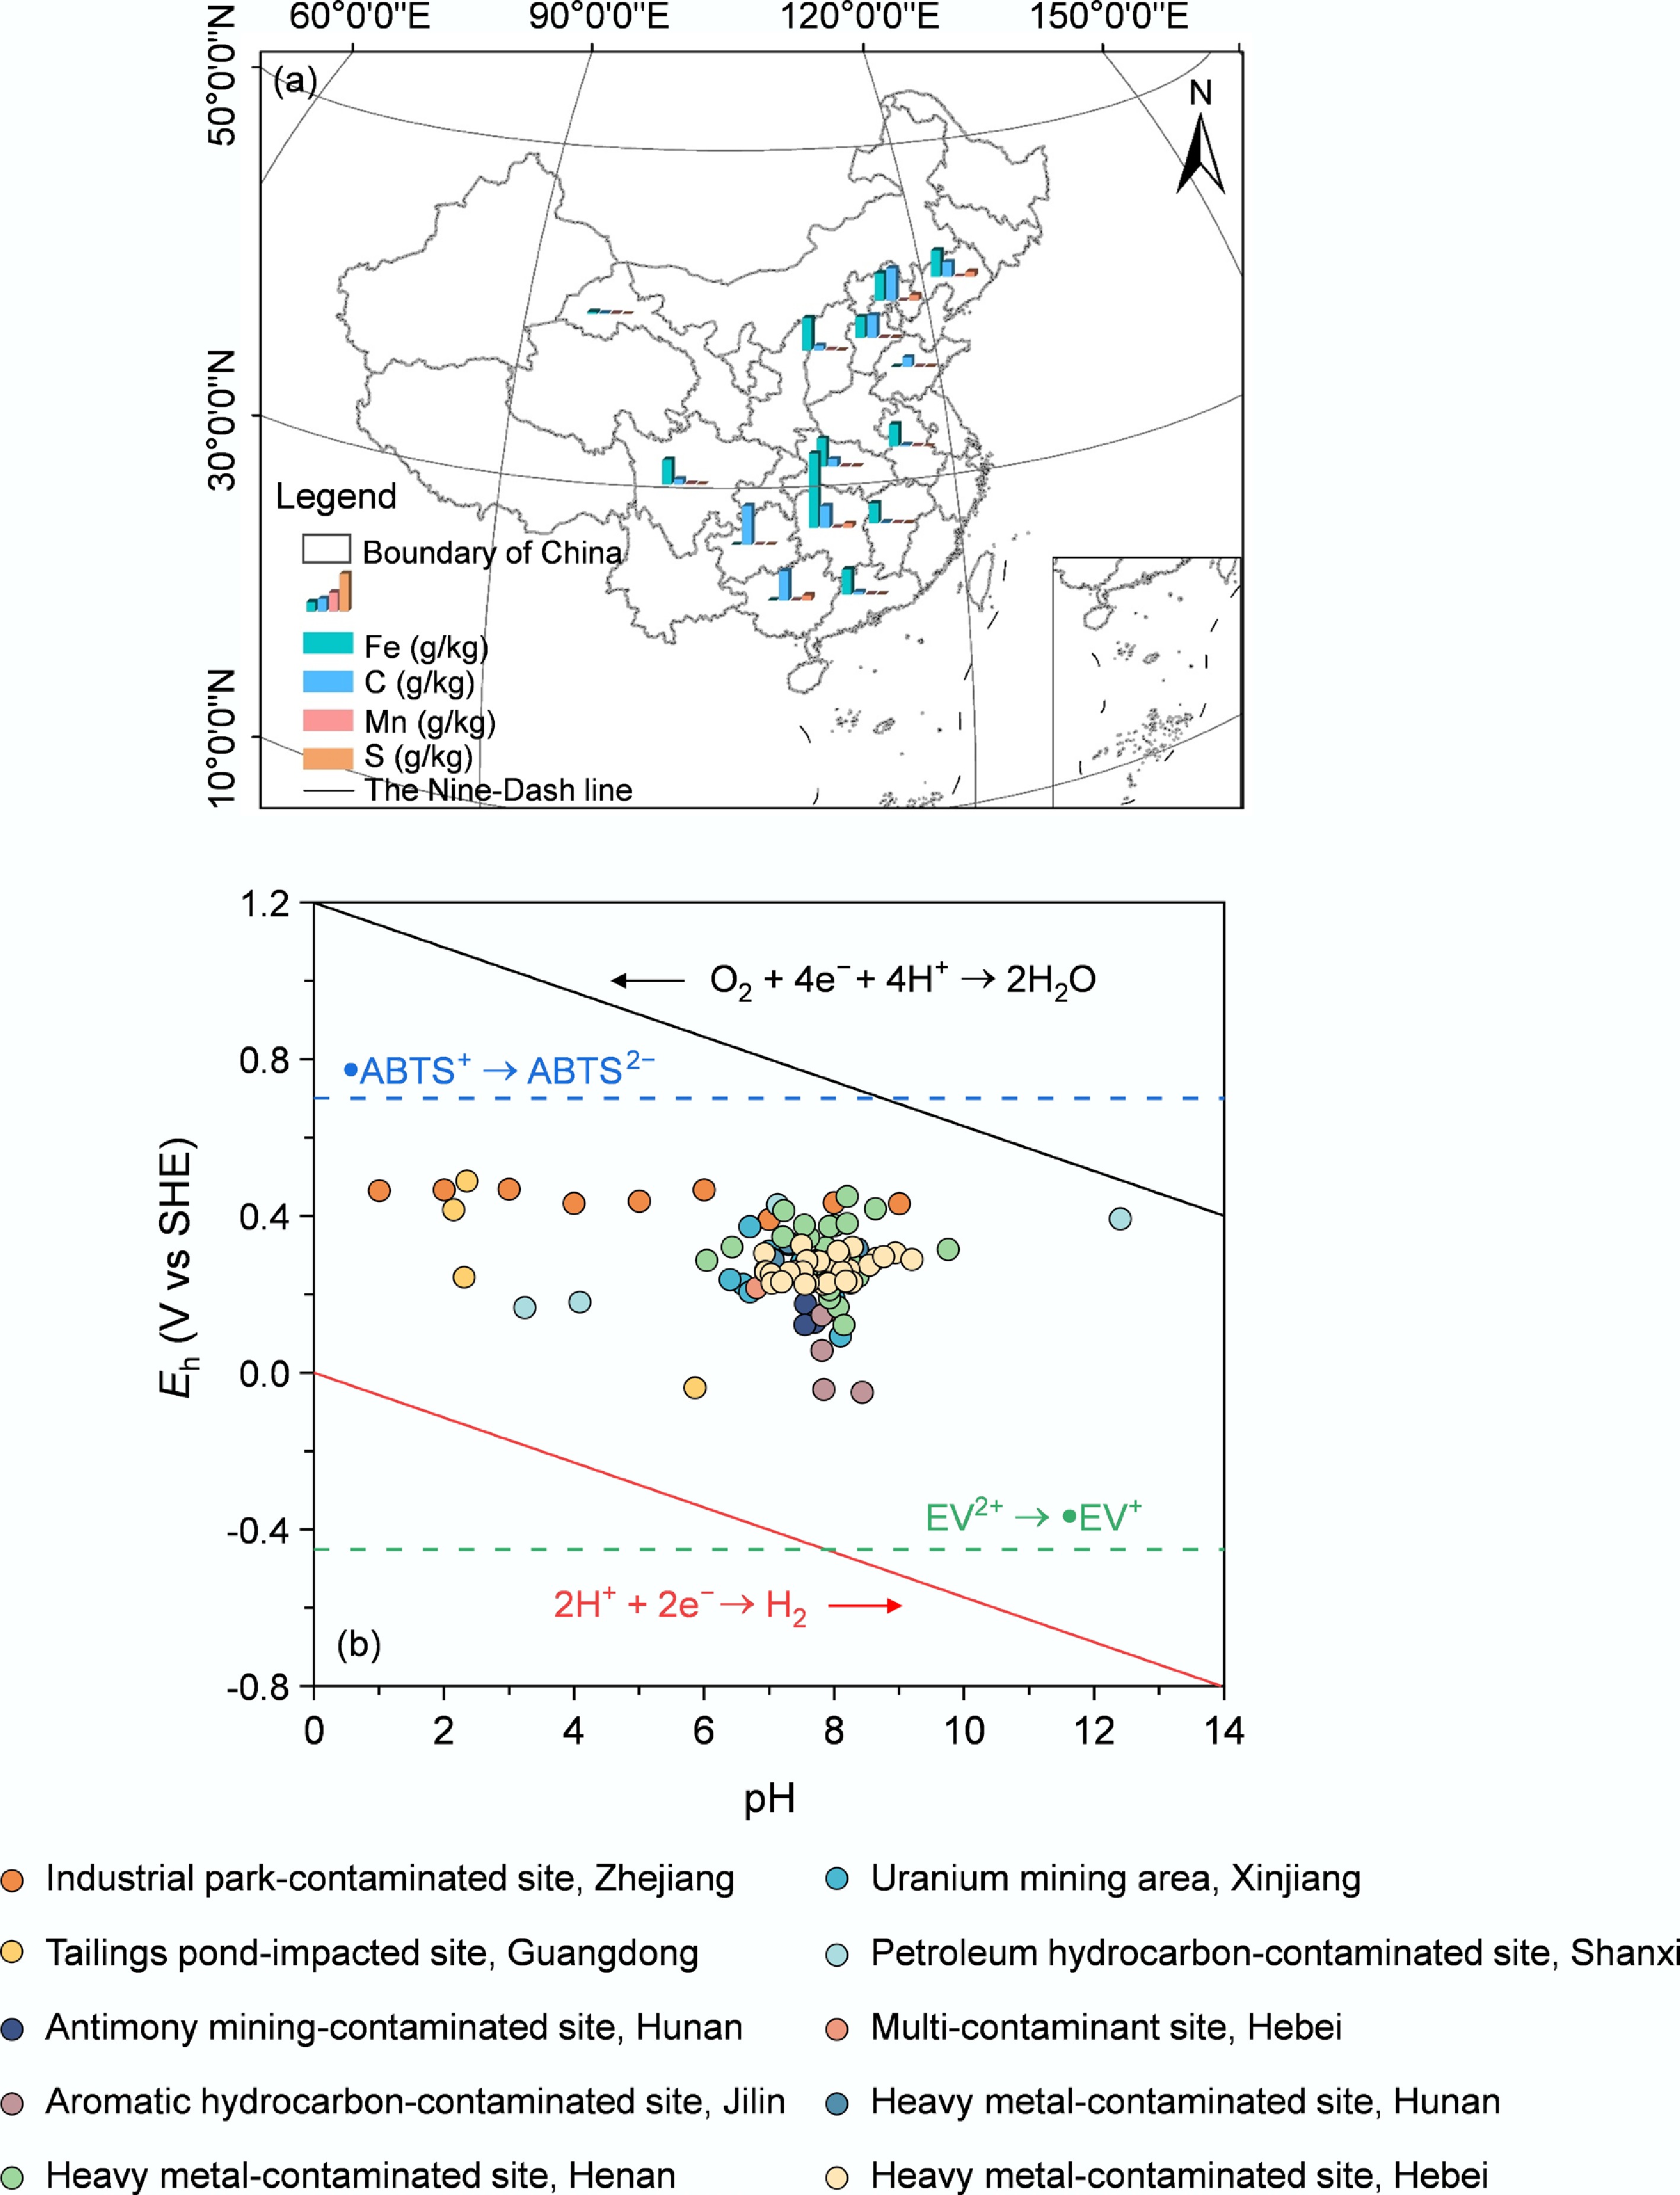
Figure 3.
EEC-related compositions in contaminated aquifers in China. (a) Contents of redox-active elements in contaminated aquifers. (b) Eh values of groundwater in contaminated aquifers. The basemap of China was obtained from the Ministry of Natural Resources CNR2024), China (GS 2024 0650).
In addition to the distribution of chemical compounds, redox potentials are also critical in determining the extent to which sediment EEC can participate in redox reactions. Both mediated electrochemical analysis and modified chemical probe methods use a potential range of −0.45 to +0.70 V (vs SHE) for EEC measurement. This range falls within the thermodynamic stability region of water, avoiding interference from side reactions such as hydrogen or oxygen evolution (Fig. 3b). To verify the environmental representativeness of this potential range, we conducted a statistical analysis of Eh-pH data from 161 groundwater samples collected from typical contaminated sites in China (Supplementary Table S2). The results show that pH values are mainly distributed between 5.0 and 9.0, with Eh values concentrated between −0.4 and +0.6 V. Most samples are in reducing to neutral oxidizing environments (Eh < 0.2 V), while a small number fall under slightly oxidizing conditions (Eh > 0.3 V). This distribution aligns well with the thermodynamic stability region of water, indicating that the selected range of −0.45 to +0.70 V not only meets theoretical requirements of EEC measurements but also reflects the actual redox state of contaminated aquifers. At these applied potentials, all Fe(III) in (oxyhydr)oxides and clays, and oxidized quinone groups in organic matter were expected to be reduced in EAC measurements[35]. All Fe(II), sulfide, and reduced quinone groups in organic matter were expected to be oxidized in EDC measurements[35]. Therefore, the maximum EEC was obtained to infer the maximum extent of redox reactions in which a sediment can participate.
Influence of sediment EEC on ISCO
-
ISCO involves injecting and transporting soluble oxidants (e.g., persulfate and H2O2) to contamination plumes. A major limitation of ISCO is the non-target consumption of oxidants before they reach the contamination plume[116]. Various redox-active substances, including Fe/Mn minerals, organic matter, and S-containing compounds, can consume oxidants through redox reactions. For example, persulfate is highly stable in sand aquifers, with a half-life of more than two years, but its half-life decreases to less than one year in sediments containing more than 2% Fe or Mn oxides[28]. By quantifying sediment EEC as an indicator of chemical heterogeneity, researchers established a linear correlation between persulfate consumption and sediment EDC[34]. The slope of 1.8 from the linear fit is close to the theoretical value of 2, indicating that this quantitative relationship accurately describes oxidant consumption by heterogeneous sediments (Eq. [2])[34]. It has also been proposed that sediment EAC may influence the long-term, slow consumption of oxidants (Eq. [3])[19].
$ \mathrm{S}_{2} \mathrm{O}_{8}^{2-} +2{\mathrm{EDC}}\rightarrow {\mathrm{SO}}_{4}^{2-} +{\mathrm{SO}}_{ 4} {\text{•}}^{ -} +2{\mathrm{EAC}} $ (2) $ \mathrm{S}_{2} \mathrm{O}_{8}^{2-} +{\mathrm{EAC}}\rightarrow {\mathrm{S}}_{2} {\mathrm{O}}_{ 8} {\text{•}}^{-} +{\mathrm{EDC}} $ (3) In addition to consuming oxidants upstream of the contamination plume, sediment EEC can activate oxidants to generate reactive oxidant species (e.g., SO4•− and •OH) for contaminant degradation. For example, when biochar was used to activate persulfate during ISCO, the degradation rate of pesticides correlated with both the EDC and EAC of biochar[117]. This indicates that EEC effectively reflects the redox properties of biochar that drive both radical and non-radical pathways for groundwater contaminant degradation in persulfate-based ISCO. Therefore, conducting EEC surveys at contamination sites is valuable for characterizing the ability of heterogeneous aquifers to consume and activate oxidants (Fig. 4).
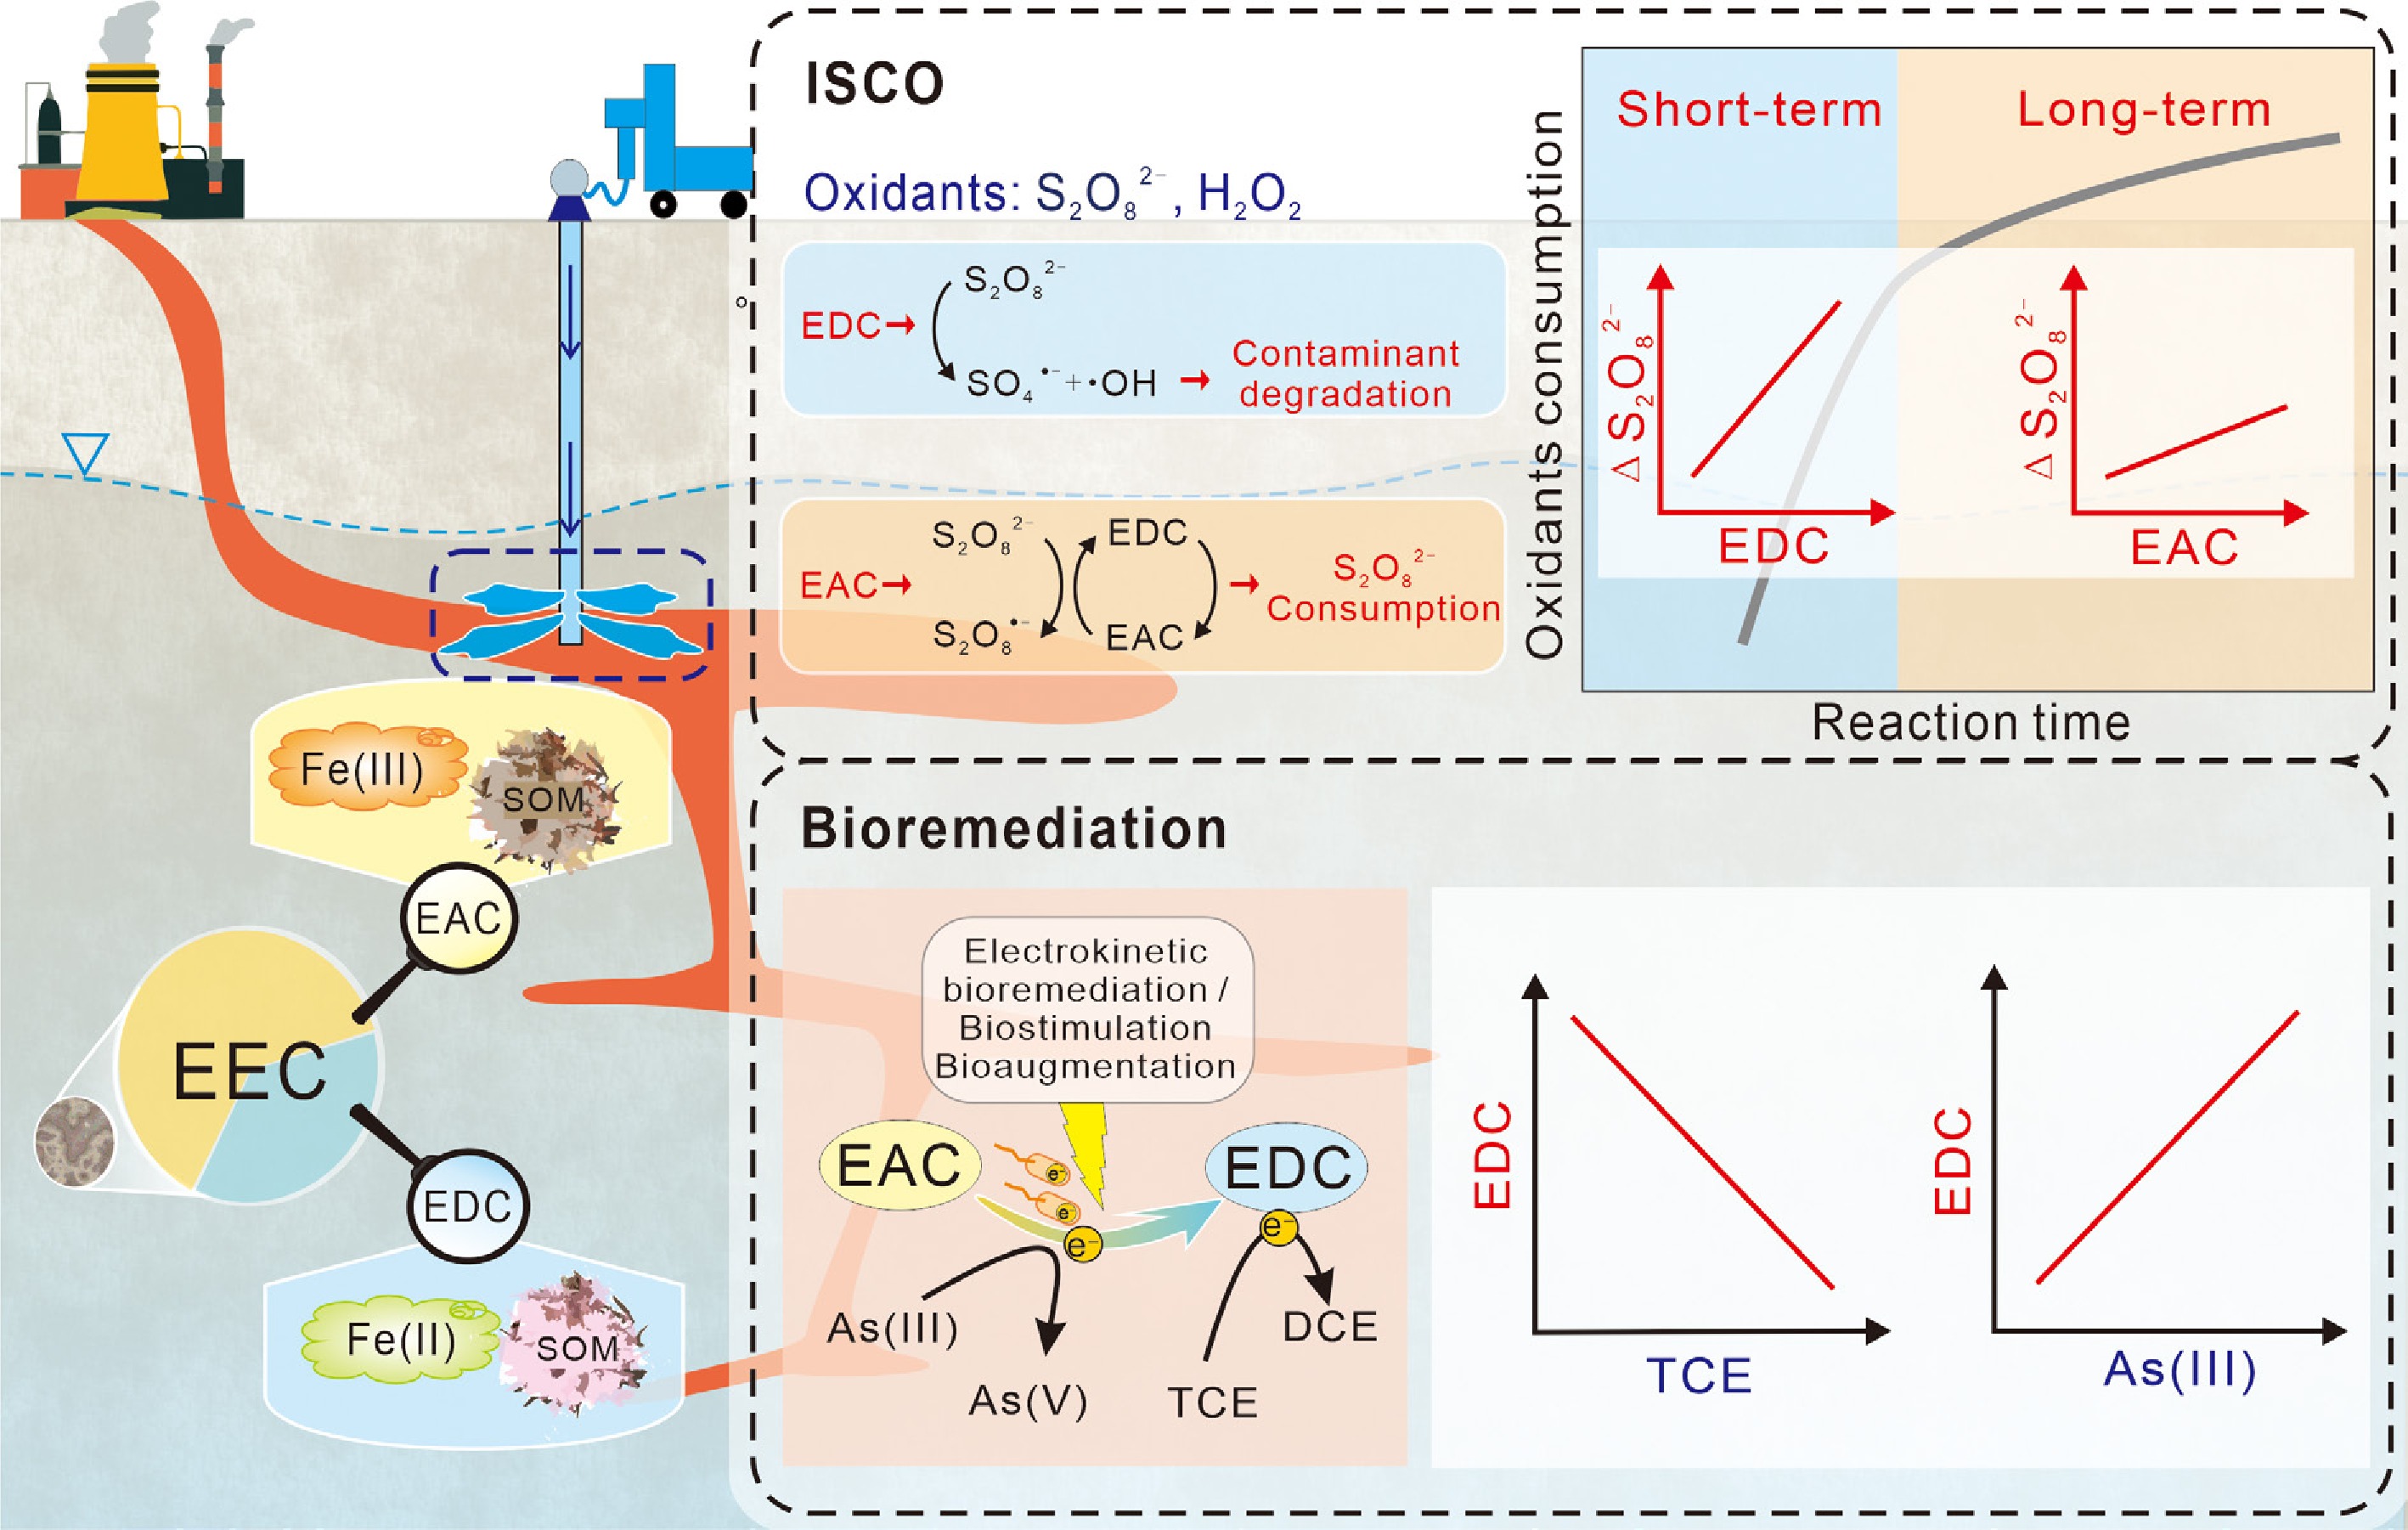
Figure 4.
Chemical controls of sediment EEC on contaminant remediation.
Influence of sediment EEC on bioremediation
-
Bioremediation is a low-cost in situ technology for soils and aquifers contaminated with organic compounds[118] and redox-active metals[119]. Microbial metabolism driven by electron transfer reactions is a key factor determining remediation performance. For example, in microbial reductive dechlorination, the dechlorination rate is positively correlated with Fe(III) concentration (i.e., EAC)[120]. Microbial reduction of Fe(III) upregulates genes associated with extracellular electron transfer, enhancing dechlorination[120]. In electrokinetically enhanced bioremediation of trichloroethylene-contaminated soils[121], the reduction of Fe(III) and trichloroethylene correlates closely with the formation of soil EDC (Supplementary Fig. S1). Initially, iron-reducing bacteria consume EAC to create a suitable reductive environment, which is subsequently colonized by organohalide-respiring bacteria[121]. In these studies, microbial oxidation of carbon sources (e.g., lactate) was the dominant energy acquisition pathway. In contrast, recent studies reveal that electrons stored in electron-donating moieties (e.g., phenolic hydroxyl and amino groups) can also support organohalide respiration[122]. This pathway is reflected by the correlation between dechlorination rates and the EDC of sediment humin[122]. For bioremediation of aquifers contaminated with redox-active metals or metalloids such as As, sediment EEC has emerged as a useful parameter for performance prediction. Researchers used variations in aquifer sediment EDC to track the oxidation of As(III) to the less toxic As(V) during biostimulation and bioaugmentation, based on observed correlations[123]. Thus, sediment EEC can serve as a simple and practical indicator for evaluating bioremediation performance (Fig. 4), without the need to explicitly identify complex reaction networks involving carbon metabolism, extracellular electron transfer, and redox transformation of contaminants.
Outlook for future research
-
Quantifying sediment chemical heterogeneity through EEC measurements has increasingly been shown to effectively describe and predict contaminant transformation and remediation in aquifers. Compared with characterizing elemental, mineral, and organic matter composition in sediments, EEC measurement is simpler and more cost-effective. Importantly, EEC can be directly linked to the rate and extent of biogeochemical transformations of contaminants, whereas bulk composition measurements often fail to provide such predictive power. Nevertheless, the application of sediment EEC is currently limited by several factors: the need for wider adoption of measurement methods, the lack of a clear mechanistic understanding of EEC-related biogeochemical processes, and the limited integration of EEC into data-driven models for predicting contaminant transformation and remediation. Addressing these challenges could significantly advance soil and groundwater contamination remediation, shifting practices from high-consumption, low-efficiency approaches toward low-consumption, high-efficiency, and sustainable strategies.
Method improvement for EEC measurement
-
A suitable method for EEC measurements, such as mediated electrochemical analysis, has been established for more than a decade[32]. However, their application in hydrogeochemistry research and remediation projects remains limited. One reason is that these methods require strict reaction conditions (e.g., anoxic environments) and a basic understanding of chemical and electrochemical principles. Therefore, developing portable equipment for easy on-site measurement is urgently needed. A recent study reported an EEC measurement device that integrates improved core electrodes, a reaction cell that maintains an anaerobic microenvironment, and data analysis software[124]. This device can be used both in the laboratory and directly at contaminated sites, enabling accurate and convenient quantification of EAC and EDC in sediments, soils, and porewater. Such portable equipment is highly valuable for geological EEC surveys and should be promoted and widely applied in future contamination remediation efforts.
Mechanistic understanding of the constitution of sediment EEC and its environmental significance
-
Currently, the environmental significance of measured sediment EEC is not fully understood. First, it remains unclear whether measured EEC accurately represents the real extent of redox reactions in aquifers under natural conditions. Generally, EEC measured using highly reactive probes or at extreme Eh values is considered to represent the maximum capacity of a soil or sediment to exchange electrons in biogeochemical processes. When measured at different Eh potentials, varied EEC values can be obtained, reflecting different reactivities. For example, Aeppli et al. reported that EAC measured at −0.14 and −0.36 V, rather than the commonly used ~−0.5 V, represents the microbial-available fraction of sediment electrons[35]. Future research should aim to identify and quantify the fractions of reactive EEC that are relevant to specific biogeochemical processes under natural thermodynamic conditions. Second, current EEC measurement methods cannot quantify certain key electron acceptors, such as O2, NO3−, and SO42−, nor electrons stored within the carbon bonds of organic matter. These electron donors and acceptors play important roles in microbial respiration and metabolism but are often not considered in the current framework of aquifer EEC. Future research should establish a unified understanding of the constitution and environmental significance of aquifer EEC, encompassing both commonly measured EEC and other important components (e.g., O2, NO3−, SO42−, and organic carbon). Third, the long-term stability of sediment EEC in subsurface environments remains unclear. Considering redox couples in sediments (e.g., Fe(II)/Fe(III) and quinone/hydroquinone), EDC and EAC can reversibly convert, resulting in a constant overall EEC[19,61,125]. However, during long-term water-rock interactions and biogeochemical processes, a fraction of EEC may be irreversibly lost or newly created. For example, enzymatic oxidation of NOM by laccase significantly decreases EEC due to irreversible oxidation of phenols[62], whereas microbial dissolution and transformation of redox-inert clay minerals may generate new EEC sites[126]. Therefore, sediment EEC is not a fixed value but a dynamic property in subsurface environments. This temporal evolution of EEC may have profound effects on the long-term remediation of contaminated aquifers. Overall, further studies are needed to gain a clearer understanding of the constitution and environmental impacts of sediment EEC in the dynamic context of contaminated aquifers.
Incorporating EEC into numerical models for process prediction in contaminated aquifers
-
Modeling solute transport, particularly reactive transport, remains challenging[116,127] due to the chemical heterogeneity of contaminated aquifers and the complexity of biogeochemical reaction networks. To better simulate the transport of redox-active solutes, it is necessary to integrate coupled hydro-biogeochemical processes. For example, recent studies have incorporated biogeochemical reactions into reactive transport models to simulate the transport of contaminants such as nitrate, Cr(VI), and Fe[128,129], as well as remediation oxidants[130]. In more recent work, researchers have incorporated EEC into the biogeochemical reactions within reactive transport models, successfully describing the transport of H2O2, dissolved O2, and nitrate[37]. Using EEC as a modeling parameter offers two key advantages. First, EEC can capture the chemical heterogeneity of aquifers without requiring detailed input of the concentrations of multiple chemical species. Second, EEC can represent complex biogeochemical reaction networks, significantly reducing modeling complexity. For instance, the reactive transport of dissolved O2 can be predicted based on the reaction of EDC with O2, without accounting for the numerous O2-consumption reactions by different electron-donating species[37,38]. Thus, incorporating EEC opens a new avenue for developing EEC-based reactive transport models, enabling improved prediction of contaminant and oxidant transport in aquifers. Building on geological EEC surveys, future research should focus on simulating EEC-incorporated reactive solute transport to guide green and sustainable remediation of contaminated aquifers.
We thank the reviewers whose input greatly improved the manuscript.
-
It accompanies this paper at: https://doi.org/10.48130/ebp-0026-0013.
-
Not applicable.
-
The authors confirm their contributions to the paper as follows: Chenglong Yu: writing – review, conceptualization and design, analysis of studies, and draft manuscript preparation; Lijun Fan: writing – research collection, analysis of studies, and draft manuscript preparation; Peng Zhang: review and editing, methodology; Yanping Cui: review and editing, methodology; Shengyan Pu: conceptualization, review and editing. All authors reviewed the results and approved the final version of the manuscript.
-
Data sharing not applicable to this article as no datasets were generated or analyzed during the current study.
-
The authors declare that they have no known competing financial interests or personal relationships that could have appeared to influence the work reported in this paper.
-
Full list of author information is available at the end of the article.
- The supplementary files can be downloaded from here.
- Copyright: © 2026 by the author(s). Published by Maximum Academic Press, Fayetteville, GA. This article is an open access article distributed under Creative Commons Attribution License (CC BY 4.0), visit https://creativecommons.org/licenses/by/4.0/.
-
About this article
Cite this article
Yu C, Fan L, Zhang P, Cui Y, Pu S. 2026. Electron exchange capacities of sediments as chemical controls on natural attenuation and engineered remediation of contaminated aquifers. Environmental and Biogeochemical Processes 2: e018 doi: 10.48130/ebp-0026-0013 

Electron exchange capacities of sediments as chemical controls on natural attenuation and engineered remediation of contaminated aquifers
- Received: 12 May 2026
- Revised: 06 June 2026
- Accepted: 24 July 2026
- Published online: 03 September 2026
Abstract: Natural attenuation and remediation of contaminants in aquifers are strongly constrained by both structural and chemical heterogeneity. While structural heterogeneity has been extensively characterized, chemical heterogeneity remains poorly quantified and insufficiently understood. In this review, we use electron exchange capacities (EEC) as a proxy for chemical heterogeneity and demonstrate how sediment EEC influences contaminant transformation and remediation performance. The evolution of EEC measurement approaches, including chemical probe and mediated electrochemical methods, is summarized. Sediment EEC is linked to the transport and transformation of groundwater contaminants through EEC-driven processes such as anaerobic microbial respiration, mineral dissolution, direct contaminant reduction, and reactive oxygen species-mediated oxidation. In terms of remediation, sediment EEC quantitatively governs the consumption and activation of injected oxidants during in situ chemical oxidation and serves as a robust parameter for evaluating bioremediation performance. Overall, EEC provides a quantitative measure of aquifer chemical heterogeneity, offering mechanistic insights into natural attenuation and guidance for green and sustainable remediation strategies. Future research should focus on methodological improvements, mechanistic elucidation of EEC composition and its environmental significance, and the development of predictive models incorporating EEC.