




-
Insights into reaction kinetics and mechanisms provide a foundation for understanding the molecular origins of chemical reactivity and for guiding the rational development of more efficient and selective synthetic transformations[1−3]. Such understanding is particularly important for reactions performed in aqueous media, where productive bond-forming processes often compete with solvent-induced decomposition pathways[4−6].
Amide bond formation is one of the most widely utilized transformations in organic synthesis and is indispensable for the preparation of pharmaceuticals, functional materials, and biologically active molecules[7−9]. Among the available synthetic methods, the Schotten–Baumann reaction remains a practical and widely adopted protocol, enabling the conversion of amines and acyl chlorides into amides under aqueous basic conditions[10,11]. Despite its practicality and widespread use, the reaction is constrained by an inherent incompatibility between the requirements of acid scavenging and acylating-agent stability[12−14]. Efficient amidation requires an aqueous basic phase to continuously neutralize the HCl generated during acylation and preserve amine nucleophilicity[12,13,15]. At the same time, highly reactive acyl chlorides readily undergo hydrolysis in water, leading to carboxylic acid formation and loss of reaction efficiency[16,17]. Consequently, successful Schotten–Baumann acylation depends on maintaining a delicate balance between productive amidation and competing hydrolysis.
To overcome the inherent incompatibility between acid scavenging and acyl chloride stability, Schotten–Baumann reactions are conventionally performed in biphasic aqueous–organic systems[10,18,19]. Although phase separation suppresses hydrolysis and preserves acylation efficiency, it does so through extensive reliance on organic solvents, including environmentally hazardous halogenated media such as CH2Cl2 and highly flammable ethers such as Et2O, as well as on interfacial transport processes[18,20,21]. As a consequence, the sustainability of the transformation remains fundamentally tied to solvent-assisted hydrolysis control, motivating the search for acylation strategies capable of operating efficiently in water without the need for a separate organic phase[22−24].
Acyl fluorides have recently attracted considerable interest owing to their unique synthetic versatility, as they can serve as acyl ('RCO'), alkyl/aryl ('R'), and fluorine ('F') sources in a wide range of organic transformations[25−27]. In particular, they have emerged as highly attractive acylating agents because fluorine incorporation imparts unique electronic and physicochemical properties[28−30], giving rise to an exceptional combination of hydrolytic stability and acyl-transfer reactivity[26,31,32]. The synthetic utility of acyl fluorides in amide bond formation has been demonstrated in several studies. SuFEx-mediated amidation enables the in situ conversion of carboxylic acids into acyl fluorides for the synthesis of sterically hindered and electron-deficient amides[33], while PFP-mediated deoxyfluorination–amidation proceeds through in situ-generated acyl fluoride intermediates[34]. These approaches demonstrated the utility of acyl fluorides for amide synthesis but relied on their in situ generation and subsequent amidation in organic media. In contrast, the present study examines acyl fluoride amidation in water, with particular focus on the origin of the unusual selectivity observed under aqueous carbonate-mediated conditions. Notably, K2CO3, which afforded only 4% yield in PFP-mediated amidation in MeCN[34], promotes highly selective N-functionalization of acyl fluorides in water in the present system. Compared with acyl chlorides, their greater compatibility with aqueous media removes a fundamental barrier to efficient acylation under aqueous conditions[35,36]. However, the exceptional reactivity and selectivity observed in water cannot be explained solely by the hydrolytic stability or electrophilic character of the acyl fluoride[37,38]. Instead, accumulating evidence suggests that reaction selectivity is governed by factors beyond the intrinsic properties of the acylating agent, particularly the inorganic cation and the surrounding aqueous solvation environment[36,39,40]. Despite their potential importance, the mechanistic roles of these factors in controlling acyl transfer and determining the balance between productive N-functionalization and competing hydrolysis remain largely unexplored.
Herein, we report a kinetic and mechanistic investigation of carbonate-mediated amidation of amines with acyl fluorides in water. By integrating reaction kinetics, spectroscopic characterization, control experiments, and density functional theory (DFT) calculations, we identify a water-solvated coordination environment resembling an 18-crown-6–K+ complex, arising from the cooperative assembly of an acyl fluoride, a potassium ion, and water. This unique structural motif enables carbonate-driven acyl transfer and the formation of a hydrolytically stable carbonic mixed anhydride, thereby achieving highly selective N-functionalization under fully aqueous conditions. More broadly, this work uncovers a previously unrecognized mode of aqueous acyl transfer and identifies cooperative acyl fluoride–K+–H2O assembly as a key determinant of reaction pathway selection and chemoselectivity in water, establishing a viable mechanistic alternative to classical Schotten–Baumann chemistry (Fig. 1).
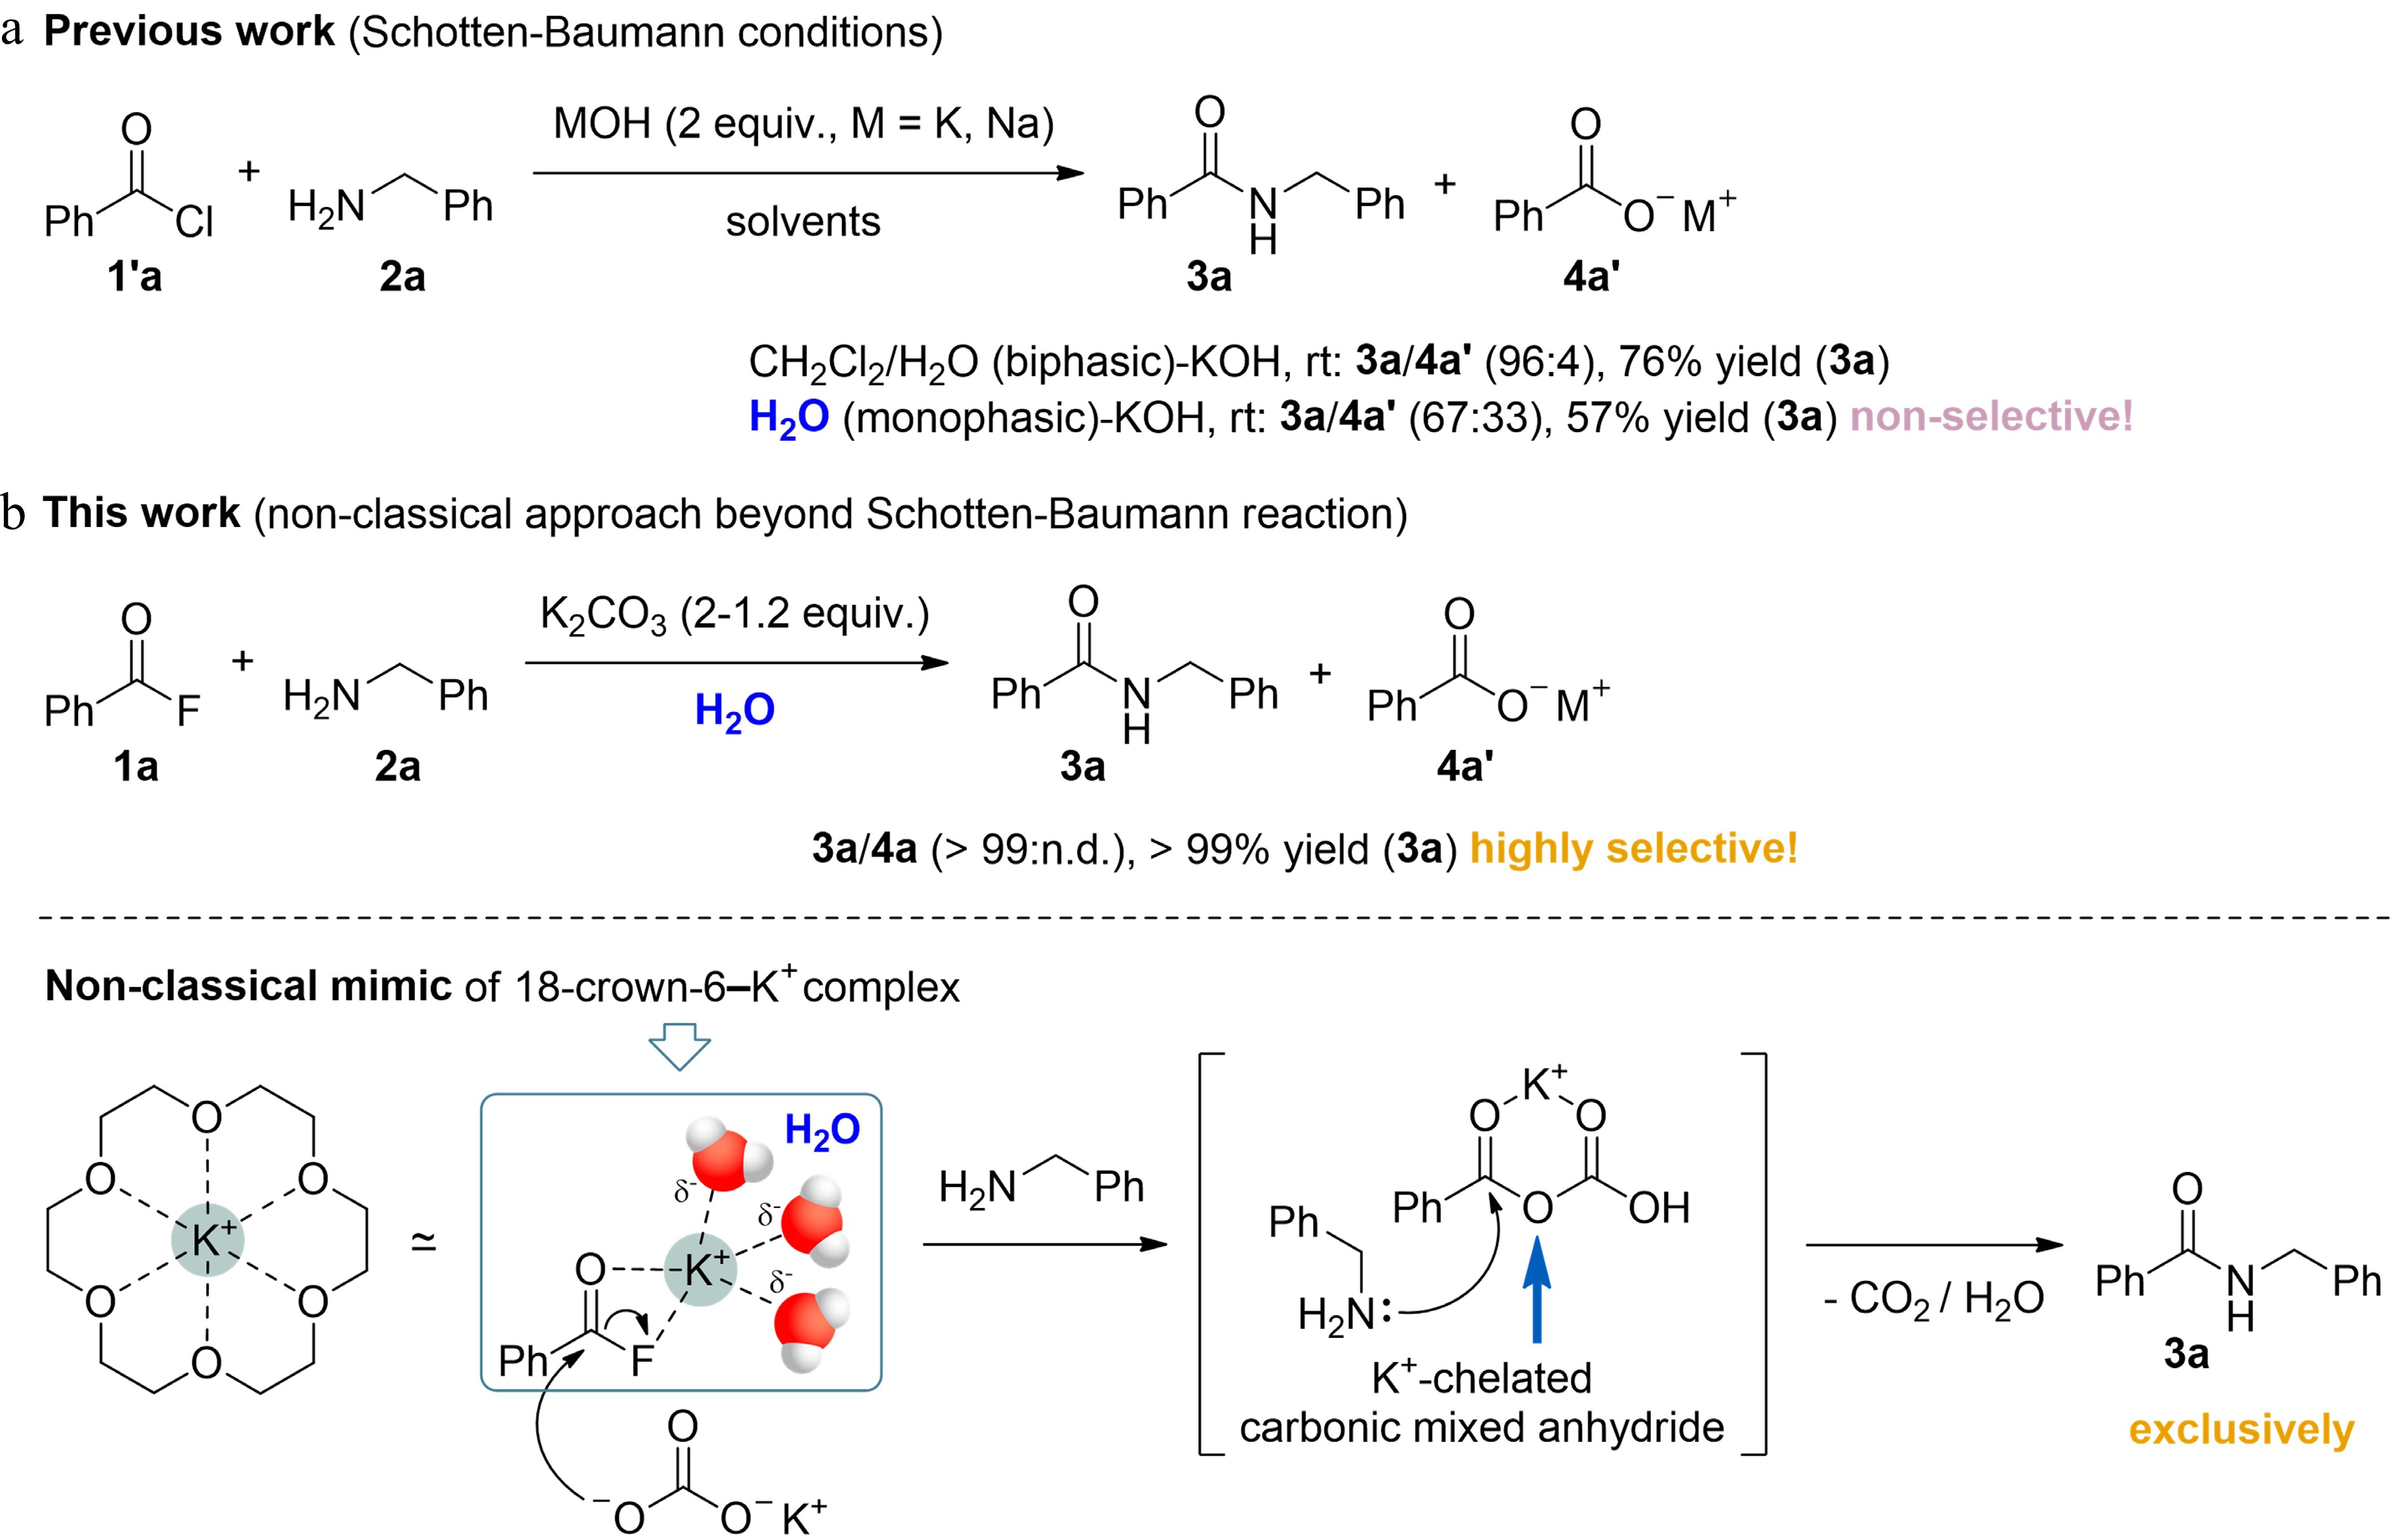
Figure 1.
(a) Classical Schotten–Baumann reaction employing a typical biphasic system (CH2Cl2/H2O, KOH). (b) Beyond Schotten–Baumann chemistry: a non-classical 18-crown-6–K+ mimic converts carbonate from a simple acid scavenger into a nucleophilic acyl-transfer partner, generating a hydrolytically stable carbonic mixed anhydride that redirects acyl-transfer reactivity toward productive N-functionalization while suppressing hydrolytic side reactions.
-
All commercially available reagents were used as received unless otherwise specified. Acyl chlorides used for the synthesis of the corresponding acyl fluorides, benzoyl fluoride (> 98.0%, GC), and amines (> 99.0%, GC) were purchased from Tokyo Chemical Industry Co., Ltd. (TCI). Anhydrous potassium fluoride (99%) was purchased from Thermo Scientific Chemicals, and anhydrous potassium carbonate (99.5%) was purchased from Daejung Chemicals & Metals Co., Ltd. Deionized water was used as the reaction solvent for all aqueous amidation reactions.
Experimental procedures
General procedure A for the synthesis of acyl fluorides 1
-
Potassium fluoride (1.0 mmol, 2.0 equiv.) was added to a stirred solution of the corresponding acyl chloride (0.5 mmol) in anhydrous acetone (2.5 mL, 0.2 M). The reaction mixture was stirred at room temperature for the specified time. After completion of the reaction, the resulting inorganic salts were removed by filtration, and the filtrate was purified according to the previously reported procedure to afford the corresponding acyl fluoride 1.
General procedure B for the synthesis of amides 3
-
To a stirred solution of amine 2 (1.0 mmol) in deionized water (5 mL, 0.2 M) were added potassium carbonate (K2CO3, 1.2 mmol, 1.2 equiv.) and acyl fluoride 1 (1.0 mmol, 1.0 equiv.). The mixture was stirred at room temperature for 1 h. Upon completion of the reaction, the reaction mixture was acidified with 1 N HCl, resulting in precipitation of the corresponding amide. The resulting solid was isolated by vacuum filtration, washed thoroughly with water, and dried under reduced pressure to afford the corresponding amide 3.
-
To delineate the factors governing selective N-functionalization, the reaction of benzoyl fluoride 1a with benzylamine 2a was systematically investigated, with the corresponding benzoyl chloride 1′a serving as a comparative reference, under a range of solvent and base conditions (Table 1). Under classical Schotten–Baumann conditions employing a biphasic CH2Cl2-H2O or Et2O-H2O system, hydrolysis of benzoyl chloride 1′a was effectively suppressed, with benzoic acid 4a formed in less than 10% yield (particularly in the CH2Cl2–H2O biphasic system, where the yield of 4a was below 5%). Nevertheless, the desired amide 3a was obtained in only moderate yields (61%–76%), presumably owing to the inherent limitations of biphasic operation, where productive acylation relies on interfacial mass transfer between the aqueous and organic phases (entries 1–2). To eliminate the use of environmentally hazardous halogenated solvents such as CH2Cl2 and highly flammable solvents such as Et2O, the reaction was subsequently conducted in water as the sole reaction medium using benzylamine 2a in the presence of 2.0 equivalents of KOH. Under these conditions, productive amidation was markedly diminished, affording 3a in only moderate yield (57%) together with benzoic acid 4a (19% yield), indicating that hydrolysis became the dominant competing pathway (entry 3). To our surprise, despite the superior hydrolytic stability of benzoyl fluoride relative to benzoyl chloride, the reaction exhibited a comparable level of hydrolysis, affording product distributions similar to those obtained with the corresponding acyl chloride (entry 4). Interestingly, the reaction outcome changed dramatically when hydroxide was replaced with potassium carbonate (K2CO3). Under identical conditions, benzoyl fluoride 1a exhibited an abrupt enhancement in selectivity, furnishing amide 3a in near-quantitative yield while completely suppressing the formation of benzoic acid 4a (entry 6). In contrast, the corresponding acyl chloride remained susceptible to hydrolysis, revealing a fundamental difference in reactivity between the two acylating agents (entry 5). Notably, reducing the amount of K2CO3 from 2.0 to 1.2 equivalents afforded the optimal result, delivering amide 3a in > 99% yield while completely eliminating the formation of the hydrolytic by-product 4a, with the reaction reaching completion within 1 h. Under otherwise identical conditions, replacing K2CO3 with KOH markedly reduced both the yield of amide 3a and the chemoselectivity of the reaction (entry 8). These results identify carbonate as an essential promoter of productive N-functionalization.
Table 1. Optimization of reaction conditionsa.
Entry Halide source Base (equiv.) Solvent Ratio of 3a : 4a Yield of 3a (%) 1 Cl (1′a) KOH (2.0) CH2Cl2/H2O (v/v = 1:1) 96:4 76 2 Cl (1′a) KOH (2.0) Et2O/H2O (v/v = 1:1) 90:10 61 3 Cl (1′a) KOH (2.0) H2O 67:33 57 4 F (1a) KOH (2.0) H2O 68:32 65 5c Cl (1′a) K2CO3 (2.0) H2O 80:20 72 6c F (1a) K2CO3 (2.0) H2O > 99:n.d. > 99d 7c F (1a) K2CO3 (1.2) H2O > 99:n.d. > 99d 8 F (1a) KOH (1.2) H2O 84:16 83 
a General conditions: benzoyl halide (1a or 1′a, 1 mmol), base (1.2–2 equiv.), solvent (0.2 M), rt, 6 h. b Yields and product ratios were determined by 1H NMR spectroscopy analysis using 1,3,5-trimethoxybenzene as an internal standard. c Reaction time: 1 h. d Isolated yield. Having established the optimized conditions, the substrate scope of the aqueous N-functionalization protocol was investigated. Acyl fluorides bearing para-substituted aryl groups with either electron-withdrawing (–Br, –CN, –NO2) or electron-donating (–OMe) substituents underwent smooth coupling with benzylamine, and high selectivity for N-functionalization was maintained irrespective of the electronic nature of the aryl substituents. Broad substrate compatibility was further demonstrated with structurally diverse amines, including simple primary amines and chiral primary amines, all of which furnished the desired amides in excellent yields. Notably, methylamine hydrochloride, a bench-stable surrogate for volatile methylamine, proved to be an equally competent coupling partner. Even when employed directly under the standard aqueous basic conditions, without prior liberation of the free amine, it afforded the corresponding amide 3f in > 99% yield. The scope was further extended to acyclic secondary amines with different steric and electronic properties. Dimethylamine, diisopropylamine, diphenylamine, and dibenzylamine furnished the corresponding tertiary amides 3l–3o in 98%–99% isolated yields. Notably, both sterically hindered diisopropylamine and the less nucleophilic diphenylamine underwent efficient amidation without appreciable competing hydrolysis, showing that increased steric demand and reduced amine nucleophilicity did not significantly compromise the selectivity of the aqueous carbonate-mediated reaction.
Overall, the protocol accommodated electronically differentiated acyl fluorides 1 and structurally diverse primary and secondary amines 2, providing the corresponding amides 3 in generally high yields (Fig. 2).
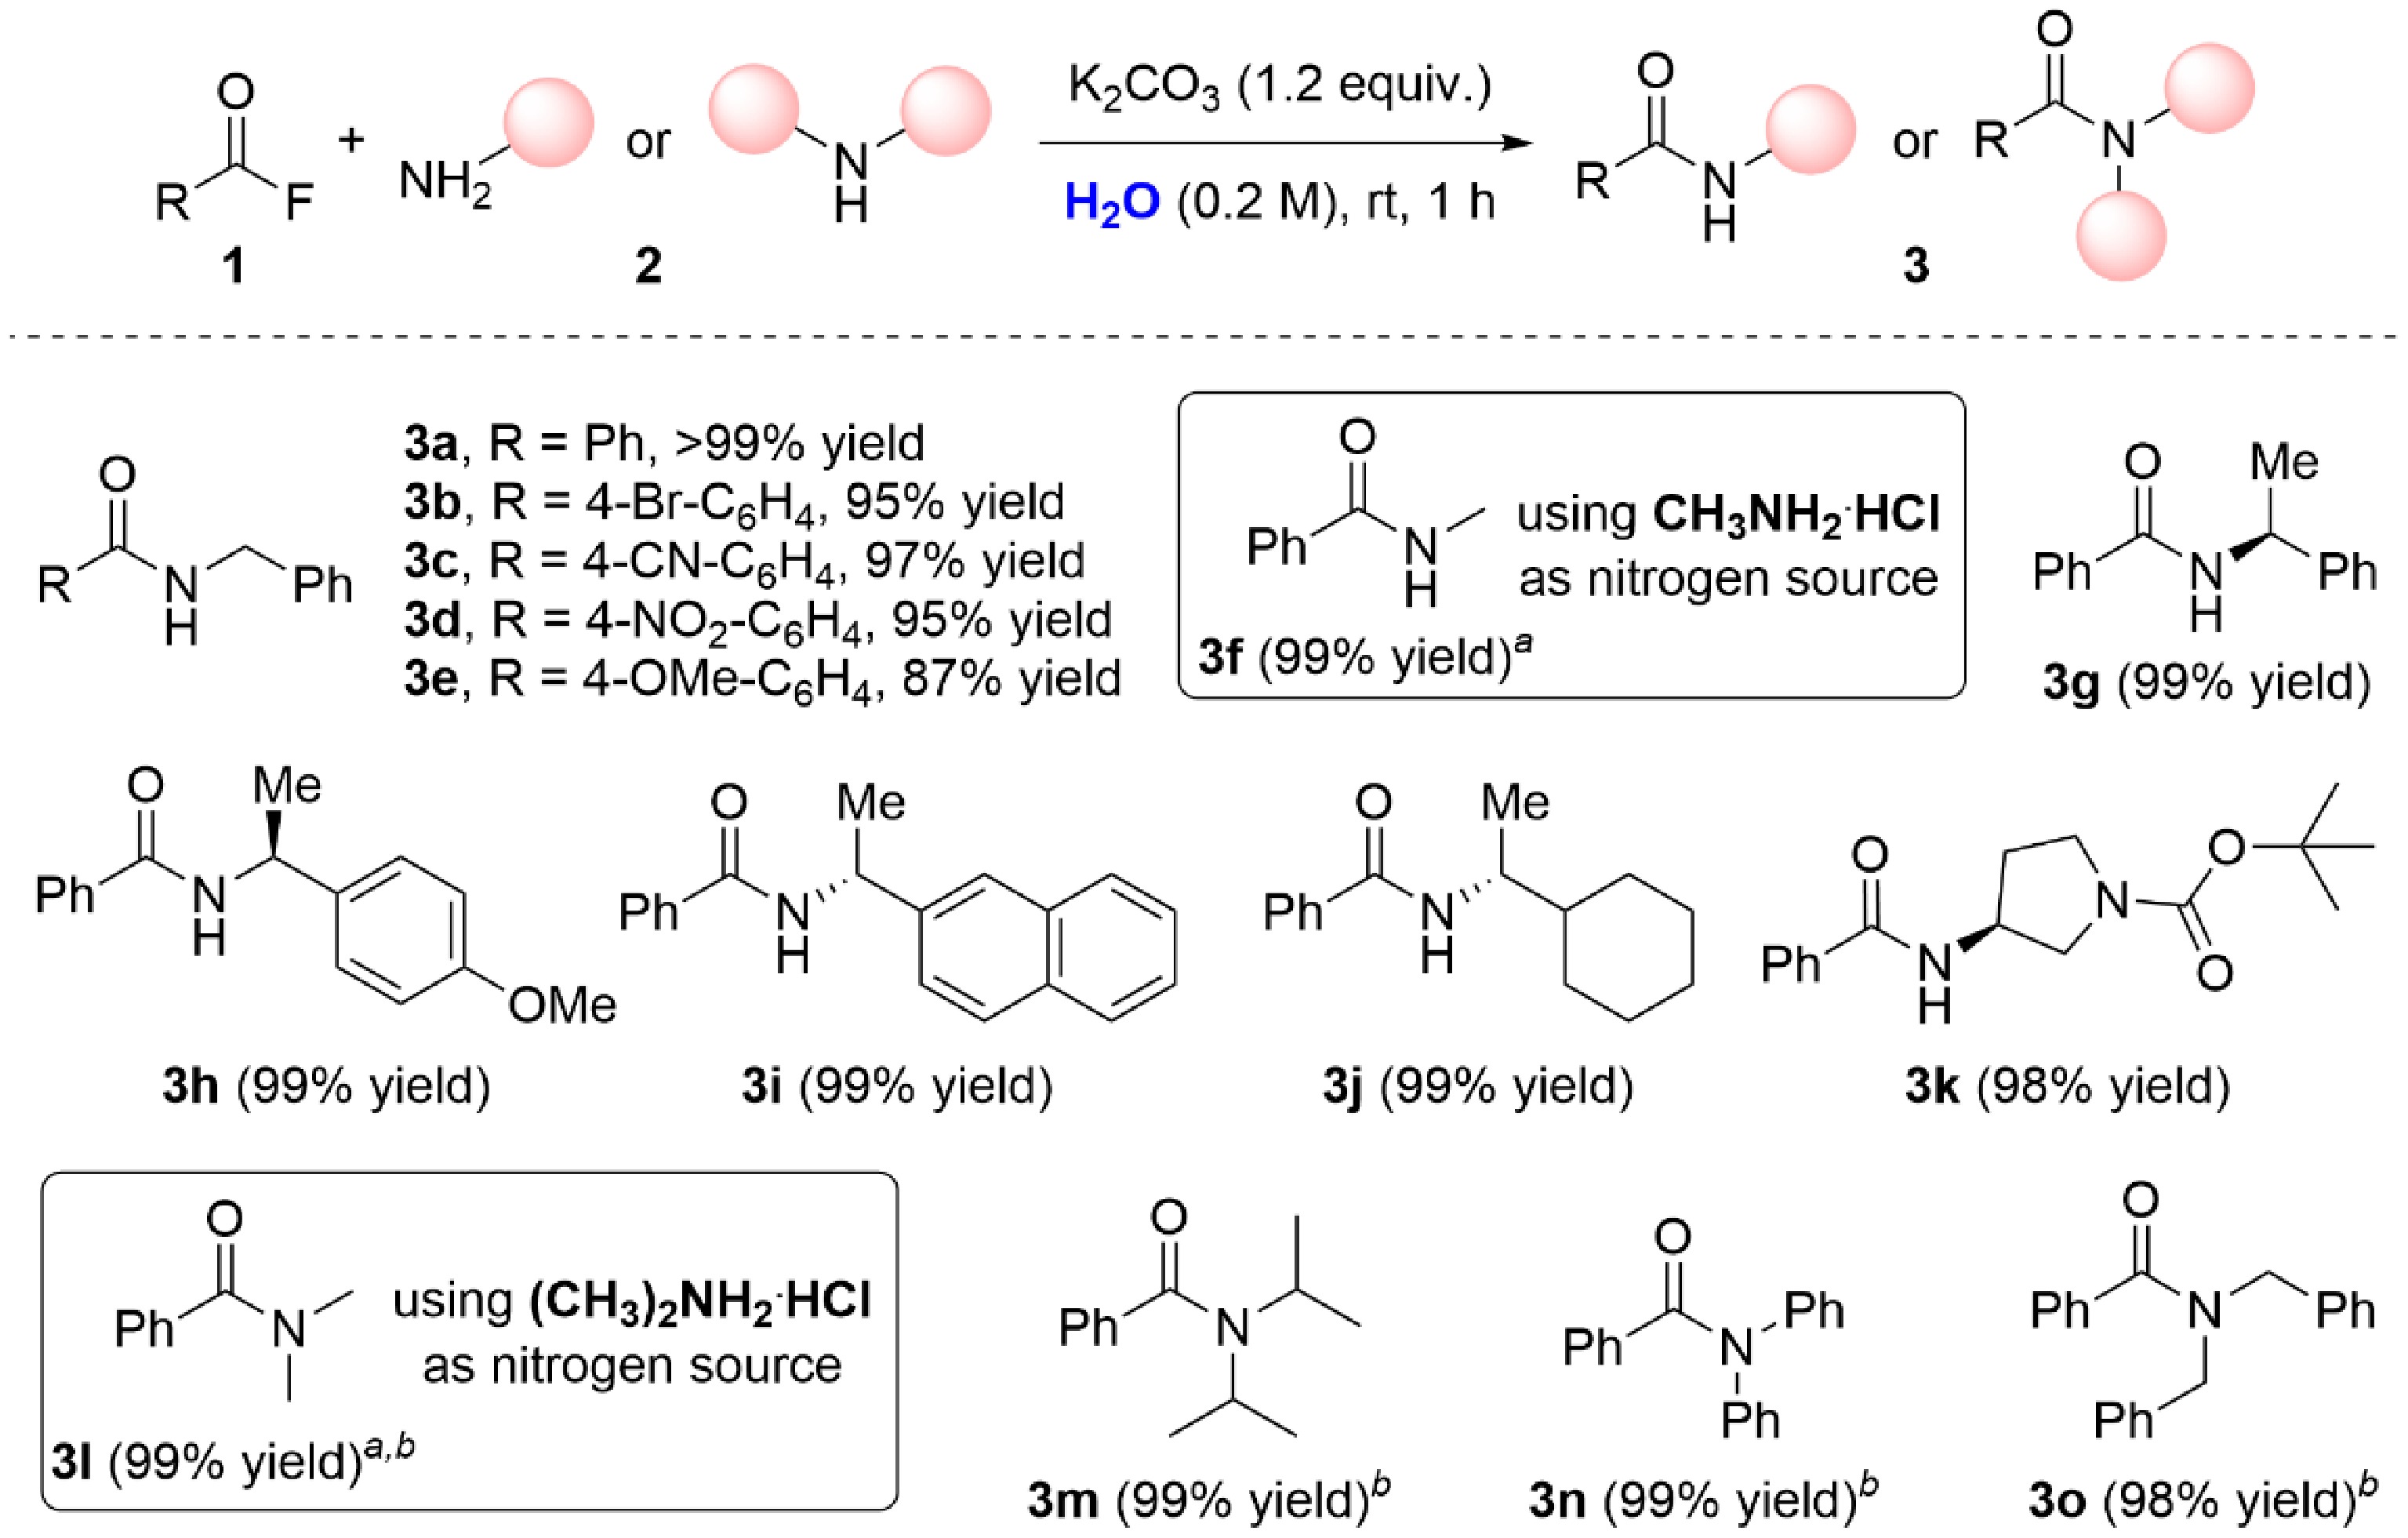
Figure 2.
Reaction scope and general conditions: acyl fluoride 1 (1 equiv.), amine 2 (1 mmol), K2CO3 (1.2 equiv.), H2O (0.2 M, 5 mL), rt, 1 h. Yields refer to isolated yields. a Using 2.2 equiv. of K2CO3. b Reaction time is 6 h.
-
To evaluate this non-classical alternative to the Schotten–Baumann reaction, the reactivity of benzoyl fluoride 1a under aqueous carbonate and hydroxide conditions was directly compared with that of the corresponding benzoyl chloride 1′a. The initial screening studies revealed that benzoyl chloride 1′a remained highly susceptible to hydrolysis under aqueous conditions, whereas benzoyl fluoride 1a exhibited markedly higher chemoselectivity toward productive N-functionalization in the presence of K2CO3. To examine the kinetic basis of this difference, time-course experiments were performed for the reaction of benzoyl fluoride 1a with benzylamine 2a using K2CO3 or KOH under identical aqueous conditions (Fig. 3).
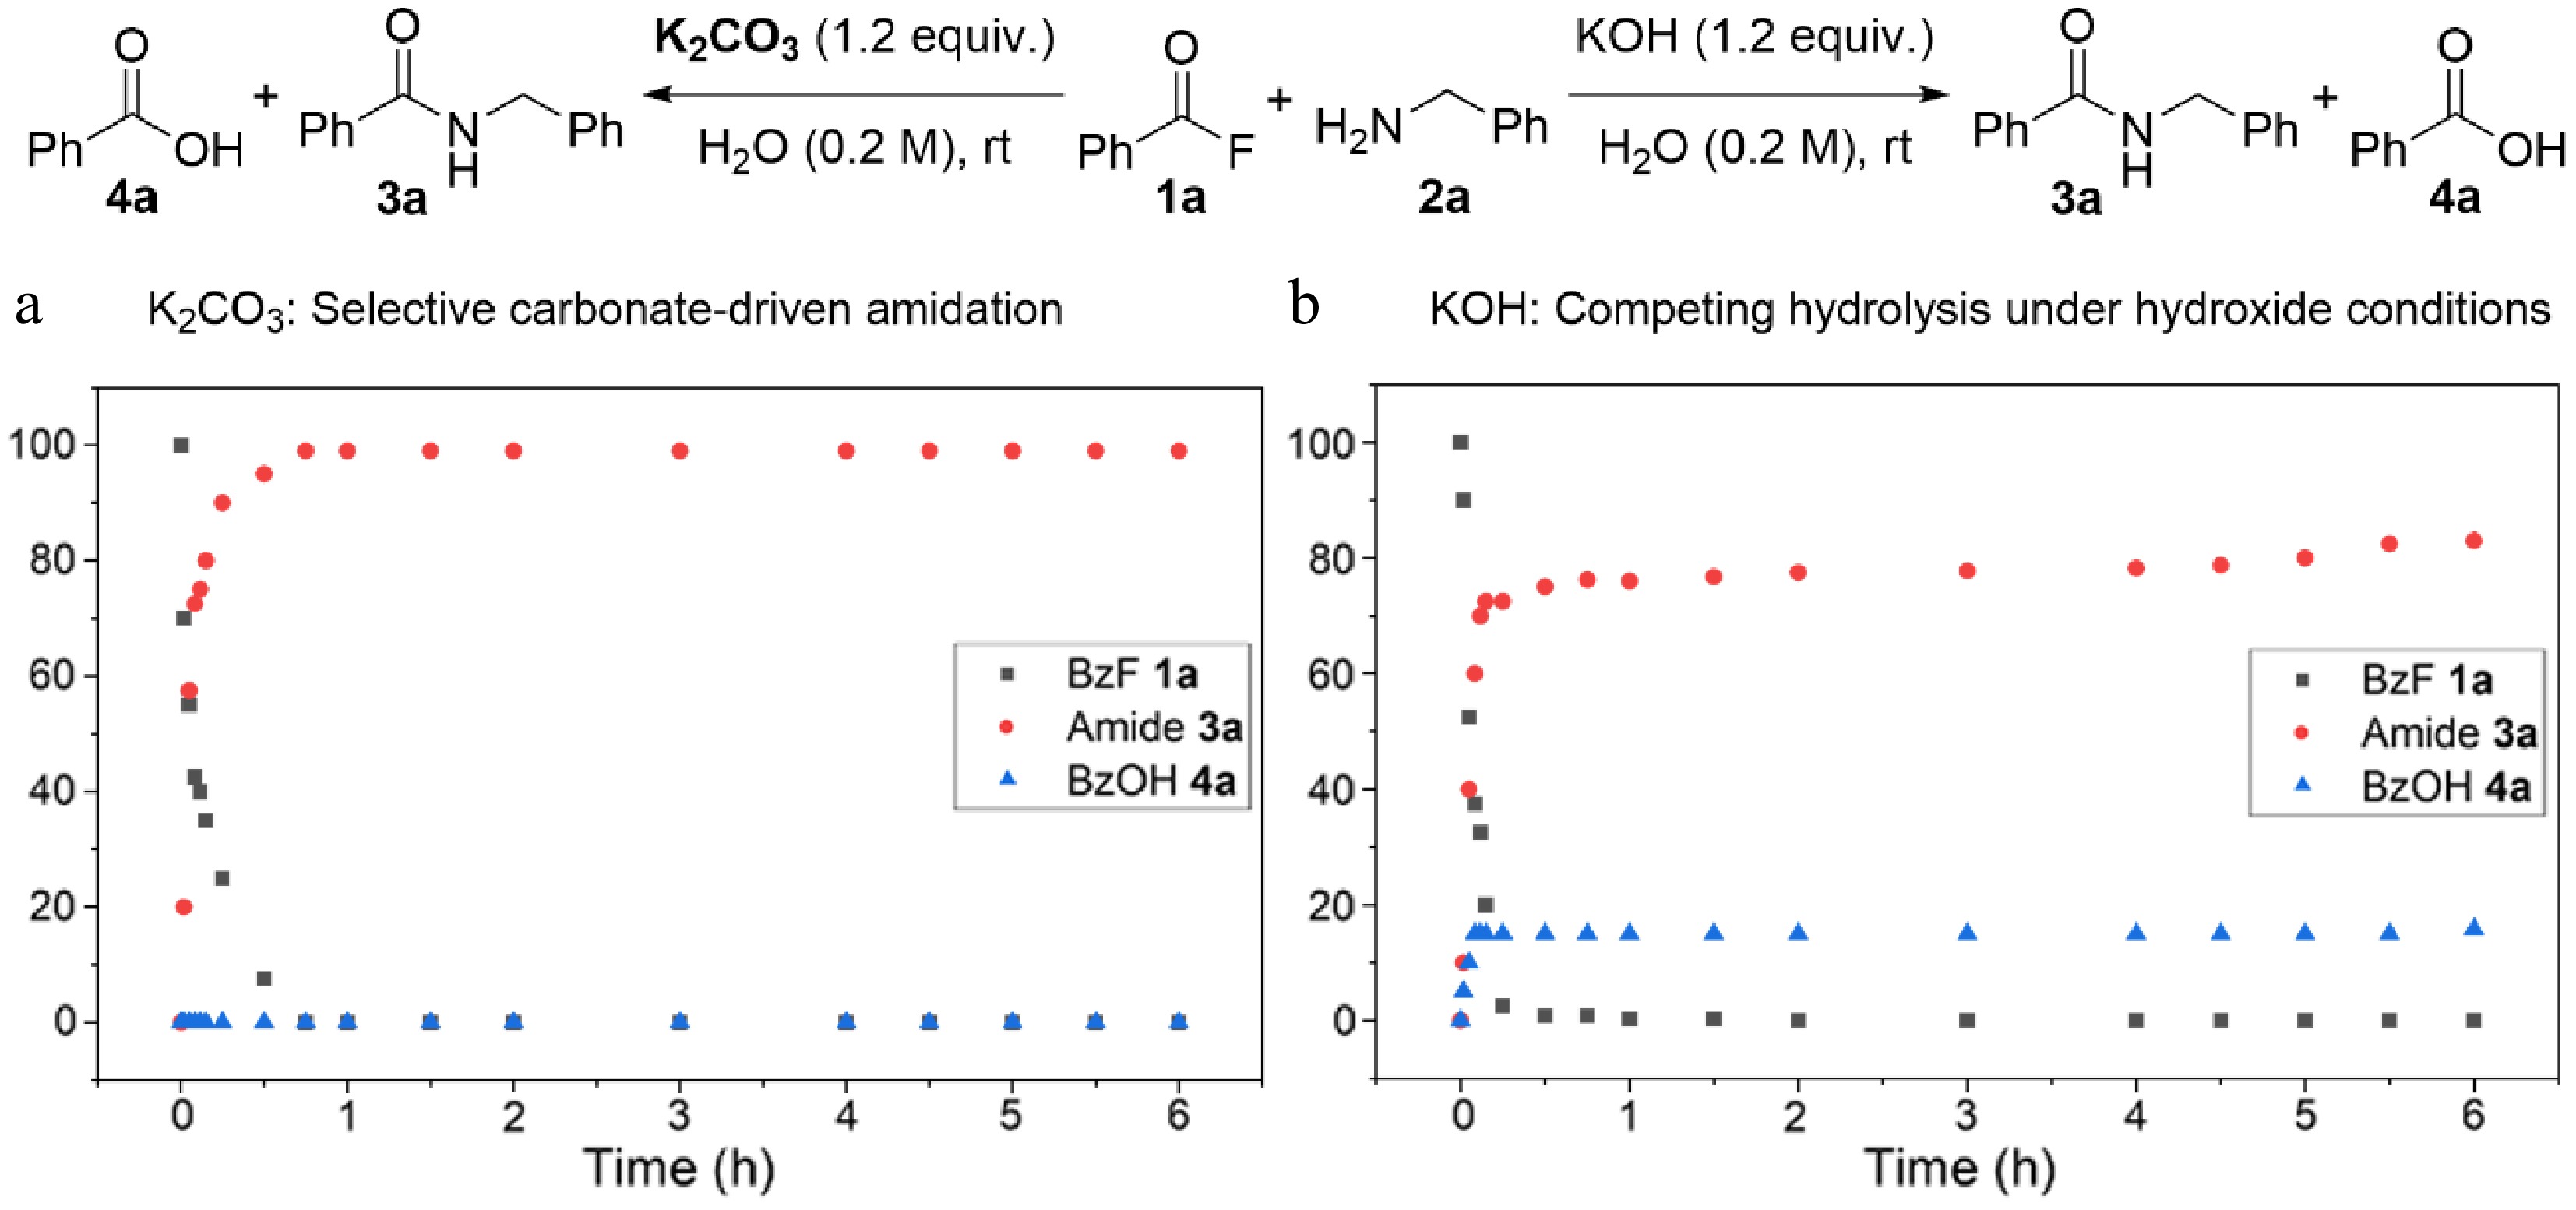
Figure 3.
Time-course profiles for the aqueous amidation of benzoyl fluoride 1a with benzylamine 2a using (a) K2CO3 or (b) KOH. The K2CO3-mediated reaction rapidly affords amide 3a in near-quantitative yield within 0.75–1 h without detectable formation of benzoic acid 4a, whereas the KOH-mediated reaction is accompanied by competing hydrolysis, leading to the accumulation of benzoic acid 4a and requiring approximately 2 h for complete conversion.
Under K2CO3-mediated conditions, rapid consumption of benzoyl fluoride 1a was accompanied by near-quantitative formation of amide 3a within 0.75–1 h, whereas benzoic acid 4a remained undetectable throughout the reaction course. This result indicates that hydrolysis of the acyl fluoride is effectively suppressed, thereby favoring productive N-functionalization over the competing hydrolysis pathway. In sharp contrast, replacement of K2CO3 with KOH led to rapid formation of benzoic acid 4a, which reached 15%–16% during the initial stage of the reaction and persisted throughout the reaction course. Although amide 3a was formed concurrently, complete consumption of benzoyl fluoride 1a required approximately 2 h, and the final yield of amide 3a reached only 83%.
To quantitatively compare the reaction profiles, the early-time decay of benzoyl fluoride 1a (0–15 min) was fitted to a single-exponential decay function,
$ {\left[\mathbf{1a}\right]}_{t}/{\left[\mathbf{1a}\right]}_{0}={\text{e}}^{-{{k}_{\text{app}}}t} $ These results show that the exceptional chemoselectivity under K2CO3-mediated aqueous conditions does not arise from accelerated consumption of benzoyl fluoride. Instead, substrate consumption is directed predominantly toward productive N-functionalization with minimal competing hydrolysis. This behavior differs markedly from the KOH-mediated reaction, in which amidation and hydrolysis occur concurrently. Unlike the classical Schotten–Baumann reaction, in which hydroxide promotes competing hydrolysis of the acyl halide, this distinct reaction-pathway partitioning supports the involvement of a non-classical carbonate-mediated acyl-transfer process. As discussed below, spectroscopic and computational studies further support the involvement of a carbonic mixed anhydride as the key intermediate in this pathway.
The striking reversal in reactivity observed under carbonate-mediated aqueous conditions cannot be rationalized solely by the intrinsic leaving-group ability of the acyl halide (Fig. 4). Under classical Schotten–Baumann conditions, acyl chlorides and acyl fluorides exhibited comparable amidation efficiencies together with similar levels of hydrolytic by-product formation, despite the substantially stronger C–F bond of acyl fluorides, as reflected by the markedly higher bond dissociation energy (BDE) of the C–F bond (121.6 kcal·mol−1) relative to the corresponding C–Cl bond in acyl chlorides (83.8 kcal·mol−1). These results indicate that hydroxide-mediated acyl transfer and hydrolysis dominate the reaction outcome irrespective of the intrinsic leaving-group properties of the acyl halide (see Table 1, entries 3–4).
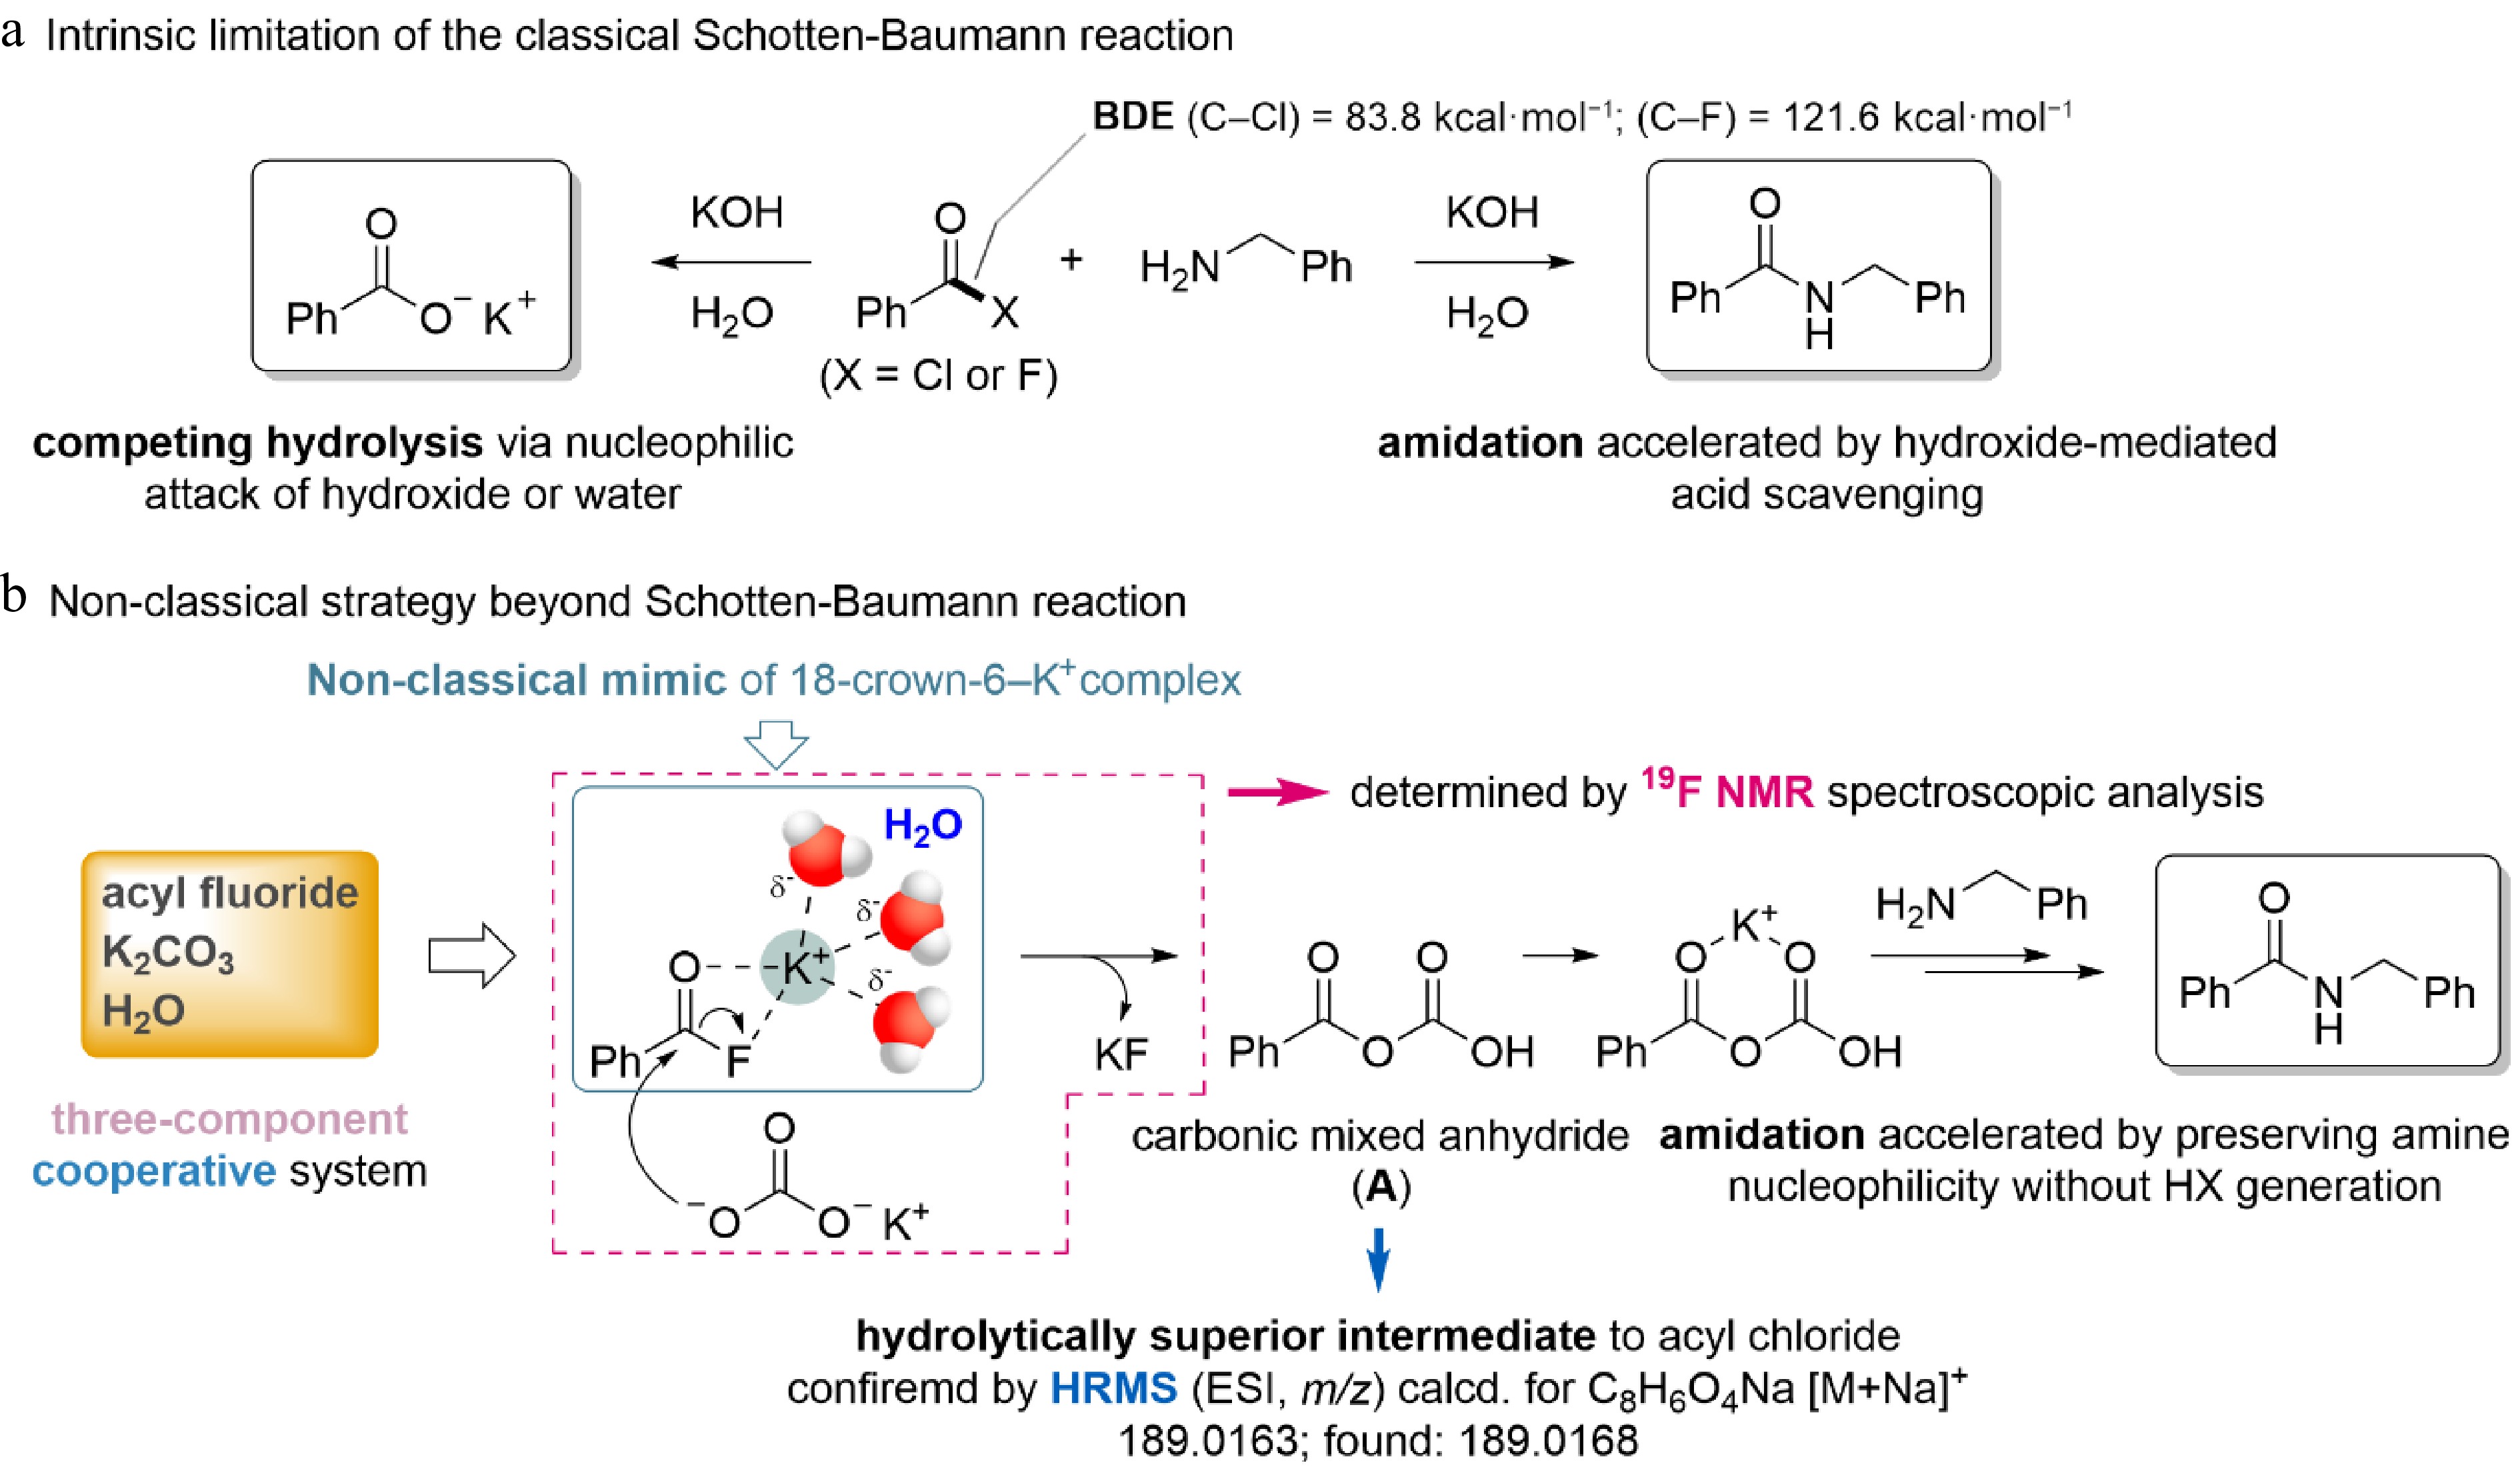
Figure 4.
Mechanistic distinction between the classical Schotten–Baumann reaction and the carbonate-mediated non-classical acyl-transfer pathway.
In sharp contrast, this reactivity trend was completely reversed in the presence of K2CO3. Although acyl fluorides are intrinsically less reactive acylating agents than acyl chlorides, they exhibited exceptional N-functionalization efficiency under aqueous carbonate-mediated conditions. To experimentally examine the role of the alkali-metal countercation, Li2CO3, Na2CO3, K2CO3, and Cs2CO3 were evaluated under otherwise identical conditions, with carbonate maintained as the common anion (Supplmentary File 1: Table S3). Li2CO3 afforded amide 3a in 91% yield with a 3a : 4a ratio of 97:3, whereas Na2CO3 and K2CO3 afforded 3a in 96% and > 99% yield, respectively, with no detectable hydrolysis. In contrast, Cs2CO3 afforded 3a in 71% yield with a 3a : 4a ratio of 96:4. This countercation dependence shows that the identity of the alkali-metal cation influences the efficiency and selectivity of the carbonate-mediated acyl-transfer process, providing experimental support for cation participation in the reaction.
Consistent with the experimentally observed countercation effect and previous reports of alkali-metal coordination to anhydride-type species[41], the unusual reactivity arises from the cooperative organization of an acyl fluoride, a potassium ion (derived from K2CO3), and water, which activates the acyl fluoride toward nucleophilic attack by carbonate despite the intrinsically lower nucleophilicity of carbonate relative to hydroxide. Consequently, carbonate functions as a nucleophilic acyl-transfer partner to generate a carbonic mixed anhydride (A). DFT calculations further suggest that the carbonic mixed anhydride can associate with K+ in a water-solvated coordination environment, providing a structural model consistent with the experimentally observed cation effect. To experimentally verify the formation of this transient intermediate, in situ 1H NMR spectroscopy was employed (Fig. 5). Owing to its high reactivity and rapid decomposition during workup, the carbonic mixed anhydride was monitored directly using aliquots withdrawn from the reaction mixture.
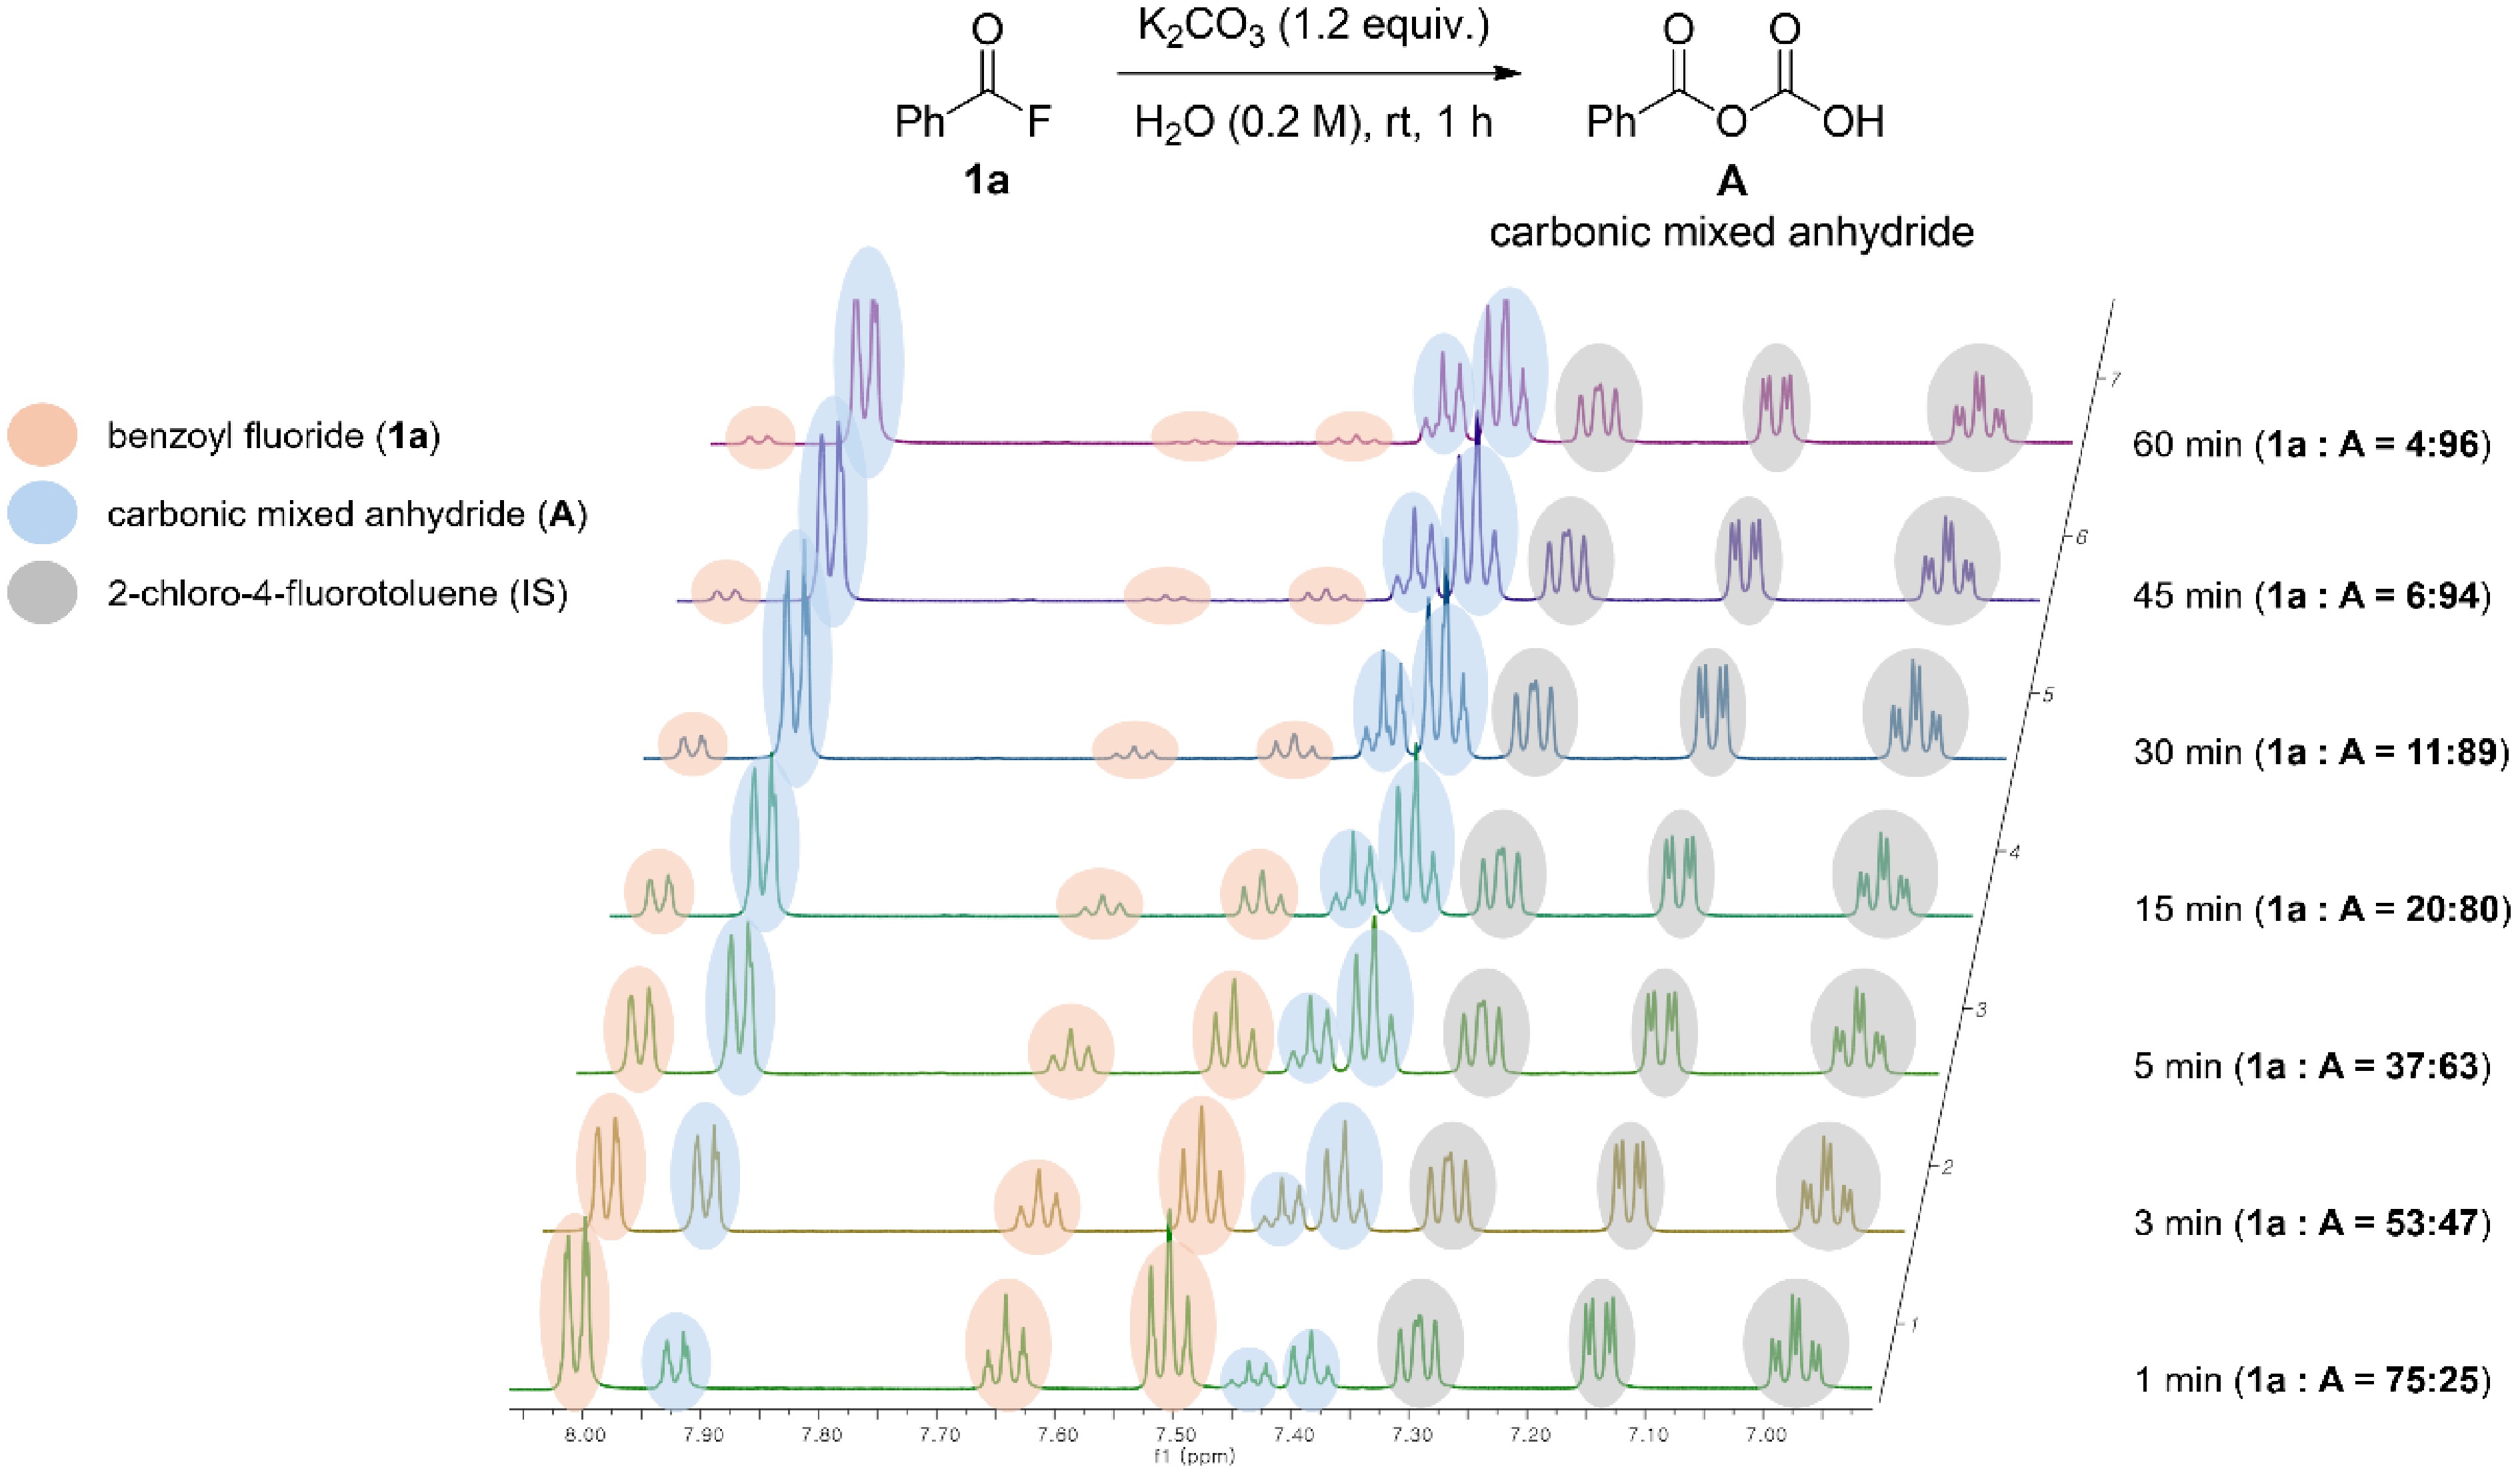
Figure 5.
1H NMR spectroscopic evidence for the formation of the carbonic mixed anhydride intermediate during the carbonate-mediated acyl-transfer reaction.
The appearance of the characteristic 1H NMR signals of the carbonic mixed anhydride, accompanied by the simultaneous disappearance of the acyl fluoride signals, provides direct experimental evidence for its transient formation under the reaction conditions. The formation of this intermediate was further supported by high-resolution mass spectrometry (HRMS), which showed an ion at m/z 189.0168 corresponding to [M + Na]+ (calcd for C8H6O4Na [M + Na]+: 189.0163).
In addition, carbonate-mediated C–F bond cleavage was directly monitored by 19F NMR spectroscopy, which revealed concomitant consumption of the acyl fluoride and formation of KF during the reaction (Fig. 6). These observations provide direct experimental evidence that carbonate participates as a nucleophile in the initial acyl-transfer step. DFT calculations further provide a molecular-level rationale for the proposed activation mode and the subsequent carbonate-mediated acyl-transfer pathway (Fig. 7). The resulting carbonic mixed anhydride exhibits enhanced hydrolytic stability relative to the corresponding acyl halide, thereby favoring productive N-functionalization over competing hydrolysis. Furthermore, because the carbonate-mediated acyl-transfer pathway proceeds without the generation of HX (e.g., HCl or HF), amine nucleophilicity is preserved throughout the reaction, enabling near-quantitative amidation under fully aqueous conditions.
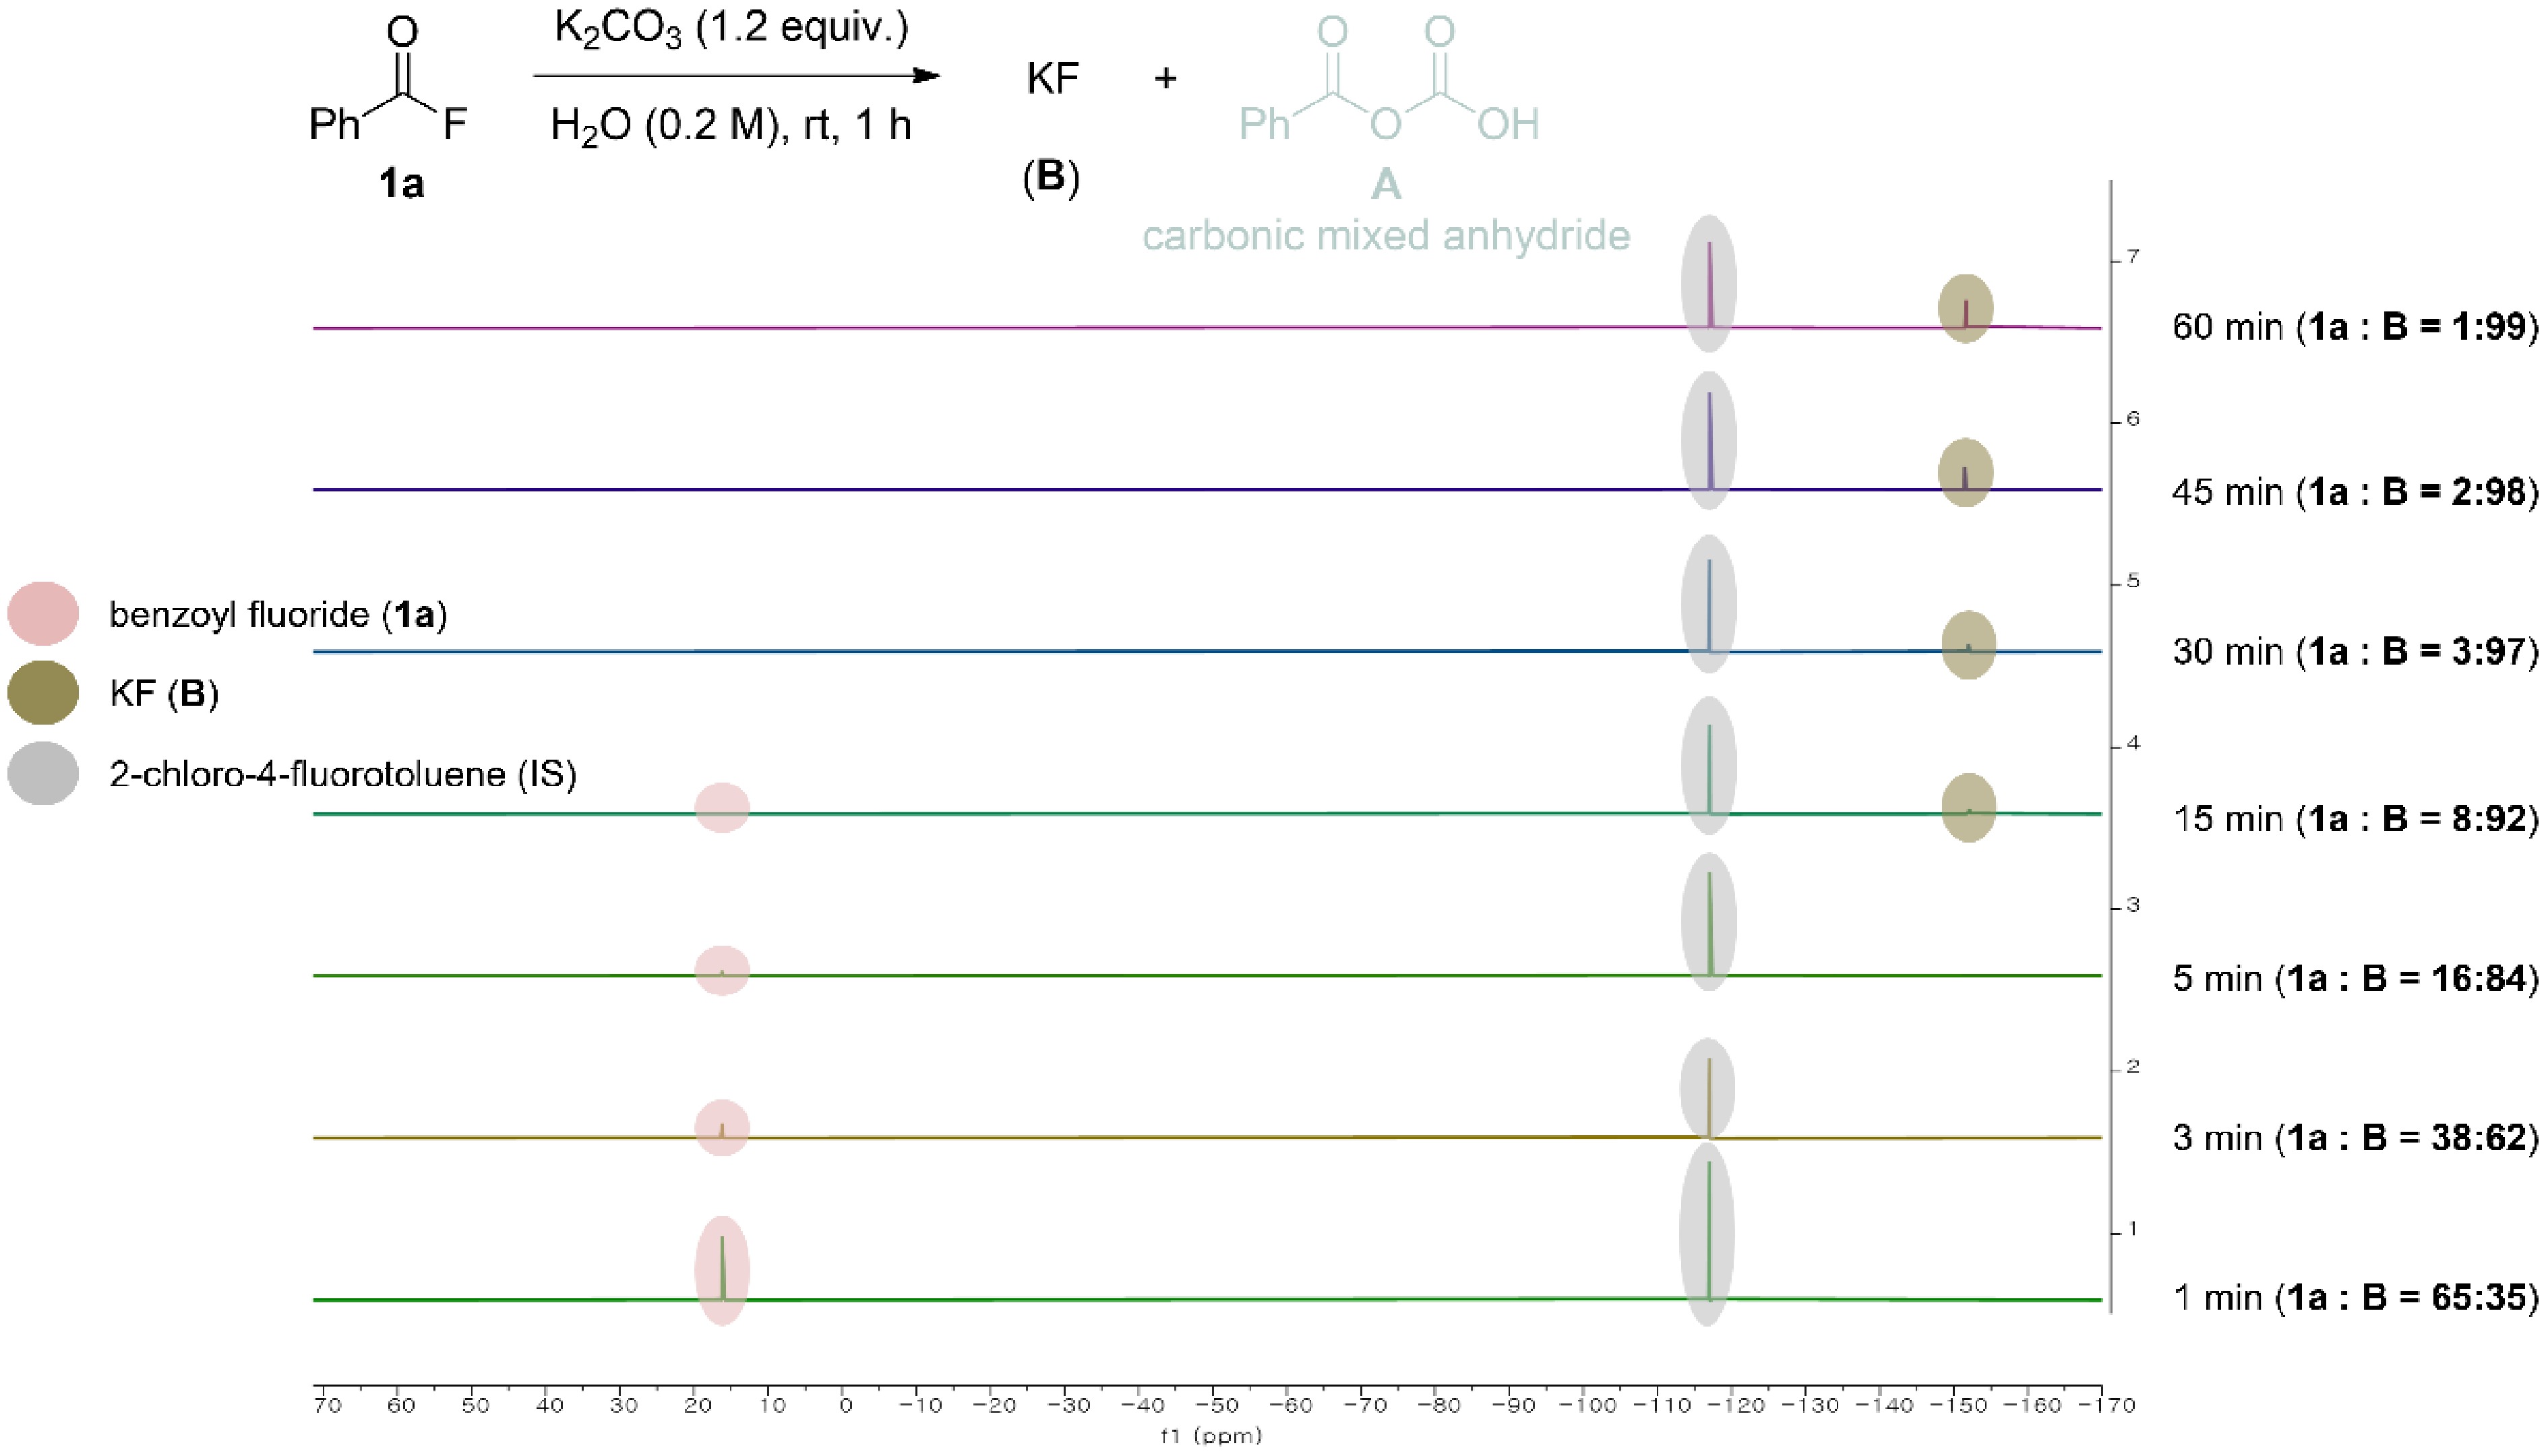
Figure 6.
19F NMR spectroscopic evidence for carbonate-mediated C–F bond cleavage through the concomitant consumption of acyl fluoride and formation of KF during the reaction.
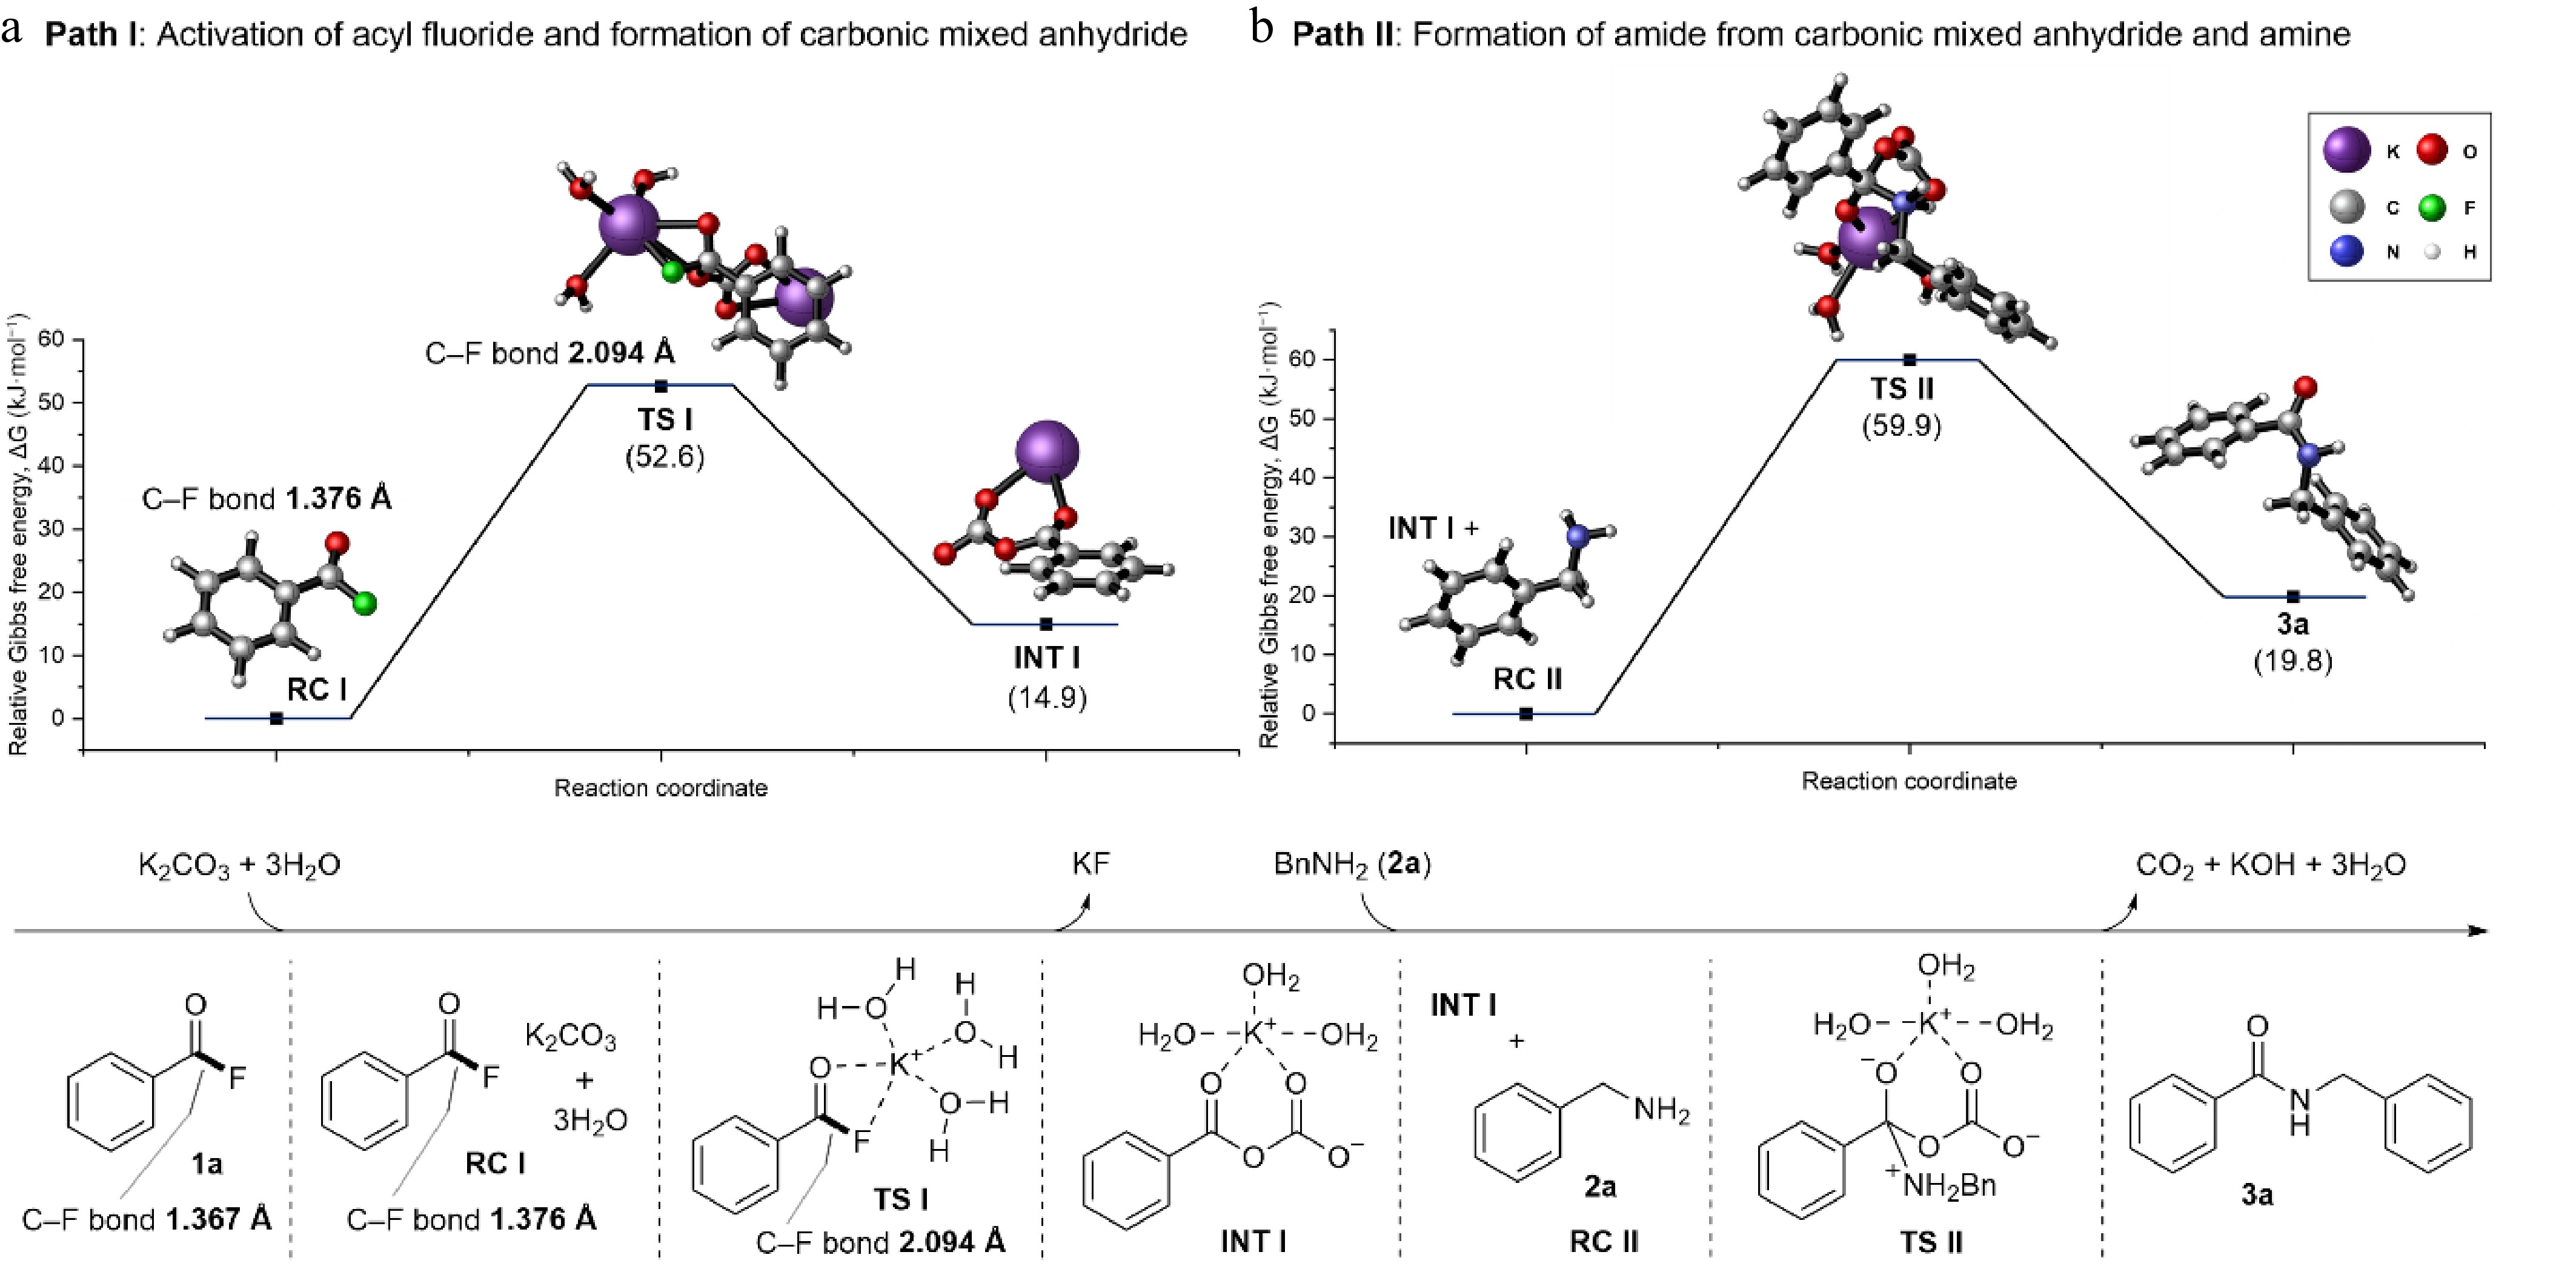
Figure 7.
DFT-calculated Gibbs free-energy profiles and representative optimized structures for the carbonate-mediated amidation of benzoyl fluoride. (a) Path I: carbonate-mediated activation of benzoyl fluoride within the non-classical 18-crown-6–K+ mimic incorporating three explicit water molecules, leading to C–F bond cleavage and formation of the K+–associated carbonic mixed anhydride. (b) Path II: amide formation via nucleophilic acyl substitution of the K+–associated carbonic mixed anhydride by benzylamine.
-
For both 1H and 19F NMR spectroscopic analyses, aliquots were withdrawn directly from the reaction mixture, immediately dissolved in CD3OD containing 2-chloro-4-fluorotoluene as an internal standard (IS), and analyzed without workup or purification.
In situ observation of carbonic mixed anhydride formation by 1H NMR spectroscopy
-
The appearance of characteristic 1H NMR signals attributable to the carbonic mixed anhydride, accompanied by the simultaneous disappearance of the corresponding acyl fluoride signals, confirms its in situ formation under the reaction conditions (Fig. 5).
Direct observation of carbonate-mediated C–F bond cleavage and KF formation by 19F NMR spectroscopy
-
Progressive disappearance of the acyl fluoride resonance at δ 15.5 ppm was accompanied by the simultaneous emergence of a new signal at δ −165.9 ppm, which matched that of an authentic KF (B) reference (Fig. 6). These correlated spectral changes provide direct experimental evidence that carbonate attacks the activated acyl fluoride to initiate C–F bond cleavage, leading to KF formation and revealing its previously unrecognized role as the nucleophile responsible for the initial acyl-transfer event.
-
All density functional theory (DFT) calculations were performed using the ORCA 6.1.1 program package[42]. Geometry optimizations and transition-state searches were carried out using the B3LYP functional in conjunction with Grimme's D3BJ dispersion correction and the def2-SVP basis set[43]. Solvation effects were incorporated using the conductor-like polarizable continuum model (CPCM) with water as the implicit solvent[44]. Harmonic vibrational frequency calculations were performed at the same level of theory to verify the nature of the stationary points and to obtain Gibbs free energy corrections[45].
All local minima were confirmed by the absence of imaginary frequencies, whereas each transition state was characterized by a single imaginary frequency corresponding to the reaction coordinate. The carbonate-mediated activation step and the subsequent amine-addition step were modeled separately to avoid artificial distortions in the calculated energy profile arising from the spectator amine, KF, and water-cluster rearrangements. Three explicit water molecules were incorporated into the carbonate- and potassium-associated structures to represent the local aqueous coordination environment. Relative Gibbs free energies were obtained at the B3LYP-D3BJ/def2-SVP/CPCM (water) level and are reported in kJ·mol−1 relative to the corresponding reactant complex within each mechanistic section. The calculated energy profiles are used to compare relative energetic trends and structural features along the proposed reaction pathway rather than represent method-independent absolute activation barriers. Molecular structures were visualized using CYLview[46].
To gain mechanistic insight into the carbonate-mediated amidation of acyl fluorides, the reaction pathway was divided into two mechanistic stages for DFT analysis. Path I describes the carbonate-mediated activation of benzoyl fluoride through the formation of the non-classical 18-crown-6–K+ mimic incorporating three explicit water molecules, culminating in C–F bond cleavage and the generation of a K+–chelated carbonic mixed anhydride. Path II describes the subsequent nucleophilic attack of benzylamine on the K+–chelated carbonic mixed anhydride, leading to the amide product accompanied by the release of CO2, KOH, and H2O. This separate treatment allows the energetics of each elementary process to be evaluated relative to its own reactant complex, thereby avoiding artificial energy distortions arising from spectator species and cluster rearrangements. The resulting Gibbs free-energy profiles together with representative optimized structures are summarized in Fig. 7.
The DFT calculations reveal that carbonate-mediated activation of benzoyl fluoride within the non-classical 18-crown-6–K+ mimic proceeds via TS I to generate the K+-associated carbonic mixed anhydride, which serves as the key intermediate in the proposed carbonate-driven acyl-transfer pathway (Fig. 7a). TS I is located 52.6 kJ·mol−1 above the reactant complex, corresponding to the activation barrier for carbonate-mediated C–F bond cleavage. Structural analysis of the optimized geometries along the reaction coordinate provides further insight into the activation process. In the reactant complex (RC I), the calculated C–F bond length of benzoyl fluoride is 1.376 Å, essentially identical to that of free benzoyl fluoride (1.367 Å), indicating that little or no C–F bond activation occurs prior to carbonate attack. At TS I, however, the C–F bond is substantially elongated to 2.094 Å. This marked elongation reflects significant activation of the acyl fluoride within the cooperative acyl fluoride–K+–H2O assembly of the non-classical 18-crown-6–K+ mimic, accompanied by increased electrophilicity of the carbonyl carbon. Consequently, despite the resonance-stabilized and inherently weak nucleophilic character of carbonate, the activated acyl fluoride undergoes efficient carbonate-mediated acyl transfer to generate the K+–associated carbonic mixed anhydride. These results provide computational evidence that the non-classical assembly enables a distinct carbonate-driven acyl-transfer pathway through concerted C–F bond activation and carbonyl polarization.
The subsequent amide-forming step was investigated in Path II (Fig. 7b), where the K+-associated carbonic mixed anhydride generated in Path I serves as the electrophilic acyl-transfer intermediate. In the reactant complex (RC II), benzylamine is positioned in close proximity to the acyl carbon of INT I, providing a preorganized geometry for nucleophilic attack. TS II is located 59.9 kJ·mol−1 above RC II and corresponds to the C–N bond-forming transition state. The relatively high free-energy barrier reflects the energetic demand associated with the tetrahedral transition-state geometry required for nucleophilic acyl substitution. Following passage through TS II, the reaction proceeds directly to the desired amide product 3a with concomitant release of CO2, KOH, and H2O. The product state lies 19.8 kJ·mol−1 above RC II but 40.1 kJ·mol−1 below TS II, demonstrating that the reaction proceeds downhill in free energy after crossing the C–N bond-forming transition state. Together with the Path I calculations, these results support a stepwise carbonate-driven amidation mechanism in which potassium carbonate first converts the acyl fluoride into a K+–associated carbonic mixed anhydride, which subsequently undergoes nucleophilic acyl substitution by benzylamine 2a to furnish the amide product 3a.
-
In summary, we have developed a sustainable aqueous amidation protocol based on carbonate-mediated acyl transfer of acyl fluorides and elucidated its kinetic and mechanistic origin through reaction kinetics, NMR spectroscopy, control experiments, HRMS, and DFT calculations. Rather than proceeding through the conventional hydroxide-mediated pathway characteristic of the classical Schotten–Baumann reaction, the transformation operates through a non-classical acyl-transfer mechanism initiated by the cooperative organization of an acyl fluoride, a potassium ion (derived from K2CO3), and water. This unique activation mode enables carbonate to participate directly in acyl transfer, leading to the formation of a hydrolytically stable carbonic mixed anhydride while avoiding HX generation. The combination of enhanced resistance toward hydrolysis and preservation of amine nucleophilicity enables productive N-functionalization to consistently outcompete competing hydrolysis under fully aqueous conditions. Beyond providing a molecular-level rationale for the unusual reactivity of acyl fluorides in water, this work establishes a mechanistic foundation for the rational design of selective aqueous acyl-transfer chemistry and expands the conceptual boundaries of classical Schotten–Baumann methodology. Ongoing studies in our laboratory are directed toward applying this non-classical acyl-transfer strategy to a broader range of bond-forming reactions and sustainable synthetic transformations in aqueous media.
-
The authors confirm their contributions to the paper as follows: experiments, data analysis, writing – original draft: Kim GH; manuscript review and editing, conceptualization, project supervision, funding acquisition: Yang JW. All authors reviewed the results and approved the final version of the manuscript.
-
All data generated or analyzed during this study are included in this published article and its supplementary information files.
-
The authors declare that they have no conflict of interest.
-
accompanies this paper online at: https://doi.org/10.48130/prkm-0026-0023.
- The supplementary files can be downloaded from here.
- Supplmentary File 1
- Copyright: © 2026 by the author(s). Published by Maximum Academic Press, Fayetteville, GA. This article is an open access article distributed under Creative Commons Attribution License (CC BY 4.0), visit https://creativecommons.org/licenses/by/4.0/.
-
About this article
Cite this article
Kim GH, Yang JW. 2026. A non-classical pathway for selective N-functionalization in water: kinetic and mechanistic evidence beyond Schotten–Baumann reaction. Progress in Reaction Kinetics and Mechanism 51: e029 doi: 10.48130/prkm-0026-0023 
A non-classical pathway for selective N-functionalization in water: kinetic and mechanistic evidence beyond Schotten–Baumann reaction
- Received: 15 July 2026
- Revised: 26 August 2026
- Accepted: 09 September 2026
- Published online: 03 November 2026
Abstract: Efficient and selective amidation in water remains challenging because productive N-acylation competes with hydrolysis of the acylating agent. Although acyl fluorides exhibit greater hydrolytic stability than acyl chlorides, the mechanistic origins of their unusual reactivity and selectivity in water remain poorly understood. Combined kinetic, spectroscopic, and computational investigations reveal a previously unknown acyl-transfer pathway arising from the cooperative assembly of an acyl fluoride, a potassium ion (derived from K2CO3), and water, generating a water-solvated mimic of the 18-crown-6–K+ complex that fundamentally alters acyl-transfer reactivity under aqueous conditions. This activation mode enables carbonate-mediated formation of a carbonic mixed anhydride, whose enhanced resistance to hydrolysis relative to the corresponding acyl halide decisively favors productive N-functionalization over competing hydrolysis. Under the optimized K2CO3-mediated conditions, the model amidation afforded the amide in > 99% yield within 1 h, with no detectable hydrolysis product (> 99:n.d.), whereas the classical biphasic Schotten–Baumann reaction using benzoyl chloride and KOH in CH2Cl2/H2O afforded the corresponding amide in 76% yield with an amide-to-hydrolysis-product ratio of 96:4. The resulting mechanism enables highly selective amidation under fully aqueous conditions and reveals a previously unrecognized mode of aqueous acyl transfer. Beyond elucidating the origins of acyl fluoride reactivity in water, this work establishes a kinetic and mechanistic foundation for the rational design of sustainable acylation chemistry and reveals a non-classical acyl-transfer mechanism that transcends the inherent limitations of classical Schotten–Baumann methodology.